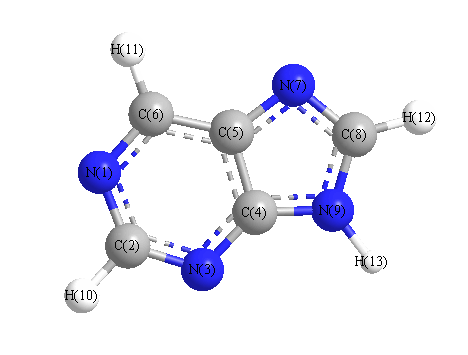
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3930 |
3555 |
105.57 |
|
|
|
| 2 |
A' |
3419 |
3092 |
5.73 |
|
|
|
| 3 |
A' |
3378 |
3055 |
28.08 |
|
|
|
| 4 |
A' |
3363 |
3042 |
22.33 |
|
|
|
| 5 |
A' |
1828 |
1653 |
122.89 |
|
|
|
| 6 |
A' |
1795 |
1624 |
168.39 |
|
|
|
| 7 |
A' |
1712 |
1548 |
43.92 |
|
|
|
| 8 |
A' |
1623 |
1468 |
11.22 |
|
|
|
| 9 |
A' |
1562 |
1413 |
31.75 |
|
|
|
| 10 |
A' |
1553 |
1405 |
108.43 |
|
|
|
| 11 |
A' |
1490 |
1347 |
77.20 |
|
|
|
| 12 |
A' |
1424 |
1288 |
28.31 |
|
|
|
| 13 |
A' |
1403 |
1269 |
6.67 |
|
|
|
| 14 |
A' |
1308 |
1183 |
5.89 |
|
|
|
| 15 |
A' |
1260 |
1139 |
44.90 |
|
|
|
| 16 |
A' |
1223 |
1106 |
11.93 |
|
|
|
| 17 |
A' |
1162 |
1051 |
30.27 |
|
|
|
| 18 |
A' |
1022 |
924 |
1.16 |
|
|
|
| 19 |
A' |
988 |
894 |
18.63 |
|
|
|
| 20 |
A' |
869 |
786 |
21.92 |
|
|
|
| 21 |
A' |
713 |
645 |
0.07 |
|
|
|
| 22 |
A' |
615 |
556 |
3.17 |
|
|
|
| 23 |
A' |
480 |
434 |
13.41 |
|
|
|
| 24 |
A" |
1129 |
1021 |
0.40 |
|
|
|
| 25 |
A" |
1068 |
966 |
5.30 |
|
|
|
| 26 |
A" |
1021 |
923 |
3.12 |
|
|
|
| 27 |
A" |
902 |
816 |
17.52 |
|
|
|
| 28 |
A" |
721 |
652 |
16.96 |
|
|
|
| 29 |
A" |
672 |
608 |
21.34 |
|
|
|
| 30 |
A" |
559 |
505 |
126.87 |
|
|
|
| 31 |
A" |
464 |
420 |
17.39 |
|
|
|
| 32 |
A" |
277 |
251 |
0.06 |
|
|
|
| 33 |
A" |
254 |
229 |
7.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22592.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 20432.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.420 |
|
|
|
| 2 |
C |
0.111 |
|
|
|
| 3 |
N |
-0.451 |
|
|
|
| 4 |
C |
0.587 |
|
|
|
| 5 |
C |
-0.040 |
|
|
|
| 6 |
C |
0.071 |
|
|
|
| 7 |
N |
-0.428 |
|
|
|
| 8 |
C |
0.165 |
|
|
|
| 9 |
N |
-0.716 |
|
|
|
| 10 |
H |
0.227 |
|
|
|
| 11 |
H |
0.249 |
|
|
|
| 12 |
H |
0.246 |
|
|
|
| 13 |
H |
0.398 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.256 |
2.934 |
0.000 |
3.701 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.226 |
6.428 |
0.000 |
| y |
6.428 |
-47.538 |
0.000 |
| z |
0.000 |
0.000 |
-52.418 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.248 |
6.428 |
0.000 |
| y |
6.428 |
3.784 |
0.000 |
| z |
0.000 |
0.000 |
-3.536 |
|
| Polar |
|---|
| 3z2-r2 | -7.072 |
|---|
| x2-y2 | -2.688 |
|---|
| xy | 6.428 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.441 |
1.346 |
0.000 |
| y |
1.346 |
11.005 |
0.000 |
| z |
0.000 |
0.000 |
4.879 |
<r2> (average value of r
2) Å
2
| <r2> |
249.783 |
| (<r2>)1/2 |
15.805 |