return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: MP2=FULL/6-311G*

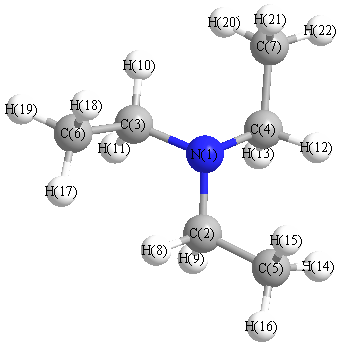
19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-291.570191
Energy at 298.15K-291.587567
HF Energy-290.424906
Nuclear repulsion energy337.344328
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3184 3015 36.11      
2 A 3168 3001 7.27      
3 A 3136 2970 0.56      
4 A 3083 2920 7.05      
5 A 2987 2829 190.61      
6 A 1552 1470 16.97      
7 A 1530 1449 0.98      
8 A 1525 1445 1.64      
9 A 1449 1372 22.47      
10 A 1429 1354 0.01      
11 A 1346 1275 4.85      
12 A 1200 1136 10.58      
13 A 1115 1056 1.38      
14 A 1061 1005 10.04      
15 A 832 788 0.23      
16 A 773 732 11.98      
17 A 459 434 3.19      
18 A 311 295 0.86      
19 A 205 195 0.49      
20 A 111 105 0.01      
21 E 3185 3017 28.34      
22 E 3171 3003 42.52      
23 E 3136 2970 21.84      
24 E 3083 2920 26.62      
25 E 2979 2822 13.36      
26 E 1550 1468 9.56      
26 E 1550 1468 9.56      
27 E 1539 1458 1.96      
27 E 1539 1458 1.96      
28 E 1524 1444 4.04      
28 E 1524 1444 4.04      
29 E 1447 1371 22.13      
30 E 1417 1342 4.11      
30 E 1417 1342 4.11      
31 E 1356 1284 17.64      
32 E 1264 1197 20.84      
33 E 1133 1073 11.26      
34 E 1118 1059 18.59      
35 E 953 903 1.30      
35 E 953 903 1.30      
36 E 814 771 3.00      
36 E 814 771 3.00      
37 E 484 458 2.51      
37 E 484 458 2.51      
38 E 327 309 0.50      
38 E 327 309 0.50      
39 E 221 210 0.06      
39 E 221 210 0.06      
40 E 99 93 0.24      
40 E 99 93 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 46028.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 43593.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.07750 0.07750 0.04465

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.008
C2 0.328 1.351 -0.452
C3 1.006 -0.959 -0.452
C4 -1.334 -0.391 -0.452
C5 -0.281 2.425 0.438
C6 2.241 -0.969 0.438
C7 -1.960 -1.456 0.438
H8 1.415 1.461 -0.433
H9 0.018 1.492 -1.504
H10 0.557 -1.956 -0.433
H11 1.283 -0.762 -1.504
H12 -1.972 0.495 -0.433
H13 -1.301 -0.730 -1.504
H14 -1.372 2.375 0.452
H15 0.076 2.301 1.462
H16 0.000 3.422 0.086
H17 2.742 0.001 0.452
H18 1.954 -1.216 1.462
H19 2.963 -1.711 0.086
H20 -1.371 -2.375 0.452
H21 -2.030 -1.084 1.462
H22 -2.963 -1.711 0.086

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.46371.46371.46372.47912.47912.47912.08092.12392.08092.12392.08092.12392.77812.72273.42262.77812.72273.42262.77812.72273.4226
C21.46372.40732.40731.52233.13563.72861.09271.10583.31442.54602.45442.84472.18062.15112.16482.91013.59094.07514.19333.89264.5271
C31.46372.40732.40733.72861.52233.13562.45442.84471.09271.10583.31442.54604.19333.89264.52712.18062.15112.16482.91013.59094.0751
C41.46372.40732.40733.13563.72861.52233.31442.54602.45442.84471.09271.10582.91013.59094.07514.19333.89264.52712.18062.15112.1648
C52.47911.52233.72863.13564.22884.22882.13692.17494.54484.04672.71023.84321.09191.09141.09363.87554.39375.26854.92264.05294.9423
C62.47913.13561.52233.72864.22884.22882.71023.84322.13692.17494.54484.04674.92264.05294.94231.09191.09141.09363.87554.39375.2685
C72.47913.72863.13561.52234.22884.22884.54484.04672.71023.84322.13692.17493.87554.39375.26854.92264.05294.94231.09191.09141.0936
H82.08091.09272.45443.31442.13692.71024.54481.76023.52242.47043.52243.65053.06362.46772.47342.16273.32373.56724.82264.68395.4312
H92.12391.10582.84472.54602.17493.84324.04671.76023.65052.58502.47042.58502.55663.07422.49993.67054.45854.63264.55114.43064.6558
H102.08093.31441.09272.45444.54482.13692.71023.52243.65051.76023.52242.47044.82264.68395.43123.06362.46772.47342.16273.32373.5672
H112.12392.54601.10582.84474.04672.17493.84322.47042.58501.76023.65052.58504.55114.43064.65582.55663.07422.49993.67054.45854.6326
H122.08092.45443.31441.09272.71024.54482.13693.52242.47043.52243.65051.76022.16273.32373.56724.82264.68395.43123.06362.46772.4734
H132.12392.84472.54601.10583.84324.04672.17493.65052.58502.47042.58501.76023.67054.45854.63264.55114.43064.65582.55663.07422.4999
H142.77812.18064.19332.91011.09194.92263.87553.06362.55664.82264.55112.16273.67051.76641.76424.74984.99755.96804.74983.66304.3999
H152.72272.15113.89263.59091.09144.05294.39372.46773.07424.68394.43063.32374.45851.76641.77653.66303.98695.13044.99753.98695.2176
H163.42262.16484.52714.07511.09364.94235.26852.47342.49995.43124.65583.56724.63261.76421.77654.39995.21765.92665.96805.13045.9266
H172.77812.91012.18064.19333.87551.09194.92262.16273.67053.06362.55664.82264.55114.74983.66304.39991.76641.76424.74984.99755.9680
H182.72273.59092.15113.89264.39371.09144.05293.32374.45852.46773.07424.68394.43064.99753.98695.21761.76641.77653.66303.98695.1304
H193.42264.07512.16484.52715.26851.09364.94233.56724.63262.47342.49995.43124.65585.96805.13045.92661.76421.77654.39995.21765.9266
H202.77814.19332.91012.18064.92263.87551.09194.82264.55112.16273.67053.06362.55664.74984.99755.96804.74983.66304.39991.76641.7642
H212.72273.89263.59092.15114.05294.39371.09144.68394.43063.32374.45852.46773.07423.66303.98695.13044.99753.98695.21761.76641.7765
H223.42264.52714.07512.16484.94235.26851.09365.43124.65583.56724.63262.47342.49994.39995.21765.92665.96805.13045.92661.76421.7765

picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.232 N1 C2 H8 108.092
N1 C2 H9 110.728 N1 C3 C6 112.232
N1 C3 H10 108.092 N1 C3 H11 110.728
N1 C4 C7 112.232 N1 C4 H12 108.092
N1 C4 H13 110.728 C2 N1 C3 110.633
C2 N1 C4 110.633 C2 C5 H14 111.997
C2 C5 H15 109.675 C2 C5 H16 110.622
C3 N1 C4 110.633 C3 C6 H17 111.997
C3 C6 H18 109.675 C3 C6 H19 110.622
C4 C7 H20 111.997 C4 C7 H21 109.674
C4 C7 H22 110.622 C5 C2 H8 108.483
C5 C2 H9 110.696 C6 C3 H10 108.483
C6 C3 H11 110.696 C7 C4 H12 108.483
C7 C4 H13 110.696 H8 C2 H9 106.374
H10 C3 H11 106.374 H12 C4 H13 106.374
H14 C5 H15 108.011 H14 C5 H16 107.647
H15 C5 H16 108.791 H17 C6 H18 108.011
H17 C6 H19 107.647 H18 C6 H19 108.791
H20 C7 H21 108.011 H20 C7 H22 107.647
H21 C7 H22 108.791
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability