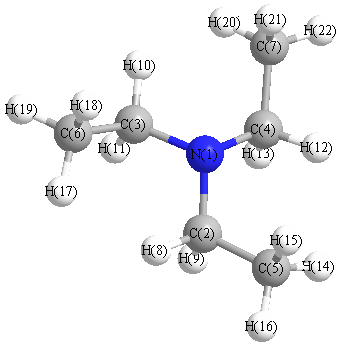
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3137 |
3010 |
65.17 |
|
|
|
| 2 |
A |
3122 |
2996 |
0.44 |
|
|
|
| 3 |
A |
3096 |
2971 |
0.02 |
|
|
|
| 4 |
A |
3053 |
2930 |
10.71 |
|
|
|
| 5 |
A |
2941 |
2822 |
234.12 |
|
|
|
| 6 |
A |
1529 |
1467 |
16.14 |
|
|
|
| 7 |
A |
1504 |
1443 |
3.91 |
|
|
|
| 8 |
A |
1498 |
1438 |
0.30 |
|
|
|
| 9 |
A |
1432 |
1375 |
16.61 |
|
|
|
| 10 |
A |
1409 |
1352 |
1.82 |
|
|
|
| 11 |
A |
1335 |
1281 |
5.37 |
|
|
|
| 12 |
A |
1179 |
1131 |
12.01 |
|
|
|
| 13 |
A |
1100 |
1056 |
2.14 |
|
|
|
| 14 |
A |
1050 |
1008 |
9.77 |
|
|
|
| 15 |
A |
817 |
784 |
0.08 |
|
|
|
| 16 |
A |
765 |
734 |
12.11 |
|
|
|
| 17 |
A |
442 |
424 |
3.13 |
|
|
|
| 18 |
A |
305 |
293 |
1.06 |
|
|
|
| 19 |
A |
192 |
184 |
0.58 |
|
|
|
| 20 |
A |
93 |
89 |
0.03 |
|
|
|
| 21 |
E |
3137 |
3010 |
27.14 |
|
|
|
| 21 |
E |
3137 |
3010 |
27.16 |
|
|
|
| 22 |
E |
3126 |
3000 |
75.95 |
|
|
|
| 22 |
E |
3126 |
3000 |
75.94 |
|
|
|
| 23 |
E |
3095 |
2970 |
17.81 |
|
|
|
| 23 |
E |
3095 |
2970 |
17.80 |
|
|
|
| 24 |
E |
3053 |
2930 |
32.00 |
|
|
|
| 24 |
E |
3053 |
2930 |
32.00 |
|
|
|
| 25 |
E |
2929 |
2810 |
18.67 |
|
|
|
| 25 |
E |
2929 |
2810 |
18.67 |
|
|
|
| 26 |
E |
1523 |
1462 |
8.46 |
|
|
|
| 26 |
E |
1523 |
1462 |
8.46 |
|
|
|
| 27 |
E |
1514 |
1453 |
3.68 |
|
|
|
| 27 |
E |
1514 |
1453 |
3.68 |
|
|
|
| 28 |
E |
1498 |
1438 |
2.12 |
|
|
|
| 28 |
E |
1498 |
1438 |
2.12 |
|
|
|
| 29 |
E |
1429 |
1372 |
27.16 |
|
|
|
| 29 |
E |
1429 |
1372 |
27.16 |
|
|
|
| 30 |
E |
1403 |
1346 |
0.57 |
|
|
|
| 30 |
E |
1403 |
1346 |
0.57 |
|
|
|
| 31 |
E |
1342 |
1288 |
24.03 |
|
|
|
| 31 |
E |
1342 |
1288 |
24.03 |
|
|
|
| 32 |
E |
1252 |
1202 |
24.46 |
|
|
|
| 32 |
E |
1252 |
1202 |
24.46 |
|
|
|
| 33 |
E |
1117 |
1072 |
19.95 |
|
|
|
| 33 |
E |
1117 |
1072 |
19.95 |
|
|
|
| 34 |
E |
1107 |
1062 |
11.22 |
|
|
|
| 34 |
E |
1107 |
1062 |
11.22 |
|
|
|
| 35 |
E |
943 |
905 |
1.09 |
|
|
|
| 35 |
E |
943 |
905 |
1.09 |
|
|
|
| 36 |
E |
802 |
770 |
3.32 |
|
|
|
| 36 |
E |
802 |
770 |
3.32 |
|
|
|
| 37 |
E |
480 |
460 |
2.33 |
|
|
|
| 37 |
E |
480 |
460 |
2.33 |
|
|
|
| 38 |
E |
313 |
300 |
0.38 |
|
|
|
| 38 |
E |
313 |
300 |
0.38 |
|
|
|
| 39 |
E |
216 |
207 |
0.04 |
|
|
|
| 39 |
E |
216 |
207 |
0.04 |
|
|
|
| 40 |
E |
87 |
83 |
0.25 |
|
|
|
| 40 |
E |
87 |
83 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 45365.4 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 43532.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.263 |
|
|
|
| 2 |
C |
-0.355 |
|
|
|
| 3 |
C |
-0.355 |
|
|
|
| 4 |
C |
-0.355 |
|
|
|
| 5 |
C |
-0.641 |
|
|
|
| 6 |
C |
-0.641 |
|
|
|
| 7 |
C |
-0.641 |
|
|
|
| 8 |
H |
0.223 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.223 |
|
|
|
| 11 |
H |
0.195 |
|
|
|
| 12 |
H |
0.223 |
|
|
|
| 13 |
H |
0.195 |
|
|
|
| 14 |
H |
0.219 |
|
|
|
| 15 |
H |
0.228 |
|
|
|
| 16 |
H |
0.218 |
|
|
|
| 17 |
H |
0.219 |
|
|
|
| 18 |
H |
0.228 |
|
|
|
| 19 |
H |
0.218 |
|
|
|
| 20 |
H |
0.219 |
|
|
|
| 21 |
H |
0.228 |
|
|
|
| 22 |
H |
0.218 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.554 |
0.554 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.627 |
0.000 |
0.000 |
| y |
0.000 |
-46.627 |
0.000 |
| z |
0.000 |
0.000 |
-49.262 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.317 |
0.000 |
0.000 |
| y |
0.000 |
1.317 |
0.000 |
| z |
0.000 |
0.000 |
-2.635 |
|
| Polar |
|---|
| 3z2-r2 | -5.270 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.248 |
0.000 |
0.000 |
| y |
0.000 |
12.249 |
-0.000 |
| z |
0.000 |
-0.000 |
10.605 |
<r2> (average value of r
2) Å
2
| <r2> |
286.423 |
| (<r2>)1/2 |
16.924 |