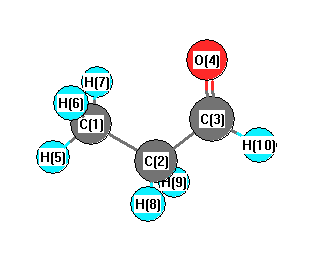
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3129 |
3012 |
24.08 |
|
|
|
| 2 |
A' |
3057 |
2943 |
20.12 |
|
|
|
| 3 |
A' |
3018 |
2906 |
29.93 |
|
|
|
| 4 |
A' |
2879 |
2771 |
159.79 |
|
|
|
| 5 |
A' |
1835 |
1767 |
159.43 |
|
|
|
| 6 |
A' |
1514 |
1457 |
10.28 |
|
|
|
| 7 |
A' |
1460 |
1406 |
17.28 |
|
|
|
| 8 |
A' |
1430 |
1377 |
19.54 |
|
|
|
| 9 |
A' |
1416 |
1363 |
1.27 |
|
|
|
| 10 |
A' |
1372 |
1321 |
12.96 |
|
|
|
| 11 |
A' |
1119 |
1077 |
14.88 |
|
|
|
| 12 |
A' |
1008 |
971 |
0.89 |
|
|
|
| 13 |
A' |
861 |
829 |
22.72 |
|
|
|
| 14 |
A' |
678 |
652 |
5.48 |
|
|
|
| 15 |
A' |
254 |
245 |
8.82 |
|
|
|
| 16 |
A" |
3132 |
3016 |
25.23 |
|
|
|
| 17 |
A" |
3042 |
2929 |
11.70 |
|
|
|
| 18 |
A" |
1507 |
1451 |
9.72 |
|
|
|
| 19 |
A" |
1287 |
1239 |
0.15 |
|
|
|
| 20 |
A" |
1146 |
1103 |
0.34 |
|
|
|
| 21 |
A" |
906 |
872 |
2.43 |
|
|
|
| 22 |
A" |
668 |
643 |
4.70 |
|
|
|
| 23 |
A" |
233 |
225 |
0.54 |
|
|
|
| 24 |
A" |
136 |
131 |
1.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18543.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 17851.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.651 |
|
|
|
| 2 |
C |
-0.525 |
|
|
|
| 3 |
C |
0.127 |
|
|
|
| 4 |
O |
-0.287 |
|
|
|
| 5 |
H |
0.224 |
|
|
|
| 6 |
H |
0.235 |
|
|
|
| 7 |
H |
0.235 |
|
|
|
| 8 |
H |
0.239 |
|
|
|
| 9 |
H |
0.239 |
|
|
|
| 10 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.079 |
2.609 |
0.000 |
2.610 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.119 |
-1.590 |
0.000 |
| y |
-1.590 |
-28.307 |
0.000 |
| z |
0.000 |
0.000 |
-24.187 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.128 |
-1.590 |
0.000 |
| y |
-1.590 |
-4.154 |
0.000 |
| z |
0.000 |
0.000 |
2.026 |
|
| Polar |
|---|
| 3z2-r2 | 4.052 |
|---|
| x2-y2 | 4.188 |
|---|
| xy | -1.590 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.862 |
-0.043 |
0.000 |
| y |
-0.043 |
5.945 |
0.000 |
| z |
0.000 |
0.000 |
4.224 |
<r2> (average value of r
2) Å
2
| <r2> |
84.628 |
| (<r2>)1/2 |
9.199 |