return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CHO (Propanal)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-192.669047
Energy at 298.15K-192.675460
HF Energy-191.992416
Nuclear repulsion energy118.517476
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3189 3020 18.13      
2 A' 3099 2935 16.23      
3 A' 3070 2907 22.16      
4 A' 2951 2795 140.72      
5 A' 1787 1692 93.71      
6 A' 1541 1459 10.67      
7 A' 1498 1419 16.17      
8 A' 1459 1382 14.15      
9 A' 1435 1359 2.06      
10 A' 1396 1322 14.13      
11 A' 1141 1081 13.86      
12 A' 1024 970 1.27      
13 A' 884 837 19.56      
14 A' 683 647 5.32      
15 A' 267 253 8.25      
16 A" 3192 3023 18.62      
17 A" 3111 2946 8.97      
18 A" 1538 1457 9.71      
19 A" 1308 1239 0.30      
20 A" 1168 1106 0.73      
21 A" 918 869 1.56      
22 A" 686 650 4.44      
23 A" 246 233 0.28      
24 A" 146 139 2.15      

Unscaled Zero Point Vibrational Energy (zpe) 18867.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17869.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.55404 0.19803 0.15412

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.445 0.447 0.000
C2 0.000 0.920 0.000
C3 -0.998 -0.210 0.000
O4 -0.702 -1.387 0.000
H5 2.132 1.295 0.000
H6 1.652 -0.166 0.879
H7 1.652 -0.166 -0.879
H8 -0.220 1.549 0.872
H9 -0.220 1.549 -0.872
H10 -2.066 0.096 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.52072.52992.82361.09141.09071.09072.17942.17943.5284
C21.52071.50752.41122.16442.16312.16311.09781.09782.2238
C32.52991.50751.21363.47282.79212.79212.11212.11211.1109
O42.82362.41121.21363.90172.79352.79353.10083.10082.0150
H51.09142.16443.47283.90171.77101.77102.52112.52114.3652
H61.09072.16312.79212.79351.77101.75712.53913.08403.8291
H71.09072.16312.79212.79351.77101.75713.08402.53913.8291
H82.17941.09782.11213.10082.52112.53913.08401.74372.5055
H92.17941.09782.11213.10082.52113.08402.53911.74372.5055
H103.52842.22381.11092.01504.36523.82913.82912.50552.5055

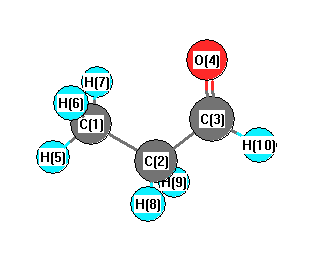
picture of Propanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 113.328 C1 C2 H8 111.660
C1 C2 H9 111.660 C2 C1 H5 110.846
C2 C1 H6 110.783 C2 C1 H7 110.783
C2 C3 O4 124.426 C2 C3 H10 115.441
C3 C2 H8 107.283 C3 C2 H9 107.283
O4 C3 H10 120.134 H5 C1 H6 108.502
H5 C1 H7 108.502 H6 C1 H7 107.309
H8 C2 H9 105.159
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability