Jump to
S1C2
S1C3
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -208.008273 |
| Energy at 298.15K | -208.014359 |
| HF Energy | -208.008273 |
| Nuclear repulsion energy | 120.529528 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3870 |
3500 |
29.10 |
|
|
|
| 2 |
A' |
3276 |
2963 |
40.70 |
|
|
|
| 3 |
A' |
3193 |
2888 |
72.96 |
|
|
|
| 4 |
A' |
3170 |
2867 |
79.13 |
|
|
|
| 5 |
A' |
1973 |
1784 |
670.35 |
|
|
|
| 6 |
A' |
1678 |
1518 |
26.62 |
|
|
|
| 7 |
A' |
1641 |
1484 |
8.35 |
|
|
|
| 8 |
A' |
1612 |
1458 |
6.78 |
|
|
|
| 9 |
A' |
1538 |
1391 |
7.72 |
|
|
|
| 10 |
A' |
1405 |
1271 |
189.21 |
|
|
|
| 11 |
A' |
1266 |
1145 |
52.72 |
|
|
|
| 12 |
A' |
1091 |
987 |
51.37 |
|
|
|
| 13 |
A' |
660 |
597 |
18.24 |
|
|
|
| 14 |
A' |
374 |
338 |
10.37 |
|
|
|
| 15 |
A" |
3243 |
2933 |
56.00 |
|
|
|
| 16 |
A" |
1624 |
1469 |
5.51 |
|
|
|
| 17 |
A" |
1257 |
1137 |
2.21 |
|
|
|
| 18 |
A" |
1175 |
1063 |
1.37 |
|
|
|
| 19 |
A" |
654 |
591 |
169.88 |
|
|
|
| 20 |
A" |
191 |
173 |
0.75 |
|
|
|
| 21 |
A" |
50 |
45 |
1.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17468.9 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 15798.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
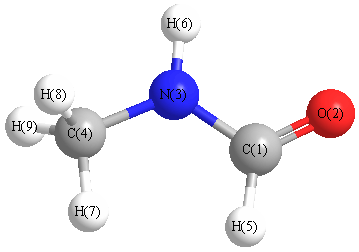
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.291 |
-0.753 |
0.000 |
| O2 |
1.389 |
-1.211 |
0.000 |
| N3 |
0.000 |
0.562 |
0.000 |
| C4 |
-1.330 |
1.120 |
0.000 |
| H5 |
-0.599 |
-1.387 |
0.000 |
| H6 |
0.781 |
1.176 |
0.000 |
| H7 |
-2.055 |
0.317 |
0.000 |
| H8 |
-1.503 |
1.727 |
0.881 |
| H9 |
-1.503 |
1.727 |
-0.881 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.1889 | 1.3470 | 2.4775 | 1.0931 | 1.9903 | 2.5796 | 3.1857 | 3.1857 |
O2 | 1.1889 | | 2.2519 | 3.5811 | 1.9957 | 2.4627 | 3.7681 | 4.2155 | 4.2155 | N3 | 1.3470 | 2.2519 | | 1.4424 | 2.0389 | 0.9937 | 2.0700 | 2.0960 | 2.0960 | C4 | 2.4775 | 3.5811 | 1.4424 | | 2.6115 | 2.1121 | 1.0819 | 1.0839 | 1.0839 | H5 | 1.0931 | 1.9957 | 2.0389 | 2.6115 | | 2.9111 | 2.2419 | 3.3604 | 3.3604 | H6 | 1.9903 | 2.4627 | 0.9937 | 2.1121 | 2.9111 | | 2.9639 | 2.5096 | 2.5096 | H7 | 2.5796 | 3.7681 | 2.0700 | 1.0819 | 2.2419 | 2.9639 | | 1.7519 | 1.7519 | H8 | 3.1857 | 4.2155 | 2.0960 | 1.0839 | 3.3604 | 2.5096 | 1.7519 | | 1.7620 | H9 | 3.1857 | 4.2155 | 2.0960 | 1.0839 | 3.3604 | 2.5096 | 1.7519 | 1.7620 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
125.260 |
|
C1 |
N3 |
H6 |
115.665 |
| O2 |
C1 |
N3 |
125.131 |
|
O2 |
C1 |
H5 |
121.922 |
| N3 |
C1 |
H5 |
112.947 |
|
N3 |
C4 |
H7 |
109.344 |
| N3 |
C4 |
H8 |
111.337 |
|
N3 |
C4 |
H9 |
111.337 |
| C4 |
N3 |
H6 |
119.075 |
|
H7 |
C4 |
H8 |
107.982 |
| H7 |
C4 |
H9 |
107.982 |
|
H8 |
C4 |
H9 |
108.743 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.364 |
|
|
|
| 2 |
O |
-0.471 |
|
|
|
| 3 |
N |
-0.649 |
|
|
|
| 4 |
C |
-0.427 |
|
|
|
| 5 |
H |
0.159 |
|
|
|
| 6 |
H |
0.369 |
|
|
|
| 7 |
H |
0.214 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
| 9 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.352 |
2.695 |
0.000 |
4.301 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.896 |
4.535 |
0.000 |
| y |
4.535 |
-24.361 |
0.000 |
| z |
0.000 |
0.000 |
-24.739 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.346 |
4.535 |
0.000 |
| y |
4.535 |
1.457 |
0.000 |
| z |
0.000 |
0.000 |
0.889 |
|
| Polar |
|---|
| 3z2-r2 | 1.779 |
|---|
| x2-y2 | -2.535 |
|---|
| xy | 4.535 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.075 |
-0.854 |
0.000 |
| y |
-0.854 |
4.978 |
0.000 |
| z |
0.000 |
0.000 |
3.073 |
<r2> (average value of r
2) Å
2
| <r2> |
87.882 |
| (<r2>)1/2 |
9.375 |
Jump to
S1C1
S1C3
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -208.008273 |
| Energy at 298.15K | -208.014359 |
| HF Energy | -208.008273 |
| Nuclear repulsion energy | 120.529528 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -208.009959 |
| Energy at 298.15K | -208.016065 |
| HF Energy | -208.009959 |
| Nuclear repulsion energy | 122.703097 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3906 |
3533 |
29.14 |
|
|
|
| 2 |
A |
3313 |
2996 |
15.33 |
|
|
|
| 3 |
A |
3257 |
2946 |
52.05 |
|
|
|
| 4 |
A |
3193 |
2888 |
48.28 |
|
|
|
| 5 |
A |
3171 |
2868 |
141.53 |
|
|
|
| 6 |
A |
1962 |
1775 |
447.70 |
|
|
|
| 7 |
A |
1708 |
1545 |
196.20 |
|
|
|
| 8 |
A |
1644 |
1487 |
9.62 |
|
|
|
| 9 |
A |
1627 |
1472 |
19.85 |
|
|
|
| 10 |
A |
1591 |
1439 |
14.88 |
|
|
|
| 11 |
A |
1558 |
1409 |
10.31 |
|
|
|
| 12 |
A |
1331 |
1204 |
123.05 |
|
|
|
| 13 |
A |
1279 |
1157 |
2.58 |
|
|
|
| 14 |
A |
1256 |
1136 |
11.27 |
|
|
|
| 15 |
A |
1162 |
1051 |
6.32 |
|
|
|
| 16 |
A |
1024 |
926 |
19.48 |
|
|
|
| 17 |
A |
838 |
758 |
9.23 |
|
|
|
| 18 |
A |
526 |
475 |
46.03 |
|
|
|
| 19 |
A |
311 |
281 |
16.59 |
|
|
|
| 20 |
A |
267 |
241 |
117.43 |
|
|
|
| 21 |
A |
53 |
48 |
0.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17487.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 15815.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.864 |
0.415 |
0.006 |
| O2 |
1.382 |
-0.656 |
-0.000 |
| N3 |
-0.463 |
0.644 |
-0.029 |
| C4 |
-1.436 |
-0.427 |
0.007 |
| H5 |
1.448 |
1.339 |
0.028 |
| H6 |
-0.774 |
1.578 |
0.078 |
| H7 |
-2.350 |
-0.095 |
-0.467 |
| H8 |
-1.049 |
-1.275 |
-0.535 |
| H9 |
-1.659 |
-0.740 |
1.022 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.1895 | 1.3472 | 2.4496 | 1.0928 | 2.0107 | 3.2893 | 2.6098 | 2.9550 |
O2 | 1.1895 | | 2.2566 | 2.8269 | 1.9959 | 3.1057 | 3.8026 | 2.5645 | 3.2088 | N3 | 1.3472 | 2.2566 | | 1.4474 | 2.0342 | 0.9906 | 2.0742 | 2.0696 | 2.1092 | C4 | 2.4496 | 2.8269 | 1.4474 | | 3.3817 | 2.1127 | 1.0821 | 1.0787 | 1.0851 | H5 | 1.0928 | 1.9959 | 2.0342 | 3.3817 | | 2.2359 | 4.0905 | 3.6586 | 3.8681 | H6 | 2.0107 | 3.1057 | 0.9906 | 2.1127 | 2.2359 | | 2.3629 | 2.9315 | 2.6548 | H7 | 3.2893 | 3.8026 | 2.0742 | 1.0821 | 4.0905 | 2.3629 | | 1.7580 | 1.7636 | H8 | 2.6098 | 2.5645 | 2.0696 | 1.0787 | 3.6586 | 2.9315 | 1.7580 | | 1.7558 | H9 | 2.9550 | 3.2088 | 2.1092 | 1.0851 | 3.8681 | 2.6548 | 1.7636 | 1.7558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
122.420 |
|
C1 |
N3 |
H6 |
117.842 |
| O2 |
C1 |
N3 |
125.534 |
|
O2 |
C1 |
H5 |
121.917 |
| N3 |
C1 |
H5 |
112.544 |
|
N3 |
C4 |
H7 |
109.323 |
| N3 |
C4 |
H8 |
109.158 |
|
N3 |
C4 |
H9 |
112.002 |
| C4 |
N3 |
H6 |
118.936 |
|
H7 |
C4 |
H8 |
108.894 |
| H7 |
C4 |
H9 |
108.938 |
|
H8 |
C4 |
H9 |
108.472 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.350 |
|
|
|
| 2 |
O |
-0.473 |
|
|
|
| 3 |
N |
-0.653 |
|
|
|
| 4 |
C |
-0.436 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
H |
0.365 |
|
|
|
| 7 |
H |
0.210 |
|
|
|
| 8 |
H |
0.247 |
|
|
|
| 9 |
H |
0.219 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.841 |
2.943 |
0.192 |
4.095 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.531 |
3.147 |
-0.051 |
| y |
3.147 |
-22.178 |
0.453 |
| z |
-0.051 |
0.453 |
-24.761 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.061 |
3.147 |
-0.051 |
| y |
3.147 |
2.968 |
0.453 |
| z |
-0.051 |
0.453 |
-0.907 |
|
| Polar |
|---|
| 3z2-r2 | -1.813 |
|---|
| x2-y2 | -3.352 |
|---|
| xy | 3.147 |
|---|
| xz | -0.051 |
|---|
| yz | 0.453 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.182 |
-0.185 |
-0.021 |
| y |
-0.185 |
4.629 |
-0.014 |
| z |
-0.021 |
-0.014 |
3.066 |
<r2> (average value of r
2) Å
2
| <r2> |
77.285 |
| (<r2>)1/2 |
8.791 |
