Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -208.714959 |
| Energy at 298.15K | -208.720942 |
| HF Energy | -208.006096 |
| Nuclear repulsion energy | 119.181472 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3668 |
3474 |
29.65 |
|
|
|
| 2 |
A' |
3191 |
3022 |
15.91 |
|
|
|
| 3 |
A' |
3082 |
2919 |
51.48 |
|
|
|
| 4 |
A' |
3003 |
2845 |
91.62 |
|
|
|
| 5 |
A' |
1824 |
1727 |
441.30 |
|
|
|
| 6 |
A' |
1574 |
1491 |
17.77 |
|
|
|
| 7 |
A' |
1517 |
1437 |
12.28 |
|
|
|
| 8 |
A' |
1506 |
1427 |
4.07 |
|
|
|
| 9 |
A' |
1423 |
1348 |
9.66 |
|
|
|
| 10 |
A' |
1332 |
1262 |
110.64 |
|
|
|
| 11 |
A' |
1195 |
1131 |
29.83 |
|
|
|
| 12 |
A' |
1034 |
980 |
48.29 |
|
|
|
| 13 |
A' |
618 |
586 |
13.55 |
|
|
|
| 14 |
A' |
349 |
331 |
7.52 |
|
|
|
| 15 |
A" |
3158 |
2991 |
27.93 |
|
|
|
| 16 |
A" |
1528 |
1447 |
7.75 |
|
|
|
| 17 |
A" |
1174 |
1112 |
0.60 |
|
|
|
| 18 |
A" |
1047 |
992 |
1.56 |
|
|
|
| 19 |
A" |
614 |
581 |
140.63 |
|
|
|
| 20 |
A" |
192 |
182 |
2.22 |
|
|
|
| 21 |
A" |
61 |
58 |
2.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16545.7 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15670.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.283 |
-0.757 |
0.000 |
| O2 |
1.401 |
-1.236 |
0.000 |
| N3 |
0.000 |
0.573 |
0.000 |
| C4 |
-1.333 |
1.135 |
0.000 |
| H5 |
-0.633 |
-1.378 |
0.000 |
| H6 |
0.801 |
1.186 |
0.000 |
| H7 |
-2.055 |
0.318 |
0.000 |
| H8 |
-1.510 |
1.744 |
0.889 |
| H9 |
-1.510 |
1.744 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2166 | 1.3602 | 2.4884 | 1.1062 | 2.0115 | 2.5736 | 3.2038 | 3.2038 |
O2 | 1.2166 | | 2.2885 | 3.6193 | 2.0390 | 2.4959 | 3.7898 | 4.2604 | 4.2604 | N3 | 1.3602 | 2.2885 | | 1.4467 | 2.0511 | 1.0086 | 2.0710 | 2.1079 | 2.1079 | C4 | 2.4884 | 3.6193 | 1.4467 | | 2.6086 | 2.1345 | 1.0903 | 1.0924 | 1.0924 | H5 | 1.1062 | 2.0390 | 2.0511 | 2.6086 | | 2.9379 | 2.2135 | 3.3628 | 3.3628 | H6 | 2.0115 | 2.4959 | 1.0086 | 2.1345 | 2.9379 | | 2.9851 | 2.5383 | 2.5383 | H7 | 2.5736 | 3.7898 | 2.0710 | 1.0903 | 2.2135 | 2.9851 | | 1.7668 | 1.7668 | H8 | 3.2038 | 4.2604 | 2.1079 | 1.0924 | 3.3628 | 2.5383 | 1.7668 | | 1.7785 | H9 | 3.2038 | 4.2604 | 2.1079 | 1.0924 | 3.3628 | 2.5383 | 1.7668 | 1.7785 | |
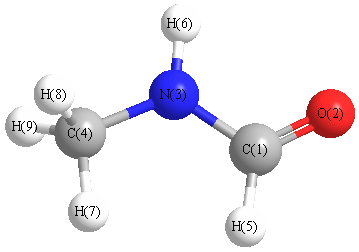 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.855 |
|
C1 |
N3 |
H6 |
115.442 |
| O2 |
C1 |
N3 |
125.185 |
|
O2 |
C1 |
H5 |
122.693 |
| N3 |
C1 |
H5 |
112.122 |
|
N3 |
C4 |
H7 |
108.623 |
| N3 |
C4 |
H8 |
111.472 |
|
N3 |
C4 |
H9 |
111.472 |
| C4 |
N3 |
H6 |
119.703 |
|
H7 |
C4 |
H8 |
108.088 |
| H7 |
C4 |
H9 |
108.088 |
|
H8 |
C4 |
H9 |
108.981 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -208.714959 |
| Energy at 298.15K | -208.720942 |
| HF Energy | -208.006096 |
| Nuclear repulsion energy | 119.181472 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -208.717005 |
| Energy at 298.15K | -208.722974 |
| HF Energy | -208.007699 |
| Nuclear repulsion energy | 121.426285 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3702 |
3506 |
31.29 |
|
|
|
| 2 |
A |
3221 |
3050 |
1.11 |
|
|
|
| 3 |
A |
3165 |
2998 |
29.06 |
|
|
|
| 4 |
A |
3089 |
2926 |
42.24 |
|
|
|
| 5 |
A |
3011 |
2852 |
123.30 |
|
|
|
| 6 |
A |
1811 |
1716 |
308.49 |
|
|
|
| 7 |
A |
1597 |
1513 |
100.23 |
|
|
|
| 8 |
A |
1544 |
1463 |
9.03 |
|
|
|
| 9 |
A |
1536 |
1454 |
39.97 |
|
|
|
| 10 |
A |
1479 |
1401 |
22.34 |
|
|
|
| 11 |
A |
1439 |
1363 |
3.92 |
|
|
|
| 12 |
A |
1251 |
1185 |
77.31 |
|
|
|
| 13 |
A |
1181 |
1119 |
18.74 |
|
|
|
| 14 |
A |
1178 |
1116 |
1.09 |
|
|
|
| 15 |
A |
1026 |
972 |
0.00 |
|
|
|
| 16 |
A |
990 |
937 |
22.68 |
|
|
|
| 17 |
A |
779 |
738 |
0.59 |
|
|
|
| 18 |
A |
528 |
500 |
48.31 |
|
|
|
| 19 |
A |
306 |
290 |
12.54 |
|
|
|
| 20 |
A |
243 |
230 |
95.00 |
|
|
|
| 21 |
A |
41 |
39 |
1.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16558.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15682.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.860 |
0.438 |
0.004 |
| O2 |
1.372 |
-0.671 |
-0.000 |
| N3 |
-0.477 |
0.662 |
-0.022 |
| C4 |
-1.413 |
-0.446 |
0.005 |
| H5 |
1.454 |
1.368 |
0.019 |
| H6 |
-0.812 |
1.606 |
0.063 |
| H7 |
-2.413 |
-0.070 |
-0.206 |
| H8 |
-1.135 |
-1.176 |
-0.755 |
| H9 |
-1.410 |
-0.947 |
0.976 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2207 | 1.3560 | 2.4390 | 1.1038 | 2.0404 | 3.3185 | 2.6761 | 2.8316 |
O2 | 1.2207 | | 2.2790 | 2.7935 | 2.0407 | 3.1551 | 3.8372 | 2.6661 | 2.9612 | N3 | 1.3560 | 2.2790 | | 1.4511 | 2.0563 | 1.0054 | 2.0774 | 2.0854 | 2.1113 | C4 | 2.4390 | 2.7935 | 1.4511 | | 3.3929 | 2.1399 | 1.0890 | 1.0895 | 1.0925 | H5 | 1.1038 | 2.0407 | 2.0563 | 3.3929 | | 2.2782 | 4.1312 | 3.7114 | 3.8055 | H6 | 2.0404 | 3.1551 | 1.0054 | 2.1399 | 2.2782 | | 2.3333 | 2.9181 | 2.7773 | H7 | 3.3185 | 3.8372 | 2.0774 | 1.0890 | 4.1312 | 2.3333 | | 1.7771 | 1.7810 | H8 | 2.6761 | 2.6661 | 2.0854 | 1.0895 | 3.7114 | 2.9181 | 1.7771 | | 1.7675 | H9 | 2.8316 | 2.9612 | 2.1113 | 1.0925 | 3.8055 | 2.7773 | 1.7810 | 1.7675 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.616 |
|
C1 |
N3 |
H6 |
118.803 |
| O2 |
C1 |
N3 |
124.280 |
|
O2 |
C1 |
H5 |
122.693 |
| N3 |
C1 |
H5 |
113.023 |
|
N3 |
C4 |
H7 |
108.906 |
| N3 |
C4 |
H8 |
109.511 |
|
N3 |
C4 |
H9 |
111.428 |
| C4 |
N3 |
H6 |
120.087 |
|
H7 |
C4 |
H8 |
109.319 |
| H7 |
C4 |
H9 |
109.454 |
|
H8 |
C4 |
H9 |
108.198 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
