Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.009271 |
| Energy at 298.15K | -209.015347 |
| HF Energy | -209.009271 |
| Nuclear repulsion energy | 119.635730 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3649 |
3498 |
18.65 |
|
|
|
| 2 |
A' |
3145 |
3015 |
19.39 |
|
|
|
| 3 |
A' |
3048 |
2921 |
58.41 |
|
|
|
| 4 |
A' |
2950 |
2828 |
103.12 |
|
|
|
| 5 |
A' |
1850 |
1773 |
503.01 |
|
|
|
| 6 |
A' |
1555 |
1490 |
24.56 |
|
|
|
| 7 |
A' |
1509 |
1446 |
7.64 |
|
|
|
| 8 |
A' |
1479 |
1417 |
4.22 |
|
|
|
| 9 |
A' |
1407 |
1348 |
8.21 |
|
|
|
| 10 |
A' |
1324 |
1269 |
105.75 |
|
|
|
| 11 |
A' |
1187 |
1138 |
25.30 |
|
|
|
| 12 |
A' |
1026 |
983 |
48.11 |
|
|
|
| 13 |
A' |
622 |
596 |
15.49 |
|
|
|
| 14 |
A' |
346 |
332 |
8.54 |
|
|
|
| 15 |
A" |
3107 |
2978 |
35.30 |
|
|
|
| 16 |
A" |
1502 |
1439 |
8.08 |
|
|
|
| 17 |
A" |
1157 |
1109 |
0.32 |
|
|
|
| 18 |
A" |
1044 |
1001 |
0.43 |
|
|
|
| 19 |
A" |
643 |
616 |
137.29 |
|
|
|
| 20 |
A" |
204 |
196 |
1.16 |
|
|
|
| 21 |
A" |
99 |
95 |
0.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16425.5 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 15743.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.283 |
-0.756 |
0.000 |
| O2 |
1.393 |
-1.232 |
0.000 |
| N3 |
0.000 |
0.569 |
0.000 |
| C4 |
-1.325 |
1.132 |
0.000 |
| H5 |
-0.636 |
-1.377 |
0.000 |
| H6 |
0.799 |
1.185 |
0.000 |
| H7 |
-2.054 |
0.319 |
0.000 |
| H8 |
-1.505 |
1.744 |
0.889 |
| H9 |
-1.505 |
1.744 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2075 | 1.3547 | 2.4799 | 1.1095 | 2.0078 | 2.5727 | 3.1995 | 3.1995 |
O2 | 1.2075 | | 2.2765 | 3.6018 | 2.0343 | 2.4883 | 3.7799 | 4.2476 | 4.2476 | N3 | 1.3547 | 2.2765 | | 1.4394 | 2.0472 | 1.0086 | 2.0688 | 2.1060 | 2.1060 | C4 | 2.4799 | 3.6018 | 1.4394 | | 2.6020 | 2.1240 | 1.0918 | 1.0943 | 1.0943 | H5 | 1.1095 | 2.0343 | 2.0472 | 2.6020 | | 2.9361 | 2.2111 | 3.3596 | 3.3596 | H6 | 2.0078 | 2.4883 | 1.0086 | 2.1240 | 2.9361 | | 2.9808 | 2.5315 | 2.5315 | H7 | 2.5727 | 3.7799 | 2.0688 | 1.0918 | 2.2111 | 2.9808 | | 1.7668 | 1.7668 | H8 | 3.1995 | 4.2476 | 2.1060 | 1.0943 | 3.3596 | 2.5315 | 1.7668 | | 1.7783 | H9 | 3.1995 | 4.2476 | 2.1060 | 1.0943 | 3.3596 | 2.5315 | 1.7668 | 1.7783 | |
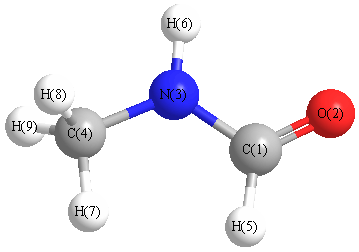 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
125.115 |
|
C1 |
N3 |
H6 |
115.557 |
| O2 |
C1 |
N3 |
125.275 |
|
O2 |
C1 |
H5 |
122.751 |
| N3 |
C1 |
H5 |
111.973 |
|
N3 |
C4 |
H7 |
108.866 |
| N3 |
C4 |
H8 |
111.726 |
|
N3 |
C4 |
H9 |
111.726 |
| C4 |
N3 |
H6 |
119.329 |
|
H7 |
C4 |
H8 |
107.843 |
| H7 |
C4 |
H9 |
107.843 |
|
H8 |
C4 |
H9 |
108.688 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.216 |
|
|
|
| 2 |
O |
-0.368 |
|
|
|
| 3 |
N |
-0.534 |
|
|
|
| 4 |
C |
-0.512 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
| 6 |
H |
0.348 |
|
|
|
| 7 |
H |
0.229 |
|
|
|
| 8 |
H |
0.237 |
|
|
|
| 9 |
H |
0.237 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.963 |
2.798 |
0.000 |
4.075 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.219 |
4.027 |
0.000 |
| y |
4.027 |
-24.233 |
0.000 |
| z |
0.000 |
0.000 |
-24.749 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.729 |
4.027 |
0.000 |
| y |
4.027 |
1.251 |
0.000 |
| z |
0.000 |
0.000 |
0.477 |
|
| Polar |
|---|
| 3z2-r2 | 0.954 |
|---|
| x2-y2 | -1.987 |
|---|
| xy | 4.027 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.604 |
-1.022 |
0.000 |
| y |
-1.022 |
5.650 |
0.000 |
| z |
0.000 |
0.000 |
3.235 |
<r2> (average value of r
2) Å
2
| <r2> |
88.565 |
| (<r2>)1/2 |
9.411 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.009271 |
| Energy at 298.15K | -209.015347 |
| HF Energy | -209.009271 |
| Nuclear repulsion energy | 119.635730 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.011098 |
| Energy at 298.15K | -209.017221 |
| HF Energy | -209.011098 |
| Nuclear repulsion energy | 121.854168 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3683 |
3530 |
18.97 |
|
|
|
| 2 |
A |
3181 |
3049 |
1.59 |
|
|
|
| 3 |
A |
3115 |
2986 |
36.10 |
|
|
|
| 4 |
A |
3055 |
2928 |
47.23 |
|
|
|
| 5 |
A |
2967 |
2844 |
135.31 |
|
|
|
| 6 |
A |
1836 |
1760 |
347.41 |
|
|
|
| 7 |
A |
1588 |
1523 |
114.36 |
|
|
|
| 8 |
A |
1518 |
1455 |
9.61 |
|
|
|
| 9 |
A |
1509 |
1447 |
29.11 |
|
|
|
| 10 |
A |
1450 |
1390 |
23.17 |
|
|
|
| 11 |
A |
1431 |
1372 |
2.38 |
|
|
|
| 12 |
A |
1246 |
1194 |
70.48 |
|
|
|
| 13 |
A |
1171 |
1122 |
22.33 |
|
|
|
| 14 |
A |
1161 |
1113 |
0.67 |
|
|
|
| 15 |
A |
1024 |
982 |
0.31 |
|
|
|
| 16 |
A |
988 |
947 |
22.19 |
|
|
|
| 17 |
A |
780 |
748 |
0.72 |
|
|
|
| 18 |
A |
552 |
529 |
55.41 |
|
|
|
| 19 |
A |
302 |
290 |
13.62 |
|
|
|
| 20 |
A |
283 |
271 |
80.10 |
|
|
|
| 21 |
A |
94 |
90 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16466.9 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 15783.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.860 |
0.433 |
0.004 |
| O2 |
1.368 |
-0.668 |
-0.000 |
| N3 |
-0.471 |
0.657 |
-0.020 |
| C4 |
-1.414 |
-0.440 |
0.005 |
| H5 |
1.454 |
1.364 |
0.017 |
| H6 |
-0.802 |
1.603 |
0.057 |
| H7 |
-2.411 |
-0.059 |
-0.217 |
| H8 |
-1.136 |
-1.178 |
-0.749 |
| H9 |
-1.425 |
-0.936 |
0.979 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2126 | 1.3498 | 2.4358 | 1.1047 | 2.0333 | 3.3150 | 2.6732 | 2.8367 |
O2 | 1.2126 | | 2.2669 | 2.7918 | 2.0337 | 3.1419 | 3.8343 | 2.6633 | 2.9725 | N3 | 1.3498 | 2.2669 | | 1.4469 | 2.0514 | 1.0052 | 2.0772 | 2.0832 | 2.1089 | C4 | 2.4358 | 2.7918 | 1.4469 | | 3.3886 | 2.1332 | 1.0900 | 1.0906 | 1.0936 | H5 | 1.1047 | 2.0337 | 2.0514 | 3.3886 | | 2.2697 | 4.1256 | 3.7093 | 3.8089 | H6 | 2.0333 | 3.1419 | 1.0052 | 2.1332 | 2.2697 | | 2.3289 | 2.9143 | 2.7723 | H7 | 3.3150 | 3.8343 | 2.0772 | 1.0900 | 4.1256 | 2.3289 | | 1.7778 | 1.7811 | H8 | 2.6732 | 2.6633 | 2.0832 | 1.0906 | 3.7093 | 2.9143 | 1.7778 | | 1.7685 | H9 | 2.8367 | 2.9725 | 2.1089 | 1.0936 | 3.8089 | 2.7723 | 1.7811 | 1.7685 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.104 |
|
C1 |
N3 |
H6 |
118.671 |
| O2 |
C1 |
N3 |
124.338 |
|
O2 |
C1 |
H5 |
122.644 |
| N3 |
C1 |
H5 |
113.014 |
|
N3 |
C4 |
H7 |
109.116 |
| N3 |
C4 |
H8 |
109.570 |
|
N3 |
C4 |
H9 |
111.460 |
| C4 |
N3 |
H6 |
119.817 |
|
H7 |
C4 |
H8 |
109.234 |
| H7 |
C4 |
H9 |
109.298 |
|
H8 |
C4 |
H9 |
108.132 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.207 |
|
|
|
| 2 |
O |
-0.371 |
|
|
|
| 3 |
N |
-0.542 |
|
|
|
| 4 |
C |
-0.530 |
|
|
|
| 5 |
H |
0.160 |
|
|
|
| 6 |
H |
0.349 |
|
|
|
| 7 |
H |
0.232 |
|
|
|
| 8 |
H |
0.262 |
|
|
|
| 9 |
H |
0.232 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.910 |
2.613 |
0.000 |
3.911 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.453 |
2.515 |
0.000 |
| y |
2.515 |
-21.954 |
0.000 |
| z |
0.000 |
0.000 |
-24.776 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.088 |
2.515 |
0.000 |
| y |
2.515 |
3.160 |
0.000 |
| z |
0.000 |
0.000 |
-1.073 |
|
| Polar |
|---|
| 3z2-r2 | -2.145 |
|---|
| x2-y2 | -3.499 |
|---|
| xy | 2.515 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.006 |
-0.090 |
0.000 |
| y |
-0.090 |
5.019 |
0.000 |
| z |
0.000 |
0.000 |
3.226 |
<r2> (average value of r
2) Å
2
| <r2> |
77.977 |
| (<r2>)1/2 |
8.830 |
