Jump to
S1C2
S1C3
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.027458 |
| Energy at 298.15K | -209.033534 |
| HF Energy | -209.027458 |
| Nuclear repulsion energy | 119.632299 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3644 |
3497 |
17.92 |
|
|
|
| 2 |
A' |
3143 |
3016 |
19.89 |
|
|
|
| 3 |
A' |
3046 |
2923 |
59.32 |
|
|
|
| 4 |
A' |
2947 |
2828 |
104.97 |
|
|
|
| 5 |
A' |
1846 |
1772 |
501.95 |
|
|
|
| 6 |
A' |
1554 |
1492 |
24.02 |
|
|
|
| 7 |
A' |
1508 |
1447 |
7.14 |
|
|
|
| 8 |
A' |
1479 |
1419 |
4.44 |
|
|
|
| 9 |
A' |
1407 |
1350 |
8.09 |
|
|
|
| 10 |
A' |
1322 |
1269 |
105.70 |
|
|
|
| 11 |
A' |
1185 |
1138 |
26.23 |
|
|
|
| 12 |
A' |
1025 |
984 |
47.97 |
|
|
|
| 13 |
A' |
622 |
596 |
15.47 |
|
|
|
| 14 |
A' |
347 |
333 |
8.63 |
|
|
|
| 15 |
A" |
3104 |
2979 |
36.16 |
|
|
|
| 16 |
A" |
1502 |
1441 |
8.04 |
|
|
|
| 17 |
A" |
1157 |
1110 |
0.32 |
|
|
|
| 18 |
A" |
1044 |
1002 |
0.44 |
|
|
|
| 19 |
A" |
643 |
617 |
136.83 |
|
|
|
| 20 |
A" |
205 |
196 |
1.14 |
|
|
|
| 21 |
A" |
99 |
95 |
0.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16415.1 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 15751.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
-0.756 |
0.000 |
| O2 |
1.394 |
-1.231 |
0.000 |
| N3 |
0.000 |
0.569 |
0.000 |
| C4 |
-1.325 |
1.132 |
0.000 |
| H5 |
-0.635 |
-1.377 |
0.000 |
| H6 |
0.799 |
1.185 |
0.000 |
| H7 |
-2.054 |
0.319 |
0.000 |
| H8 |
-1.506 |
1.743 |
0.889 |
| H9 |
-1.506 |
1.743 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2075 | 1.3546 | 2.4800 | 1.1095 | 2.0078 | 2.5725 | 3.1996 | 3.1996 |
O2 | 1.2075 | | 2.2766 | 3.6021 | 2.0344 | 2.4885 | 3.7797 | 4.2479 | 4.2479 | N3 | 1.3546 | 2.2766 | | 1.4397 | 2.0469 | 1.0086 | 2.0689 | 2.1063 | 2.1063 | C4 | 2.4800 | 3.6021 | 1.4397 | | 2.6016 | 2.1244 | 1.0917 | 1.0941 | 1.0941 | H5 | 1.1095 | 2.0344 | 2.0469 | 2.6016 | | 2.9359 | 2.2104 | 3.3591 | 3.3591 | H6 | 2.0078 | 2.4885 | 1.0086 | 2.1244 | 2.9359 | | 2.9809 | 2.5320 | 2.5320 | H7 | 2.5725 | 3.7797 | 2.0689 | 1.0917 | 2.2104 | 2.9809 | | 1.7666 | 1.7666 | H8 | 3.1996 | 4.2479 | 2.1063 | 1.0941 | 3.3591 | 2.5320 | 1.7666 | | 1.7779 | H9 | 3.1996 | 4.2479 | 2.1063 | 1.0941 | 3.3591 | 2.5320 | 1.7666 | 1.7779 | |
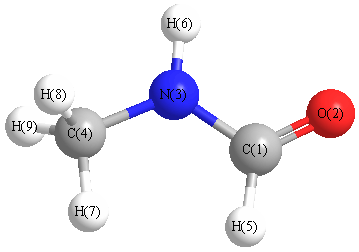 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
125.100 |
|
C1 |
N3 |
H6 |
115.559 |
| O2 |
C1 |
N3 |
125.287 |
|
O2 |
C1 |
H5 |
122.754 |
| N3 |
C1 |
H5 |
111.958 |
|
N3 |
C4 |
H7 |
108.855 |
| N3 |
C4 |
H8 |
111.736 |
|
N3 |
C4 |
H9 |
111.736 |
| C4 |
N3 |
H6 |
119.342 |
|
H7 |
C4 |
H8 |
107.844 |
| H7 |
C4 |
H9 |
107.844 |
|
H8 |
C4 |
H9 |
108.677 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.211 |
|
|
|
| 2 |
O |
-0.367 |
|
|
|
| 3 |
N |
-0.528 |
|
|
|
| 4 |
C |
-0.512 |
|
|
|
| 5 |
H |
0.147 |
|
|
|
| 6 |
H |
0.347 |
|
|
|
| 7 |
H |
0.229 |
|
|
|
| 8 |
H |
0.236 |
|
|
|
| 9 |
H |
0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.965 |
2.800 |
0.000 |
4.078 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.238 |
4.024 |
0.000 |
| y |
4.024 |
-24.244 |
0.000 |
| z |
0.000 |
0.000 |
-24.754 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.739 |
4.024 |
0.000 |
| y |
4.024 |
1.252 |
0.000 |
| z |
0.000 |
0.000 |
0.487 |
|
| Polar |
|---|
| 3z2-r2 | 0.974 |
|---|
| x2-y2 | -1.994 |
|---|
| xy | 4.024 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.616 |
-1.022 |
0.000 |
| y |
-1.022 |
5.659 |
0.000 |
| z |
0.000 |
0.000 |
3.239 |
<r2> (average value of r
2) Å
2
| <r2> |
88.578 |
| (<r2>)1/2 |
9.412 |
Jump to
S1C1
S1C3
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.027458 |
| Energy at 298.15K | -209.033534 |
| HF Energy | -209.027458 |
| Nuclear repulsion energy | 119.632299 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.029299 |
| Energy at 298.15K | -209.035424 |
| HF Energy | -209.029299 |
| Nuclear repulsion energy | 121.861103 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3678 |
3530 |
18.18 |
|
|
|
| 2 |
A |
3179 |
3051 |
1.72 |
|
|
|
| 3 |
A |
3112 |
2986 |
36.92 |
|
|
|
| 4 |
A |
3053 |
2929 |
47.85 |
|
|
|
| 5 |
A |
2964 |
2845 |
137.24 |
|
|
|
| 6 |
A |
1833 |
1759 |
345.78 |
|
|
|
| 7 |
A |
1588 |
1524 |
113.00 |
|
|
|
| 8 |
A |
1518 |
1457 |
9.56 |
|
|
|
| 9 |
A |
1509 |
1448 |
29.36 |
|
|
|
| 10 |
A |
1451 |
1392 |
23.33 |
|
|
|
| 11 |
A |
1431 |
1373 |
2.59 |
|
|
|
| 12 |
A |
1244 |
1193 |
71.33 |
|
|
|
| 13 |
A |
1171 |
1124 |
22.66 |
|
|
|
| 14 |
A |
1161 |
1114 |
0.67 |
|
|
|
| 15 |
A |
1024 |
983 |
0.30 |
|
|
|
| 16 |
A |
987 |
947 |
21.65 |
|
|
|
| 17 |
A |
780 |
749 |
0.70 |
|
|
|
| 18 |
A |
553 |
531 |
55.24 |
|
|
|
| 19 |
A |
304 |
291 |
13.68 |
|
|
|
| 20 |
A |
283 |
271 |
79.77 |
|
|
|
| 21 |
A |
94 |
90 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16456.5 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 15791.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.860 |
0.433 |
0.004 |
| O2 |
1.368 |
-0.669 |
-0.000 |
| N3 |
-0.471 |
0.657 |
-0.020 |
| C4 |
-1.414 |
-0.440 |
0.005 |
| H5 |
1.455 |
1.363 |
0.017 |
| H6 |
-0.802 |
1.603 |
0.057 |
| H7 |
-2.411 |
-0.059 |
-0.217 |
| H8 |
-1.136 |
-1.178 |
-0.748 |
| H9 |
-1.425 |
-0.936 |
0.979 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2127 | 1.3497 | 2.4359 | 1.1046 | 2.0333 | 3.3150 | 2.6732 | 2.8368 |
O2 | 1.2127 | | 2.2668 | 2.7914 | 2.0338 | 3.1418 | 3.8338 | 2.6628 | 2.9721 | N3 | 1.3497 | 2.2668 | | 1.4473 | 2.0513 | 1.0052 | 2.0775 | 2.0836 | 2.1092 | C4 | 2.4359 | 2.7914 | 1.4473 | | 3.3888 | 2.1336 | 1.0898 | 1.0904 | 1.0934 | H5 | 1.1046 | 2.0338 | 2.0513 | 3.3888 | | 2.2697 | 4.1258 | 3.7094 | 3.8090 | H6 | 2.0333 | 3.1418 | 1.0052 | 2.1336 | 2.2697 | | 2.3295 | 2.9147 | 2.7726 | H7 | 3.3150 | 3.8338 | 2.0775 | 1.0898 | 4.1258 | 2.3295 | | 1.7775 | 1.7806 | H8 | 2.6732 | 2.6628 | 2.0836 | 1.0904 | 3.7094 | 2.9147 | 1.7775 | | 1.7680 | H9 | 2.8368 | 2.9721 | 2.1092 | 1.0934 | 3.8090 | 2.7726 | 1.7806 | 1.7680 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.087 |
|
C1 |
N3 |
H6 |
118.674 |
| O2 |
C1 |
N3 |
124.321 |
|
O2 |
C1 |
H5 |
122.654 |
| N3 |
C1 |
H5 |
113.022 |
|
N3 |
C4 |
H7 |
109.124 |
| N3 |
C4 |
H8 |
109.580 |
|
N3 |
C4 |
H9 |
111.469 |
| C4 |
N3 |
H6 |
119.830 |
|
H7 |
C4 |
H8 |
109.230 |
| H7 |
C4 |
H9 |
109.289 |
|
H8 |
C4 |
H9 |
108.116 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.203 |
|
|
|
| 2 |
O |
-0.370 |
|
|
|
| 3 |
N |
-0.536 |
|
|
|
| 4 |
C |
-0.530 |
|
|
|
| 5 |
H |
0.160 |
|
|
|
| 6 |
H |
0.348 |
|
|
|
| 7 |
H |
0.232 |
|
|
|
| 8 |
H |
0.261 |
|
|
|
| 9 |
H |
0.232 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.912 |
2.615 |
-0.002 |
3.914 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.460 |
2.516 |
0.001 |
| y |
2.516 |
-21.973 |
-0.002 |
| z |
0.001 |
-0.002 |
-24.781 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.083 |
2.516 |
0.001 |
| y |
2.516 |
3.147 |
-0.002 |
| z |
0.001 |
-0.002 |
-1.064 |
|
| Polar |
|---|
| 3z2-r2 | -2.127 |
|---|
| x2-y2 | -3.487 |
|---|
| xy | 2.516 |
|---|
| xz | 0.001 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.015 |
-0.088 |
-0.000 |
| y |
-0.088 |
5.029 |
-0.000 |
| z |
-0.000 |
-0.000 |
3.229 |
<r2> (average value of r
2) Å
2
| <r2> |
77.964 |
| (<r2>)1/2 |
8.830 |
