Jump to
S1C2
S1C3
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.062329 |
| Energy at 298.15K | -209.068342 |
| HF Energy | -208.829819 |
| Nuclear repulsion energy | 119.185566 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3639 |
3639 |
19.37 |
|
|
|
| 2 |
A' |
3154 |
3154 |
20.95 |
|
|
|
| 3 |
A' |
3059 |
3059 |
57.98 |
|
|
|
| 4 |
A' |
2969 |
2969 |
100.97 |
|
|
|
| 5 |
A' |
1807 |
1807 |
451.04 |
|
|
|
| 6 |
A' |
1565 |
1565 |
17.22 |
|
|
|
| 7 |
A' |
1509 |
1509 |
7.19 |
|
|
|
| 8 |
A' |
1497 |
1497 |
5.07 |
|
|
|
| 9 |
A' |
1417 |
1417 |
8.67 |
|
|
|
| 10 |
A' |
1317 |
1317 |
109.62 |
|
|
|
| 11 |
A' |
1179 |
1179 |
35.42 |
|
|
|
| 12 |
A' |
1023 |
1023 |
46.13 |
|
|
|
| 13 |
A' |
614 |
614 |
14.18 |
|
|
|
| 14 |
A' |
347 |
347 |
8.16 |
|
|
|
| 15 |
A" |
3116 |
3116 |
36.54 |
|
|
|
| 16 |
A" |
1521 |
1521 |
7.04 |
|
|
|
| 17 |
A" |
1168 |
1168 |
0.58 |
|
|
|
| 18 |
A" |
1042 |
1042 |
0.79 |
|
|
|
| 19 |
A" |
626 |
626 |
135.86 |
|
|
|
| 20 |
A" |
195 |
195 |
1.47 |
|
|
|
| 21 |
A" |
79 |
79 |
1.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16420.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16420.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
-0.760 |
0.000 |
| O2 |
1.400 |
-1.237 |
0.000 |
| N3 |
0.000 |
0.571 |
0.000 |
| C4 |
-1.333 |
1.138 |
0.000 |
| H5 |
-0.632 |
-1.380 |
0.000 |
| H6 |
0.800 |
1.186 |
0.000 |
| H7 |
-2.060 |
0.326 |
0.000 |
| H8 |
-1.509 |
1.749 |
0.888 |
| H9 |
-1.509 |
1.749 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2138 | 1.3608 | 2.4932 | 1.1060 | 2.0126 | 2.5831 | 3.2086 | 3.2086 |
O2 | 1.2138 | | 2.2866 | 3.6207 | 2.0369 | 2.4957 | 3.7966 | 4.2619 | 4.2619 | N3 | 1.3608 | 2.2866 | | 1.4484 | 2.0507 | 1.0086 | 2.0742 | 2.1099 | 2.1099 | C4 | 2.4932 | 3.6207 | 1.4484 | | 2.6138 | 2.1331 | 1.0897 | 1.0920 | 1.0920 | H5 | 1.1060 | 2.0369 | 2.0507 | 2.6138 | | 2.9378 | 2.2249 | 3.3685 | 3.3685 | H6 | 2.0126 | 2.4957 | 1.0086 | 2.1331 | 2.9378 | | 2.9858 | 2.5366 | 2.5366 | H7 | 2.5831 | 3.7966 | 2.0742 | 1.0897 | 2.2249 | 2.9858 | | 1.7653 | 1.7653 | H8 | 3.2086 | 4.2619 | 2.1099 | 1.0920 | 3.3685 | 2.5366 | 1.7653 | | 1.7760 | H9 | 3.2086 | 4.2619 | 2.1099 | 1.0920 | 3.3685 | 2.5366 | 1.7653 | 1.7760 | |
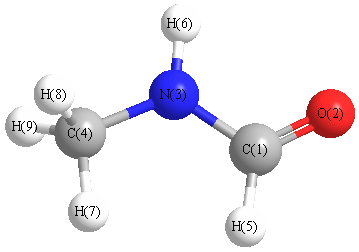 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
125.095 |
|
C1 |
N3 |
H6 |
115.494 |
| O2 |
C1 |
N3 |
125.190 |
|
O2 |
C1 |
H5 |
122.750 |
| N3 |
C1 |
H5 |
112.060 |
|
N3 |
C4 |
H7 |
108.794 |
| N3 |
C4 |
H8 |
111.535 |
|
N3 |
C4 |
H9 |
111.535 |
| C4 |
N3 |
H6 |
119.411 |
|
H7 |
C4 |
H8 |
108.022 |
| H7 |
C4 |
H9 |
108.022 |
|
H8 |
C4 |
H9 |
108.809 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.062329 |
| Energy at 298.15K | -209.068342 |
| HF Energy | -208.829819 |
| Nuclear repulsion energy | 119.185566 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -209.064231 |
| Energy at 298.15K | -209.070273 |
| HF Energy | -208.831544 |
| Nuclear repulsion energy | 121.391222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3670 |
3670 |
18.67 |
|
|
|
| 2 |
A |
3190 |
3190 |
2.03 |
|
|
|
| 3 |
A |
3123 |
3123 |
37.84 |
|
|
|
| 4 |
A |
3065 |
3065 |
47.95 |
|
|
|
| 5 |
A |
2980 |
2980 |
135.37 |
|
|
|
| 6 |
A |
1793 |
1793 |
307.15 |
|
|
|
| 7 |
A |
1588 |
1588 |
96.33 |
|
|
|
| 8 |
A |
1538 |
1538 |
8.29 |
|
|
|
| 9 |
A |
1526 |
1526 |
40.64 |
|
|
|
| 10 |
A |
1470 |
1470 |
22.30 |
|
|
|
| 11 |
A |
1434 |
1434 |
5.67 |
|
|
|
| 12 |
A |
1233 |
1233 |
85.13 |
|
|
|
| 13 |
A |
1178 |
1178 |
19.70 |
|
|
|
| 14 |
A |
1172 |
1172 |
1.05 |
|
|
|
| 15 |
A |
1022 |
1022 |
0.13 |
|
|
|
| 16 |
A |
971 |
971 |
18.65 |
|
|
|
| 17 |
A |
776 |
776 |
0.59 |
|
|
|
| 18 |
A |
538 |
538 |
49.57 |
|
|
|
| 19 |
A |
303 |
303 |
13.00 |
|
|
|
| 20 |
A |
259 |
259 |
85.88 |
|
|
|
| 21 |
A |
73 |
73 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16449.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16449.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.863 |
0.435 |
0.004 |
| O2 |
1.377 |
-0.671 |
-0.000 |
| N3 |
-0.474 |
0.660 |
-0.021 |
| C4 |
-1.422 |
-0.443 |
0.005 |
| H5 |
1.456 |
1.365 |
0.018 |
| H6 |
-0.804 |
1.606 |
0.059 |
| H7 |
-2.417 |
-0.060 |
-0.211 |
| H8 |
-1.149 |
-1.178 |
-0.750 |
| H9 |
-1.431 |
-0.941 |
0.977 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2190 | 1.3561 | 2.4484 | 1.1028 | 2.0383 | 3.3250 | 2.6875 | 2.8464 |
O2 | 1.2190 | | 2.2790 | 2.8081 | 2.0376 | 3.1531 | 3.8489 | 2.6833 | 2.9849 | N3 | 1.3561 | 2.2790 | | 1.4545 | 2.0546 | 1.0054 | 2.0813 | 2.0895 | 2.1148 | C4 | 2.4484 | 2.8081 | 1.4545 | | 3.3986 | 2.1404 | 1.0884 | 1.0886 | 1.0918 | H5 | 1.1028 | 2.0376 | 2.0546 | 3.3986 | | 2.2729 | 4.1332 | 3.7210 | 3.8167 | H6 | 2.0383 | 3.1531 | 1.0054 | 2.1404 | 2.2729 | | 2.3344 | 2.9198 | 2.7782 | H7 | 3.3250 | 3.8489 | 2.0813 | 1.0884 | 4.1332 | 2.3344 | | 1.7749 | 1.7780 | H8 | 2.6875 | 2.6833 | 2.0895 | 1.0886 | 3.7210 | 2.9198 | 1.7749 | | 1.7651 | H9 | 2.8464 | 2.9849 | 2.1148 | 1.0918 | 3.8167 | 2.7782 | 1.7780 | 1.7651 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.139 |
|
C1 |
N3 |
H6 |
118.589 |
| O2 |
C1 |
N3 |
124.423 |
|
O2 |
C1 |
H5 |
122.626 |
| N3 |
C1 |
H5 |
112.948 |
|
N3 |
C4 |
H7 |
109.024 |
| N3 |
C4 |
H8 |
109.658 |
|
N3 |
C4 |
H9 |
111.516 |
| C4 |
N3 |
H6 |
119.830 |
|
H7 |
C4 |
H8 |
109.232 |
| H7 |
C4 |
H9 |
109.280 |
|
H8 |
C4 |
H9 |
108.099 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
