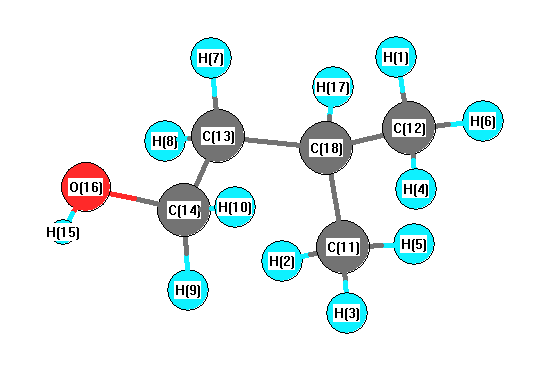
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3824 |
3681 |
10.06 |
|
|
|
| 2 |
A' |
3104 |
2988 |
64.35 |
|
|
|
| 3 |
A' |
3103 |
2987 |
87.96 |
|
|
|
| 4 |
A' |
3040 |
2926 |
22.64 |
|
|
|
| 5 |
A' |
3034 |
2921 |
43.63 |
|
|
|
| 6 |
A' |
3016 |
2904 |
11.81 |
|
|
|
| 7 |
A' |
2985 |
2874 |
51.49 |
|
|
|
| 8 |
A' |
1539 |
1481 |
0.86 |
|
|
|
| 9 |
A' |
1525 |
1468 |
9.32 |
|
|
|
| 10 |
A' |
1523 |
1466 |
15.19 |
|
|
|
| 11 |
A' |
1499 |
1443 |
2.95 |
|
|
|
| 12 |
A' |
1476 |
1421 |
2.42 |
|
|
|
| 13 |
A' |
1428 |
1375 |
5.47 |
|
|
|
| 14 |
A' |
1382 |
1331 |
0.97 |
|
|
|
| 15 |
A' |
1351 |
1300 |
35.80 |
|
|
|
| 16 |
A' |
1259 |
1212 |
32.57 |
|
|
|
| 17 |
A' |
1211 |
1166 |
4.91 |
|
|
|
| 18 |
A' |
1133 |
1090 |
1.07 |
|
|
|
| 19 |
A' |
1080 |
1039 |
100.18 |
|
|
|
| 20 |
A' |
1008 |
971 |
4.32 |
|
|
|
| 21 |
A' |
965 |
929 |
11.44 |
|
|
|
| 22 |
A' |
807 |
777 |
5.73 |
|
|
|
| 23 |
A' |
556 |
535 |
1.39 |
|
|
|
| 24 |
A' |
403 |
388 |
7.19 |
|
|
|
| 25 |
A' |
326 |
314 |
5.78 |
|
|
|
| 26 |
A' |
257 |
248 |
0.02 |
|
|
|
| 27 |
A' |
207 |
200 |
2.23 |
|
|
|
| 28 |
A" |
3102 |
2986 |
28.73 |
|
|
|
| 29 |
A" |
3097 |
2981 |
0.31 |
|
|
|
| 30 |
A" |
3078 |
2963 |
50.65 |
|
|
|
| 31 |
A" |
3030 |
2917 |
56.82 |
|
|
|
| 32 |
A" |
3014 |
2901 |
28.24 |
|
|
|
| 33 |
A" |
1511 |
1455 |
1.83 |
|
|
|
| 34 |
A" |
1499 |
1443 |
0.67 |
|
|
|
| 35 |
A" |
1409 |
1357 |
12.00 |
|
|
|
| 36 |
A" |
1375 |
1324 |
0.55 |
|
|
|
| 37 |
A" |
1332 |
1282 |
0.12 |
|
|
|
| 38 |
A" |
1261 |
1214 |
0.00 |
|
|
|
| 39 |
A" |
1183 |
1139 |
5.64 |
|
|
|
| 40 |
A" |
1013 |
975 |
0.51 |
|
|
|
| 41 |
A" |
969 |
933 |
0.00 |
|
|
|
| 42 |
A" |
930 |
895 |
0.93 |
|
|
|
| 43 |
A" |
798 |
768 |
0.03 |
|
|
|
| 44 |
A" |
371 |
357 |
0.61 |
|
|
|
| 45 |
A" |
314 |
303 |
133.11 |
|
|
|
| 46 |
A" |
242 |
233 |
0.68 |
|
|
|
| 47 |
A" |
104 |
100 |
0.03 |
|
|
|
| 48 |
A" |
72 |
69 |
9.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36370.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 35014.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.223 |
|
|
|
| 2 |
H |
0.223 |
|
|
|
| 3 |
H |
0.213 |
|
|
|
| 4 |
H |
0.213 |
|
|
|
| 5 |
H |
0.222 |
|
|
|
| 6 |
H |
0.222 |
|
|
|
| 7 |
H |
0.227 |
|
|
|
| 8 |
H |
0.227 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.195 |
|
|
|
| 11 |
C |
-0.645 |
|
|
|
| 12 |
C |
-0.645 |
|
|
|
| 13 |
C |
-0.455 |
|
|
|
| 14 |
C |
-0.184 |
|
|
|
| 15 |
H |
0.389 |
|
|
|
| 16 |
O |
-0.578 |
|
|
|
| 17 |
H |
0.218 |
|
|
|
| 18 |
C |
-0.259 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.340 |
-1.497 |
0.000 |
1.535 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.780 |
3.588 |
0.000 |
| y |
3.588 |
-43.585 |
0.000 |
| z |
0.000 |
0.000 |
-40.425 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.226 |
3.588 |
0.000 |
| y |
3.588 |
-5.983 |
0.000 |
| z |
0.000 |
0.000 |
-1.243 |
|
| Polar |
|---|
| 3z2-r2 | -2.486 |
|---|
| x2-y2 | 8.806 |
|---|
| xy | 3.588 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
223.279 |
| (<r2>)1/2 |
14.943 |