return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (1-Butanol, 3-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-272.237374
Energy at 298.15K-272.250888
HF Energy-271.230454
Nuclear repulsion energy257.135285
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3853 3649 17.87      
2 A' 3163 2996 62.17      
3 A' 3159 2992 49.93      
4 A' 3083 2920 32.14      
5 A' 3073 2910 26.93      
6 A' 3068 2906 10.98      
7 A' 3048 2887 34.95      
8 A' 1570 1487 0.71      
9 A' 1553 1471 13.93      
10 A' 1549 1467 12.71      
11 A' 1530 1450 3.18      
12 A' 1498 1419 2.45      
13 A' 1458 1381 7.86      
14 A' 1400 1326 0.39      
15 A' 1368 1296 31.91      
16 A' 1278 1210 34.11      
17 A' 1237 1172 4.11      
18 A' 1149 1089 2.16      
19 A' 1100 1042 87.03      
20 A' 1030 976 7.23      
21 A' 989 937 9.95      
22 A' 827 783 5.72      
23 A' 569 539 1.32      
24 A' 416 394 7.18      
25 A' 335 317 6.26      
26 A' 268 254 0.06      
27 A' 219 207 2.62      
28 A" 3160 2992 13.05      
29 A" 3157 2990 5.94      
30 A" 3137 2971 43.64      
31 A" 3094 2930 43.37      
32 A" 3069 2907 19.35      
33 A" 1539 1457 2.38      
34 A" 1523 1443 0.66      
35 A" 1439 1363 14.59      
36 A" 1400 1326 0.21      
37 A" 1352 1280 0.06      
38 A" 1278 1211 0.02      
39 A" 1207 1143 5.03      
40 A" 1036 982 0.79      
41 A" 988 936 0.00      
42 A" 946 896 0.90      
43 A" 808 765 0.03      
44 A" 381 361 0.02      
45 A" 284 269 130.24      
46 A" 251 238 5.42      
47 A" 108 103 0.42      
48 A" 57 54 12.60      

Unscaled Zero Point Vibrational Energy (zpe) 37001.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 35044.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.20153 0.06278 0.05629

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 1.206 -0.338 2.163
H2 1.206 -0.338 -2.163
H3 0.495 -1.694 -1.278
H4 0.495 -1.694 1.278
H5 2.244 -1.483 -1.304
H6 2.244 -1.483 1.304
H7 0.036 1.503 0.879
H8 0.036 1.503 -0.879
H9 -1.455 -0.479 -0.889
H10 -1.455 -0.479 0.889
C11 1.292 -0.946 -1.258
C12 1.292 -0.946 1.258
C13 0.000 0.849 0.000
C14 -1.348 0.156 0.000
H15 -3.200 0.763 0.000
O16 -2.336 1.182 0.000
H17 2.102 0.572 0.000
C18 1.219 -0.081 0.000

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 H7 H8 H9 H10 C11 C12 C13 C14 H15 O16 H17 C18
H14.32703.76681.76893.79651.76822.53133.74314.05192.95423.47661.09382.74653.38385.03044.41992.51212.1787
H24.32701.76893.76681.76823.79653.74312.53132.95424.05191.09383.47662.74653.38385.03044.41992.51212.1787
H33.76681.76892.55591.76163.12583.88353.25432.33063.15821.09342.76192.88872.90784.61794.23323.05782.1817
H41.76893.76682.55593.12581.76163.25433.88353.15822.33062.76191.09342.88872.90784.61794.23323.05782.1817
H53.79651.76821.76163.12582.60854.30733.73763.85534.41581.09372.78593.48904.15836.03165.45672.43822.1720
H61.76823.79653.12581.76162.60853.73764.30734.41583.85532.78591.09373.48904.15836.03165.45672.43822.1720
H72.53133.74313.88353.25434.30733.73761.75723.04552.48033.48432.77811.09592.12193.43372.54972.42982.1630
H83.74312.53133.25433.88353.73764.30731.75722.48033.04552.77813.48431.09592.12193.43372.54972.42982.1630
H94.05192.95422.33063.15823.85534.41583.04552.48031.77742.81123.51802.16111.09762.31892.07963.81382.8459
H102.95424.05193.15822.33064.41583.85532.48033.04551.77743.51802.81122.16111.09762.31892.07963.81382.8459
C113.47661.09381.09342.76191.09372.78593.48432.77812.81123.51802.51682.54453.12584.96834.39022.13151.5289
C121.09383.47662.76191.09342.78591.09372.77813.48433.51802.81122.51682.54453.12584.96834.39022.13151.5289
C132.74652.74652.88872.88873.48903.48901.09591.09592.16112.16112.54452.54451.51593.20092.35932.11981.5330
C143.38383.38382.90782.90784.15834.15832.12192.12191.09761.09763.12583.12581.51591.94851.42393.47502.5782
H155.03045.03044.61794.61796.03166.03163.43373.43372.31892.31894.96834.96833.20091.94850.96015.30484.4986
O164.41994.41994.23324.23325.45675.45672.54972.54972.07962.07964.39024.39022.35931.42390.96014.47903.7723
H172.51212.51213.05783.05782.43822.43822.42982.42983.81383.81382.13152.13152.11983.47505.30484.47901.0979
C182.17872.17872.18172.18172.17202.17202.16302.16302.84592.84591.52891.52891.53302.57824.49863.77231.0979

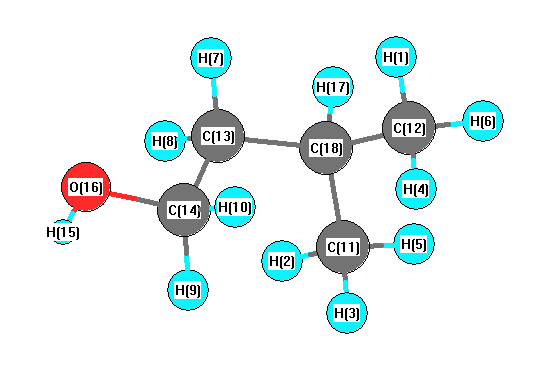
picture of 1-Butanol, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C12 H4 107.951 H1 C12 H6 107.865
H1 C12 C18 111.266 H2 C11 H3 107.951
H2 C11 H5 107.865 H2 C11 C18 111.266
H3 C11 H5 107.314 H3 C11 C18 111.532
H4 C12 H6 107.314 H4 C12 C18 111.532
H5 C11 C18 110.737 H6 C12 C18 110.737
H7 C13 H8 106.589 H7 C13 C14 107.586
H7 C13 C18 109.607 H8 C13 C14 107.586
H8 C13 C18 109.607 H9 C14 H10 108.137
H9 C14 C13 110.546 H9 C14 O16 110.460
H10 C14 C13 110.546 H10 C14 O16 110.460
C11 C18 C12 110.790 C11 C18 C13 112.411
C11 C18 H17 107.337 C12 C18 C13 112.411
C12 C18 H17 107.337 C13 C14 O16 106.708
C13 C18 H17 106.182 C14 C13 C18 115.482
C14 O16 H15 108.048
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability