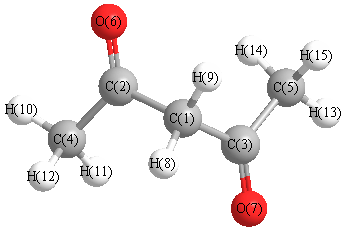
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3163 |
3045 |
9.28 |
|
|
|
| 2 |
A |
3108 |
2992 |
7.79 |
|
|
|
| 3 |
A |
3082 |
2967 |
3.61 |
|
|
|
| 4 |
A |
3043 |
2930 |
0.32 |
|
|
|
| 5 |
A |
1828 |
1759 |
23.07 |
|
|
|
| 6 |
A |
1485 |
1430 |
4.59 |
|
|
|
| 7 |
A |
1482 |
1426 |
23.66 |
|
|
|
| 8 |
A |
1468 |
1413 |
10.56 |
|
|
|
| 9 |
A |
1395 |
1343 |
7.29 |
|
|
|
| 10 |
A |
1281 |
1233 |
24.35 |
|
|
|
| 11 |
A |
1150 |
1107 |
2.37 |
|
|
|
| 12 |
A |
1081 |
1041 |
0.95 |
|
|
|
| 13 |
A |
941 |
906 |
1.14 |
|
|
|
| 14 |
A |
794 |
764 |
0.16 |
|
|
|
| 15 |
A |
626 |
603 |
2.02 |
|
|
|
| 16 |
A |
490 |
472 |
5.78 |
|
|
|
| 17 |
A |
320 |
308 |
0.80 |
|
|
|
| 18 |
A |
168 |
162 |
0.06 |
|
|
|
| 19 |
A |
142 |
137 |
0.98 |
|
|
|
| 20 |
A |
62 |
60 |
7.55 |
|
|
|
| 21 |
B |
3163 |
3045 |
11.05 |
|
|
|
| 22 |
B |
3152 |
3034 |
7.95 |
|
|
|
| 23 |
B |
3108 |
2992 |
1.03 |
|
|
|
| 24 |
B |
3043 |
2929 |
4.20 |
|
|
|
| 25 |
B |
1799 |
1732 |
349.17 |
|
|
|
| 26 |
B |
1481 |
1426 |
1.07 |
|
|
|
| 27 |
B |
1477 |
1422 |
33.03 |
|
|
|
| 28 |
B |
1394 |
1342 |
109.66 |
|
|
|
| 29 |
B |
1257 |
1210 |
135.56 |
|
|
|
| 30 |
B |
1192 |
1147 |
172.50 |
|
|
|
| 31 |
B |
1062 |
1022 |
8.64 |
|
|
|
| 32 |
B |
999 |
962 |
0.97 |
|
|
|
| 33 |
B |
896 |
863 |
24.64 |
|
|
|
| 34 |
B |
810 |
780 |
10.53 |
|
|
|
| 35 |
B |
549 |
529 |
27.10 |
|
|
|
| 36 |
B |
502 |
483 |
0.99 |
|
|
|
| 37 |
B |
409 |
393 |
1.31 |
|
|
|
| 38 |
B |
169 |
163 |
0.43 |
|
|
|
| 39 |
B |
46 |
44 |
10.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26807.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 25807.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.617 |
|
|
|
| 2 |
C |
0.319 |
|
|
|
| 3 |
C |
0.319 |
|
|
|
| 4 |
C |
-0.733 |
|
|
|
| 5 |
C |
-0.733 |
|
|
|
| 6 |
O |
-0.297 |
|
|
|
| 7 |
O |
-0.297 |
|
|
|
| 8 |
H |
0.259 |
|
|
|
| 9 |
H |
0.259 |
|
|
|
| 10 |
H |
0.253 |
|
|
|
| 11 |
H |
0.261 |
|
|
|
| 12 |
H |
0.247 |
|
|
|
| 13 |
H |
0.253 |
|
|
|
| 14 |
H |
0.261 |
|
|
|
| 15 |
H |
0.247 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.577 |
1.577 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.599 |
-9.146 |
0.000 |
| y |
-9.146 |
-45.597 |
0.000 |
| z |
0.000 |
0.000 |
-40.153 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.725 |
-9.146 |
0.000 |
| y |
-9.146 |
-3.721 |
0.000 |
| z |
0.000 |
0.000 |
4.446 |
|
| Polar |
|---|
| 3z2-r2 | 8.892 |
|---|
| x2-y2 | 1.998 |
|---|
| xy | -9.146 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.529 |
-0.410 |
0.000 |
| y |
-0.410 |
9.664 |
0.000 |
| z |
0.000 |
0.000 |
7.236 |
<r2> (average value of r
2) Å
2
| <r2> |
230.149 |
| (<r2>)1/2 |
15.171 |