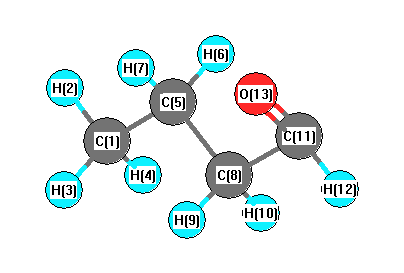
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3110 |
2994 |
40.87 |
|
|
|
| 2 |
A' |
3056 |
2942 |
20.48 |
|
|
|
| 3 |
A' |
3033 |
2920 |
30.37 |
|
|
|
| 4 |
A' |
3008 |
2896 |
26.16 |
|
|
|
| 5 |
A' |
2871 |
2763 |
171.78 |
|
|
|
| 6 |
A' |
1835 |
1767 |
163.38 |
|
|
|
| 7 |
A' |
1519 |
1462 |
9.37 |
|
|
|
| 8 |
A' |
1501 |
1445 |
3.46 |
|
|
|
| 9 |
A' |
1459 |
1405 |
16.24 |
|
|
|
| 10 |
A' |
1427 |
1374 |
5.41 |
|
|
|
| 11 |
A' |
1422 |
1369 |
8.63 |
|
|
|
| 12 |
A' |
1407 |
1354 |
19.03 |
|
|
|
| 13 |
A' |
1320 |
1270 |
9.19 |
|
|
|
| 14 |
A' |
1142 |
1100 |
11.97 |
|
|
|
| 15 |
A' |
1064 |
1025 |
0.18 |
|
|
|
| 16 |
A' |
969 |
933 |
6.42 |
|
|
|
| 17 |
A' |
864 |
832 |
8.72 |
|
|
|
| 18 |
A' |
701 |
674 |
14.64 |
|
|
|
| 19 |
A' |
347 |
334 |
0.72 |
|
|
|
| 20 |
A' |
194 |
186 |
8.57 |
|
|
|
| 21 |
A" |
3107 |
2991 |
66.79 |
|
|
|
| 22 |
A" |
3082 |
2967 |
0.01 |
|
|
|
| 23 |
A" |
3031 |
2918 |
10.97 |
|
|
|
| 24 |
A" |
1516 |
1459 |
9.87 |
|
|
|
| 25 |
A" |
1337 |
1287 |
0.16 |
|
|
|
| 26 |
A" |
1262 |
1215 |
0.10 |
|
|
|
| 27 |
A" |
1156 |
1113 |
0.02 |
|
|
|
| 28 |
A" |
955 |
919 |
0.00 |
|
|
|
| 29 |
A" |
790 |
760 |
1.65 |
|
|
|
| 30 |
A" |
665 |
641 |
4.88 |
|
|
|
| 31 |
A" |
252 |
242 |
0.04 |
|
|
|
| 32 |
A" |
184 |
177 |
1.65 |
|
|
|
| 33 |
A" |
78 |
75 |
1.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24830.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 23903.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.651 |
|
|
|
| 2 |
H |
0.229 |
|
|
|
| 3 |
H |
0.217 |
|
|
|
| 4 |
H |
0.217 |
|
|
|
| 5 |
C |
-0.438 |
|
|
|
| 6 |
H |
0.232 |
|
|
|
| 7 |
H |
0.232 |
|
|
|
| 8 |
C |
-0.524 |
|
|
|
| 9 |
H |
0.238 |
|
|
|
| 10 |
H |
0.238 |
|
|
|
| 11 |
C |
0.133 |
|
|
|
| 12 |
H |
0.163 |
|
|
|
| 13 |
O |
-0.287 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.752 |
1.847 |
0.000 |
2.546 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.832 |
-3.591 |
0.000 |
| y |
-3.591 |
-30.932 |
0.000 |
| z |
0.000 |
0.000 |
-30.711 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.010 |
-3.591 |
0.000 |
| y |
-3.591 |
2.840 |
0.000 |
| z |
0.000 |
0.000 |
3.171 |
|
| Polar |
|---|
| 3z2-r2 | 6.341 |
|---|
| x2-y2 | -5.900 |
|---|
| xy | -3.591 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
155.092 |
| (<r2>)1/2 |
12.454 |