return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH2CH2CH3 (Butanal)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-231.867729
Energy at 298.15K-231.876399
HF Energy-231.033595
Nuclear repulsion energy172.137537
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3168 3001 29.50      
2 A' 3104 2940 16.44      
3 A' 3076 2914 24.24      
4 A' 3057 2895 19.31      
5 A' 2945 2790 150.40      
6 A' 1784 1690 95.82      
7 A' 1546 1464 10.06      
8 A' 1530 1449 2.86      
9 A' 1494 1415 17.14      
10 A' 1453 1376 6.56      
11 A' 1442 1365 4.24      
12 A' 1427 1352 14.89      
13 A' 1333 1263 12.11      
14 A' 1162 1100 11.11      
15 A' 1090 1032 0.15      
16 A' 986 933 4.74      
17 A' 883 836 7.47      
18 A' 708 670 13.18      
19 A' 354 336 0.79      
20 A' 202 191 7.85      
21 A" 3168 3001 50.26      
22 A" 3146 2979 0.11      
23 A" 3098 2934 8.44      
24 A" 1543 1462 10.26      
25 A" 1356 1285 0.19      
26 A" 1283 1215 0.12      
27 A" 1176 1114 0.04      
28 A" 966 915 0.04      
29 A" 805 762 1.01      
30 A" 679 643 4.79      
31 A" 253 240 0.05      
32 A" 177 167 1.69      
33 A" 88 83 1.66      

Unscaled Zero Point Vibrational Energy (zpe) 25240.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23905.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.50146 0.08573 0.07629

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.273 -0.350 0.000
H2 2.805 -1.304 0.000
H3 2.594 0.210 0.883
H4 2.594 0.210 -0.883
C5 0.763 -0.559 0.000
H6 0.461 -1.145 -0.873
H7 0.461 -1.145 0.873
C8 0.000 0.758 0.000
H9 0.267 1.371 0.872
H10 0.267 1.371 -0.872
C11 -1.499 0.596 0.000
H12 -2.082 1.542 0.000
O13 -2.074 -0.473 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 O13
C11.09241.09341.09341.52512.16322.16322.52852.78312.78313.88854.74844.3487
H21.09241.76491.76492.17412.50672.50673.48103.78903.78904.70445.65544.9492
H31.09341.76491.76612.17383.07722.52722.79452.60083.13774.20464.94194.7994
H41.09341.76491.76612.17382.52723.07722.79453.13772.60084.20464.94194.7994
C51.52512.17412.17382.17381.09361.09361.52152.17512.17512.53913.53652.8376
H62.16322.50673.07722.52721.09361.74542.14323.06782.52322.76263.80102.7635
H72.16322.50672.52723.07721.09361.74542.14322.52323.06782.76263.80102.7635
C82.52853.48102.79452.79451.52152.14322.14321.09911.09911.50722.22502.4112
H92.78313.78902.60083.13772.17513.06782.52321.09911.74452.11592.51163.1044
H102.78313.78903.13772.60082.17512.52323.06781.09911.74452.11592.51163.1044
C113.88854.70444.20464.20462.53912.76262.76261.50722.11592.11591.11141.2138
H124.74845.65544.94194.94193.53653.80103.80102.22502.51162.51161.11142.0147
O134.34874.94924.79944.79942.83762.76352.76352.41123.10443.10441.21382.0147

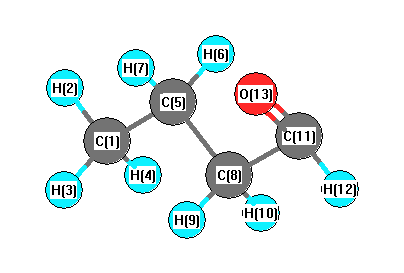
picture of Butanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.303 C1 C5 H7 110.303
C1 C5 C8 112.180 H2 C1 H3 107.688
H2 C1 H4 107.688 H2 C1 C5 111.245
H3 C1 H4 107.724 H3 C1 C5 111.161
H4 C1 C5 111.161 C5 C8 H9 111.169
C5 C8 H10 111.169 C5 C8 C11 113.931
H6 C5 H7 105.883 H6 C5 C8 108.981
H7 C5 C8 108.981 C8 C11 H12 115.534
C8 C11 O13 124.434 H9 C8 H10 105.045
H9 C8 C11 107.525 H10 C8 C11 107.525
H12 C11 O13 120.032
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability