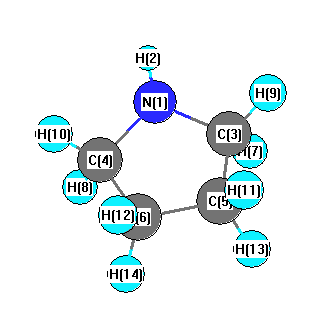
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3790 |
3427 |
0.59 |
|
|
|
| 2 |
A' |
3260 |
2948 |
130.11 |
|
|
|
| 3 |
A' |
3234 |
2925 |
5.38 |
|
|
|
| 4 |
A' |
3216 |
2908 |
73.39 |
|
|
|
| 5 |
A' |
3104 |
2807 |
180.85 |
|
|
|
| 6 |
A' |
1674 |
1514 |
0.44 |
|
|
|
| 7 |
A' |
1644 |
1487 |
3.31 |
|
|
|
| 8 |
A' |
1540 |
1393 |
2.97 |
|
|
|
| 9 |
A' |
1450 |
1311 |
1.87 |
|
|
|
| 10 |
A' |
1375 |
1243 |
3.11 |
|
|
|
| 11 |
A' |
1346 |
1217 |
11.29 |
|
|
|
| 12 |
A' |
1148 |
1038 |
0.53 |
|
|
|
| 13 |
A' |
1080 |
977 |
6.27 |
|
|
|
| 14 |
A' |
1017 |
920 |
16.68 |
|
|
|
| 15 |
A' |
967 |
875 |
40.24 |
|
|
|
| 16 |
A' |
952 |
861 |
25.06 |
|
|
|
| 17 |
A' |
833 |
754 |
22.36 |
|
|
|
| 18 |
A' |
624 |
565 |
45.43 |
|
|
|
| 19 |
A' |
307 |
277 |
4.85 |
|
|
|
| 20 |
A" |
3237 |
2928 |
45.43 |
|
|
|
| 21 |
A" |
3223 |
2915 |
88.81 |
|
|
|
| 22 |
A" |
3201 |
2895 |
23.47 |
|
|
|
| 23 |
A" |
3096 |
2800 |
42.35 |
|
|
|
| 24 |
A" |
1656 |
1498 |
2.72 |
|
|
|
| 25 |
A" |
1621 |
1466 |
0.34 |
|
|
|
| 26 |
A" |
1594 |
1442 |
9.96 |
|
|
|
| 27 |
A" |
1462 |
1322 |
9.61 |
|
|
|
| 28 |
A" |
1443 |
1305 |
21.72 |
|
|
|
| 29 |
A" |
1370 |
1239 |
5.10 |
|
|
|
| 30 |
A" |
1324 |
1197 |
0.19 |
|
|
|
| 31 |
A" |
1241 |
1122 |
10.42 |
|
|
|
| 32 |
A" |
1202 |
1087 |
4.24 |
|
|
|
| 33 |
A" |
996 |
901 |
0.06 |
|
|
|
| 34 |
A" |
941 |
851 |
2.39 |
|
|
|
| 35 |
A" |
686 |
620 |
1.19 |
|
|
|
| 36 |
A" |
56 |
51 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30453.3 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 27542.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.625 |
-0.031 |
|
0.002 |
| 2 |
H |
0.318 |
-0.017 |
|
0.009 |
| 3 |
C |
-0.238 |
0.262 |
|
0.178 |
| 4 |
C |
-0.238 |
-0.031 |
|
0.002 |
| 5 |
C |
-0.424 |
-0.017 |
|
0.009 |
| 6 |
C |
-0.424 |
0.262 |
|
0.178 |
| 7 |
H |
0.181 |
0.001 |
|
0.037 |
| 8 |
H |
0.181 |
-0.028 |
|
0.010 |
| 9 |
H |
0.206 |
0.033 |
|
-0.047 |
| 10 |
H |
0.206 |
0.001 |
|
0.037 |
| 11 |
H |
0.218 |
-0.028 |
|
0.010 |
| 12 |
H |
0.218 |
0.033 |
|
-0.047 |
| 13 |
H |
0.210 |
-0.793 |
|
-0.712 |
| 14 |
H |
0.210 |
0.354 |
|
0.334 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.042 |
-0.238 |
0.000 |
1.069 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.042 |
-0.231 |
0.000 |
1.067 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.441 |
0.983 |
0.000 |
| y |
0.983 |
-30.766 |
0.000 |
| z |
0.000 |
0.000 |
-31.793 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.161 |
0.983 |
0.000 |
| y |
0.983 |
2.351 |
0.000 |
| z |
0.000 |
0.000 |
0.810 |
|
| Polar |
|---|
| 3z2-r2 | 1.620 |
|---|
| x2-y2 | -3.675 |
|---|
| xy | 0.983 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.306 |
0.040 |
0.000 |
| y |
0.040 |
7.093 |
0.000 |
| z |
0.000 |
0.000 |
7.481 |
<r2> (average value of r
2) Å
2
| <r2> |
108.581 |
| (<r2>)1/2 |
10.420 |