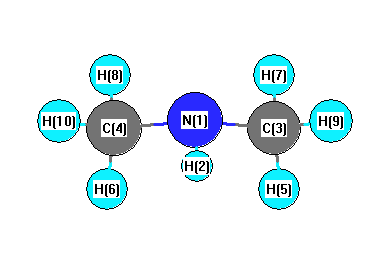
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3780 |
3418 |
0.48 |
|
|
|
| 2 |
A' |
3251 |
2940 |
74.88 |
|
|
|
| 3 |
A' |
3213 |
2906 |
79.45 |
|
|
|
| 4 |
A' |
3112 |
2815 |
185.71 |
|
|
|
| 5 |
A' |
1652 |
1494 |
2.31 |
|
|
|
| 6 |
A' |
1638 |
1481 |
10.09 |
|
|
|
| 7 |
A' |
1611 |
1457 |
1.34 |
|
|
|
| 8 |
A' |
1387 |
1254 |
0.63 |
|
|
|
| 9 |
A' |
1306 |
1181 |
18.29 |
|
|
|
| 10 |
A' |
1007 |
911 |
6.16 |
|
|
|
| 11 |
A' |
857 |
775 |
126.12 |
|
|
|
| 12 |
A' |
416 |
376 |
6.10 |
|
|
|
| 13 |
A' |
287 |
259 |
3.37 |
|
|
|
| 14 |
A" |
3249 |
2938 |
45.54 |
|
|
|
| 15 |
A" |
3208 |
2902 |
43.11 |
|
|
|
| 16 |
A" |
3103 |
2807 |
43.39 |
|
|
|
| 17 |
A" |
1665 |
1506 |
18.22 |
|
|
|
| 18 |
A" |
1630 |
1474 |
0.37 |
|
|
|
| 19 |
A" |
1620 |
1465 |
9.49 |
|
|
|
| 20 |
A" |
1579 |
1428 |
2.98 |
|
|
|
| 21 |
A" |
1274 |
1152 |
37.93 |
|
|
|
| 22 |
A" |
1194 |
1080 |
0.33 |
|
|
|
| 23 |
A" |
1127 |
1020 |
8.81 |
|
|
|
| 24 |
A" |
239 |
216 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21701.4 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 19626.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.594 |
-0.760 |
|
0.375 |
| 2 |
H |
0.312 |
0.370 |
|
-0.691 |
| 3 |
C |
-0.451 |
0.180 |
|
0.046 |
| 4 |
C |
-0.451 |
0.180 |
|
0.083 |
| 5 |
H |
0.176 |
-0.016 |
|
0.037 |
| 6 |
H |
0.176 |
-0.016 |
|
-0.014 |
| 7 |
H |
0.209 |
0.031 |
|
0.072 |
| 8 |
H |
0.209 |
0.031 |
|
0.034 |
| 9 |
H |
0.207 |
-0.000 |
|
0.024 |
| 10 |
H |
0.207 |
-0.000 |
|
0.033 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.036 |
-0.434 |
0.000 |
1.123 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.004 |
-0.270 |
1.074 |
1.107 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.784 |
0.001 |
0.001 |
| y |
0.001 |
-20.459 |
2.076 |
| z |
0.001 |
2.076 |
-21.770 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.330 |
0.001 |
0.001 |
| y |
0.001 |
0.318 |
2.076 |
| z |
0.001 |
2.076 |
-1.648 |
|
| Polar |
|---|
| 3z2-r2 | -3.296 |
|---|
| x2-y2 | 0.675 |
|---|
| xy | 0.001 |
|---|
| xz | 0.001 |
|---|
| yz | 2.076 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.884 |
0.000 |
0.000 |
| y |
0.000 |
4.264 |
0.066 |
| z |
0.000 |
0.066 |
4.020 |
<r2> (average value of r
2) Å
2
| <r2> |
56.844 |
| (<r2>)1/2 |
7.539 |