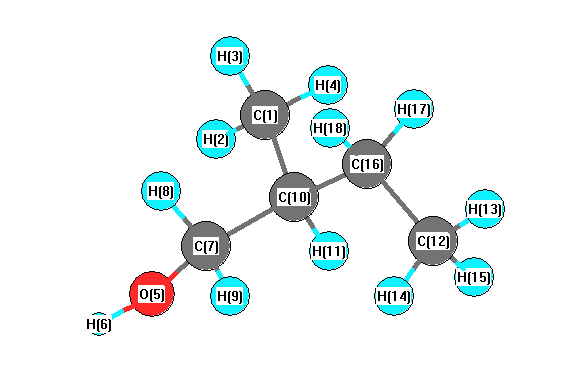
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3829 |
3687 |
12.49 |
|
|
|
| 2 |
A |
3137 |
3020 |
20.22 |
|
|
|
| 3 |
A |
3112 |
2996 |
59.01 |
|
|
|
| 4 |
A |
3103 |
2988 |
48.73 |
|
|
|
| 5 |
A |
3095 |
2980 |
43.95 |
|
|
|
| 6 |
A |
3056 |
2942 |
21.83 |
|
|
|
| 7 |
A |
3038 |
2925 |
62.63 |
|
|
|
| 8 |
A |
3033 |
2920 |
80.28 |
|
|
|
| 9 |
A |
3027 |
2914 |
10.10 |
|
|
|
| 10 |
A |
3017 |
2905 |
16.79 |
|
|
|
| 11 |
A |
3011 |
2899 |
22.47 |
|
|
|
| 12 |
A |
2965 |
2855 |
56.84 |
|
|
|
| 13 |
A |
1530 |
1473 |
2.32 |
|
|
|
| 14 |
A |
1525 |
1468 |
7.51 |
|
|
|
| 15 |
A |
1520 |
1463 |
11.69 |
|
|
|
| 16 |
A |
1514 |
1457 |
6.88 |
|
|
|
| 17 |
A |
1507 |
1450 |
9.56 |
|
|
|
| 18 |
A |
1495 |
1439 |
1.57 |
|
|
|
| 19 |
A |
1469 |
1414 |
2.92 |
|
|
|
| 20 |
A |
1423 |
1370 |
4.88 |
|
|
|
| 21 |
A |
1413 |
1360 |
9.98 |
|
|
|
| 22 |
A |
1397 |
1345 |
0.35 |
|
|
|
| 23 |
A |
1373 |
1321 |
3.63 |
|
|
|
| 24 |
A |
1338 |
1288 |
2.28 |
|
|
|
| 25 |
A |
1311 |
1263 |
14.18 |
|
|
|
| 26 |
A |
1283 |
1235 |
7.85 |
|
|
|
| 27 |
A |
1258 |
1211 |
29.89 |
|
|
|
| 28 |
A |
1192 |
1148 |
1.94 |
|
|
|
| 29 |
A |
1178 |
1134 |
4.74 |
|
|
|
| 30 |
A |
1134 |
1091 |
3.05 |
|
|
|
| 31 |
A |
1087 |
1047 |
85.64 |
|
|
|
| 32 |
A |
1055 |
1016 |
2.84 |
|
|
|
| 33 |
A |
1030 |
992 |
18.01 |
|
|
|
| 34 |
A |
982 |
945 |
6.74 |
|
|
|
| 35 |
A |
932 |
897 |
1.85 |
|
|
|
| 36 |
A |
923 |
889 |
1.76 |
|
|
|
| 37 |
A |
831 |
800 |
4.29 |
|
|
|
| 38 |
A |
765 |
737 |
6.04 |
|
|
|
| 39 |
A |
496 |
478 |
7.78 |
|
|
|
| 40 |
A |
444 |
427 |
1.55 |
|
|
|
| 41 |
A |
389 |
374 |
2.59 |
|
|
|
| 42 |
A |
279 |
269 |
40.47 |
|
|
|
| 43 |
A |
278 |
268 |
67.52 |
|
|
|
| 44 |
A |
251 |
242 |
23.29 |
|
|
|
| 45 |
A |
236 |
227 |
2.29 |
|
|
|
| 46 |
A |
224 |
216 |
2.85 |
|
|
|
| 47 |
A |
118 |
114 |
4.05 |
|
|
|
| 48 |
A |
73 |
70 |
3.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36338.4 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 34983.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.645 |
|
|
|
| 2 |
H |
0.247 |
|
|
|
| 3 |
H |
0.206 |
|
|
|
| 4 |
H |
0.217 |
|
|
|
| 5 |
O |
-0.578 |
|
|
|
| 6 |
H |
0.391 |
|
|
|
| 7 |
C |
-0.169 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.203 |
|
|
|
| 10 |
C |
-0.281 |
|
|
|
| 11 |
H |
0.218 |
|
|
|
| 12 |
C |
-0.656 |
|
|
|
| 13 |
H |
0.224 |
|
|
|
| 14 |
H |
0.217 |
|
|
|
| 15 |
H |
0.220 |
|
|
|
| 16 |
C |
-0.429 |
|
|
|
| 17 |
H |
0.222 |
|
|
|
| 18 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.388 |
-1.101 |
1.058 |
1.575 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.050 |
3.487 |
-2.664 |
| y |
3.487 |
-37.493 |
-1.691 |
| z |
-2.664 |
-1.691 |
-40.142 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.233 |
3.487 |
-2.664 |
| y |
3.487 |
2.603 |
-1.691 |
| z |
-2.664 |
-1.691 |
-1.370 |
|
| Polar |
|---|
| 3z2-r2 | -2.741 |
|---|
| x2-y2 | -2.557 |
|---|
| xy | 3.487 |
|---|
| xz | -2.664 |
|---|
| yz | -1.691 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
215.367 |
| (<r2>)1/2 |
14.675 |