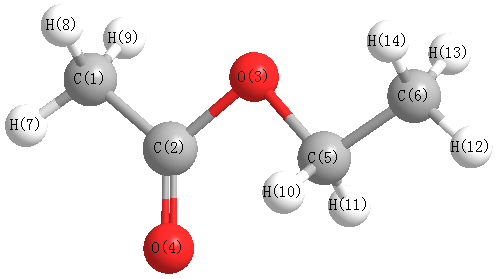
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3178 |
3059 |
9.50 |
|
|
|
| 2 |
A' |
3126 |
3010 |
29.21 |
|
|
|
| 3 |
A' |
3066 |
2952 |
20.88 |
|
|
|
| 4 |
A' |
3064 |
2949 |
0.97 |
|
|
|
| 5 |
A' |
3051 |
2937 |
19.71 |
|
|
|
| 6 |
A' |
1833 |
1765 |
258.32 |
|
|
|
| 7 |
A' |
1532 |
1475 |
7.59 |
|
|
|
| 8 |
A' |
1511 |
1454 |
3.14 |
|
|
|
| 9 |
A' |
1484 |
1429 |
16.33 |
|
|
|
| 10 |
A' |
1439 |
1385 |
6.37 |
|
|
|
| 11 |
A' |
1410 |
1358 |
64.54 |
|
|
|
| 12 |
A' |
1394 |
1342 |
23.54 |
|
|
|
| 13 |
A' |
1279 |
1232 |
412.39 |
|
|
|
| 14 |
A' |
1147 |
1104 |
16.61 |
|
|
|
| 15 |
A' |
1088 |
1047 |
105.61 |
|
|
|
| 16 |
A' |
1017 |
979 |
8.54 |
|
|
|
| 17 |
A' |
960 |
924 |
6.14 |
|
|
|
| 18 |
A' |
872 |
839 |
8.61 |
|
|
|
| 19 |
A' |
645 |
621 |
6.12 |
|
|
|
| 20 |
A' |
429 |
413 |
0.79 |
|
|
|
| 21 |
A' |
370 |
356 |
9.66 |
|
|
|
| 22 |
A' |
194 |
187 |
5.07 |
|
|
|
| 23 |
A" |
3135 |
3018 |
47.19 |
|
|
|
| 24 |
A" |
3130 |
3013 |
6.80 |
|
|
|
| 25 |
A" |
3103 |
2987 |
6.14 |
|
|
|
| 26 |
A" |
1499 |
1443 |
8.81 |
|
|
|
| 27 |
A" |
1487 |
1432 |
10.90 |
|
|
|
| 28 |
A" |
1311 |
1262 |
0.88 |
|
|
|
| 29 |
A" |
1185 |
1141 |
4.37 |
|
|
|
| 30 |
A" |
1068 |
1028 |
7.30 |
|
|
|
| 31 |
A" |
812 |
782 |
0.89 |
|
|
|
| 32 |
A" |
610 |
587 |
4.71 |
|
|
|
| 33 |
A" |
267 |
257 |
1.08 |
|
|
|
| 34 |
A" |
154 |
149 |
4.40 |
|
|
|
| 35 |
A" |
64 |
62 |
0.27 |
|
|
|
| 36 |
A" |
46 |
45 |
0.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25979.7 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 25010.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.724 |
|
|
|
| 2 |
C |
0.414 |
|
|
|
| 3 |
O |
-0.308 |
|
|
|
| 4 |
O |
-0.356 |
|
|
|
| 5 |
C |
-0.250 |
|
|
|
| 6 |
C |
-0.668 |
|
|
|
| 7 |
H |
0.253 |
|
|
|
| 8 |
H |
0.249 |
|
|
|
| 9 |
H |
0.249 |
|
|
|
| 10 |
H |
0.229 |
|
|
|
| 11 |
H |
0.229 |
|
|
|
| 12 |
H |
0.226 |
|
|
|
| 13 |
H |
0.229 |
|
|
|
| 14 |
H |
0.229 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.298 |
1.916 |
0.000 |
1.939 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.547 |
-1.049 |
0.000 |
| y |
-1.049 |
-42.608 |
0.000 |
| z |
0.000 |
0.000 |
-35.388 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.451 |
-1.049 |
0.000 |
| y |
-1.049 |
-9.640 |
0.000 |
| z |
0.000 |
0.000 |
1.189 |
|
| Polar |
|---|
| 3z2-r2 | 2.379 |
|---|
| x2-y2 | 12.061 |
|---|
| xy | -1.049 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.935 |
0.482 |
0.000 |
| y |
0.482 |
7.698 |
0.000 |
| z |
0.000 |
0.000 |
5.941 |
<r2> (average value of r
2) Å
2
| <r2> |
200.719 |
| (<r2>)1/2 |
14.168 |