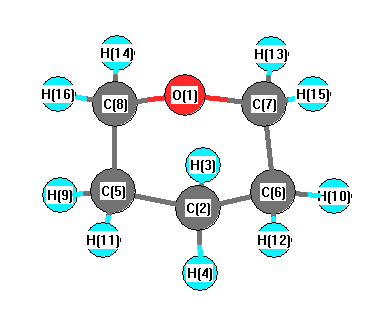
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3102 |
2987 |
41.27 |
|
|
|
| 2 |
A' |
3084 |
2969 |
78.19 |
|
|
|
| 3 |
A' |
3072 |
2958 |
44.48 |
|
|
|
| 4 |
A' |
3034 |
2921 |
18.30 |
|
|
|
| 5 |
A' |
3018 |
2906 |
29.35 |
|
|
|
| 6 |
A' |
2962 |
2852 |
119.05 |
|
|
|
| 7 |
A' |
1516 |
1460 |
4.68 |
|
|
|
| 8 |
A' |
1500 |
1444 |
8.39 |
|
|
|
| 9 |
A' |
1489 |
1433 |
9.29 |
|
|
|
| 10 |
A' |
1429 |
1375 |
6.21 |
|
|
|
| 11 |
A' |
1386 |
1334 |
1.74 |
|
|
|
| 12 |
A' |
1339 |
1289 |
4.90 |
|
|
|
| 13 |
A' |
1295 |
1247 |
5.74 |
|
|
|
| 14 |
A' |
1180 |
1136 |
3.19 |
|
|
|
| 15 |
A' |
1060 |
1020 |
10.36 |
|
|
|
| 16 |
A' |
1025 |
987 |
10.01 |
|
|
|
| 17 |
A' |
900 |
866 |
23.18 |
|
|
|
| 18 |
A' |
865 |
833 |
5.66 |
|
|
|
| 19 |
A' |
835 |
804 |
2.29 |
|
|
|
| 20 |
A' |
562 |
541 |
3.64 |
|
|
|
| 21 |
A' |
437 |
420 |
1.62 |
|
|
|
| 22 |
A' |
402 |
387 |
4.65 |
|
|
|
| 23 |
A' |
247 |
238 |
3.72 |
|
|
|
| 24 |
A" |
3100 |
2984 |
56.04 |
|
|
|
| 25 |
A" |
3077 |
2963 |
40.51 |
|
|
|
| 26 |
A" |
3033 |
2920 |
35.88 |
|
|
|
| 27 |
A" |
2955 |
2845 |
23.40 |
|
|
|
| 28 |
A" |
1502 |
1446 |
1.97 |
|
|
|
| 29 |
A" |
1479 |
1424 |
3.44 |
|
|
|
| 30 |
A" |
1400 |
1348 |
6.19 |
|
|
|
| 31 |
A" |
1385 |
1334 |
0.87 |
|
|
|
| 32 |
A" |
1358 |
1308 |
0.24 |
|
|
|
| 33 |
A" |
1313 |
1264 |
8.43 |
|
|
|
| 34 |
A" |
1238 |
1192 |
42.27 |
|
|
|
| 35 |
A" |
1201 |
1157 |
0.03 |
|
|
|
| 36 |
A" |
1133 |
1091 |
76.86 |
|
|
|
| 37 |
A" |
1070 |
1030 |
12.04 |
|
|
|
| 38 |
A" |
988 |
952 |
3.88 |
|
|
|
| 39 |
A" |
895 |
861 |
0.37 |
|
|
|
| 40 |
A" |
821 |
791 |
0.50 |
|
|
|
| 41 |
A" |
464 |
447 |
0.01 |
|
|
|
| 42 |
A" |
241 |
232 |
0.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32196.1 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 30995.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.331 |
|
|
|
| 2 |
C |
-0.455 |
|
|
|
| 3 |
H |
0.213 |
|
|
|
| 4 |
H |
0.226 |
|
|
|
| 5 |
C |
-0.438 |
|
|
|
| 6 |
C |
-0.438 |
|
|
|
| 7 |
C |
-0.239 |
|
|
|
| 8 |
C |
-0.239 |
|
|
|
| 9 |
H |
0.218 |
|
|
|
| 10 |
H |
0.218 |
|
|
|
| 11 |
H |
0.221 |
|
|
|
| 12 |
H |
0.221 |
|
|
|
| 13 |
H |
0.192 |
|
|
|
| 14 |
H |
0.192 |
|
|
|
| 15 |
H |
0.219 |
|
|
|
| 16 |
H |
0.219 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.367 |
-1.387 |
0.000 |
1.435 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.997 |
-0.336 |
0.000 |
| y |
-0.336 |
-41.470 |
0.000 |
| z |
0.000 |
0.000 |
-35.470 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.474 |
-0.336 |
0.000 |
| y |
-0.336 |
-4.737 |
0.000 |
| z |
0.000 |
0.000 |
4.263 |
|
| Polar |
|---|
| 3z2-r2 | 8.526 |
|---|
| x2-y2 | 3.473 |
|---|
| xy | -0.336 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
152.120 |
| (<r2>)1/2 |
12.334 |