Jump to
S1C2
S1C3
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -254.963898 |
| Energy at 298.15K | -254.963320 |
| HF Energy | -254.963898 |
| Nuclear repulsion energy | 46.684631 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.687 |
| N2 |
0.000 |
0.000 |
-1.850 |
| Na3 |
0.000 |
0.000 |
1.552 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1628 | 2.2396 |
N2 | 1.1628 | | 3.4024 | Na3 | 2.2396 | 3.4024 | |
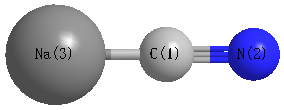 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.346 |
|
|
|
| 2 |
N |
-0.331 |
|
|
|
| 3 |
Na |
0.677 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.509 |
10.509 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.091 |
0.000 |
0.000 |
| y |
0.000 |
-17.091 |
0.000 |
| z |
0.000 |
0.000 |
-13.239 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.926 |
0.000 |
0.000 |
| y |
0.000 |
-1.926 |
0.000 |
| z |
0.000 |
0.000 |
3.852 |
|
| Polar |
|---|
| 3z2-r2 | 7.704 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.622 |
0.000 |
-0.000 |
| y |
0.000 |
2.622 |
0.000 |
| z |
-0.000 |
0.000 |
5.522 |
<r2> (average value of r
2) Å
2
| <r2> |
63.172 |
| (<r2>)1/2 |
7.948 |
Jump to
S1C1
S1C3
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -254.968681 |
| Energy at 298.15K | -254.968445 |
| HF Energy | -254.968681 |
| Nuclear repulsion energy | 51.938037 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.085 |
0.645 |
0.000 |
| N2 |
0.000 |
1.093 |
0.000 |
| Na3 |
-0.592 |
-1.047 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1741 | 2.3820 |
N2 | 1.1741 | | 2.2210 | Na3 | 2.3820 | 2.2210 | |
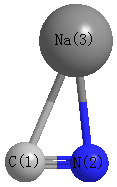 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
82.979 |
|
C1 |
Na3 |
N2 |
29.289 |
| N2 |
C1 |
Na3 |
67.731 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.189 |
|
|
|
| 2 |
N |
-0.402 |
|
|
|
| 3 |
Na |
0.591 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.567 |
-7.297 |
0.000 |
8.608 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.567 |
3.415 |
0.000 |
| y |
3.415 |
-14.098 |
0.000 |
| z |
0.000 |
0.000 |
-17.240 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.898 |
3.415 |
0.000 |
| y |
3.415 |
4.305 |
0.000 |
| z |
0.000 |
0.000 |
-0.408 |
|
| Polar |
|---|
| 3z2-r2 | -0.816 |
|---|
| x2-y2 | -5.469 |
|---|
| xy | 3.415 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.980 |
-0.016 |
-0.000 |
| y |
-0.016 |
3.550 |
0.000 |
| z |
-0.000 |
0.000 |
2.801 |
<r2> (average value of r
2) Å
2
| <r2> |
44.440 |
| (<r2>)1/2 |
6.666 |
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -254.964748 |
| Energy at 298.15K | -254.963900 |
| HF Energy | -254.964748 |
| Nuclear repulsion energy | 48.921356 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.846 |
| N2 |
0.000 |
0.000 |
-0.673 |
| Na3 |
0.000 |
0.000 |
1.435 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1734 | 3.2808 |
N2 | 1.1734 | | 2.1074 | Na3 | 3.2808 | 2.1074 | |
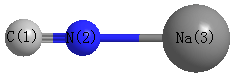 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.204 |
|
|
|
| 2 |
N |
-0.514 |
|
|
|
| 3 |
Na |
0.717 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.672 |
10.672 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.015 |
0.000 |
0.000 |
| y |
0.000 |
-17.015 |
0.000 |
| z |
0.000 |
0.000 |
-15.862 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.577 |
0.000 |
0.000 |
| y |
0.000 |
-0.577 |
0.000 |
| z |
0.000 |
0.000 |
1.153 |
|
| Polar |
|---|
| 3z2-r2 | 2.307 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.534 |
0.000 |
0.000 |
| y |
0.000 |
2.534 |
0.000 |
| z |
0.000 |
0.000 |
5.482 |
<r2> (average value of r
2) Å
2
| <r2> |
56.646 |
| (<r2>)1/2 |
7.526 |
