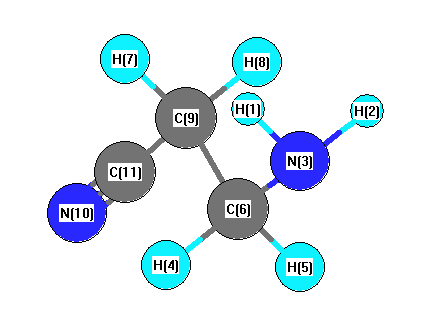
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3751 |
3392 |
0.01 |
|
|
|
| 2 |
A' |
3224 |
2916 |
45.30 |
|
|
|
| 3 |
A' |
3212 |
2905 |
4.68 |
|
|
|
| 4 |
A' |
2594 |
2346 |
14.95 |
|
|
|
| 5 |
A' |
1856 |
1679 |
37.29 |
|
|
|
| 6 |
A' |
1634 |
1478 |
3.54 |
|
|
|
| 7 |
A' |
1607 |
1454 |
3.85 |
|
|
|
| 8 |
A' |
1537 |
1391 |
16.61 |
|
|
|
| 9 |
A' |
1428 |
1291 |
4.93 |
|
|
|
| 10 |
A' |
1189 |
1076 |
15.67 |
|
|
|
| 11 |
A' |
1104 |
998 |
62.55 |
|
|
|
| 12 |
A' |
998 |
903 |
19.91 |
|
|
|
| 13 |
A' |
908 |
821 |
209.09 |
|
|
|
| 14 |
A' |
577 |
521 |
0.59 |
|
|
|
| 15 |
A' |
409 |
370 |
9.09 |
|
|
|
| 16 |
A' |
184 |
167 |
9.17 |
|
|
|
| 17 |
A" |
3829 |
3463 |
0.28 |
|
|
|
| 18 |
A" |
3271 |
2958 |
46.84 |
|
|
|
| 19 |
A" |
3244 |
2934 |
0.03 |
|
|
|
| 20 |
A" |
1519 |
1373 |
0.07 |
|
|
|
| 21 |
A" |
1429 |
1292 |
0.15 |
|
|
|
| 22 |
A" |
1270 |
1149 |
0.00 |
|
|
|
| 23 |
A" |
1056 |
955 |
0.73 |
|
|
|
| 24 |
A" |
817 |
739 |
0.94 |
|
|
|
| 25 |
A" |
437 |
395 |
0.25 |
|
|
|
| 26 |
A" |
318 |
288 |
54.39 |
|
|
|
| 27 |
A" |
117 |
106 |
2.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21758.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 19678.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.332 |
0.378 |
|
0.386 |
| 2 |
H |
0.332 |
0.378 |
|
0.386 |
| 3 |
N |
-0.815 |
-1.052 |
|
-1.057 |
| 4 |
H |
0.233 |
-0.032 |
|
-0.004 |
| 5 |
H |
0.233 |
-0.032 |
|
-0.004 |
| 6 |
C |
-0.201 |
0.532 |
|
0.476 |
| 7 |
H |
0.252 |
0.077 |
|
0.108 |
| 8 |
H |
0.252 |
0.077 |
|
0.108 |
| 9 |
C |
-0.449 |
-0.175 |
|
-0.247 |
| 10 |
N |
-0.336 |
-0.498 |
|
-0.486 |
| 11 |
C |
0.169 |
0.345 |
|
0.335 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.785 |
1.658 |
0.000 |
4.132 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
3.844 |
1.638 |
0.000 |
4.179 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.803 |
4.965 |
0.000 |
| y |
4.965 |
-31.676 |
0.000 |
| z |
0.000 |
0.000 |
-28.107 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.912 |
4.965 |
0.000 |
| y |
4.965 |
4.279 |
0.000 |
| z |
0.000 |
0.000 |
9.633 |
|
| Polar |
|---|
| 3z2-r2 | 19.265 |
|---|
| x2-y2 | -12.127 |
|---|
| xy | 4.965 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.884 |
0.161 |
0.000 |
| y |
0.161 |
4.970 |
0.000 |
| z |
0.000 |
0.000 |
4.931 |
<r2> (average value of r
2) Å
2
| <r2> |
151.519 |
| (<r2>)1/2 |
12.309 |