Jump to
S1C2
S1C3
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -692.614997 |
| Energy at 298.15K | -692.614363 |
| HF Energy | -692.418257 |
| Nuclear repulsion energy | 60.978815 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.308 |
| C2 |
0.000 |
0.000 |
-1.280 |
| N3 |
0.000 |
0.000 |
-2.453 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.5878 | 3.7606 |
C2 | 2.5878 | | 1.1727 | N3 | 3.7606 | 1.1727 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -692.616973 |
| Energy at 298.15K | |
| HF Energy | -692.423192 |
| Nuclear repulsion energy | 64.413908 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.211 |
| C2 |
0.000 |
0.000 |
-2.405 |
| N3 |
0.000 |
0.000 |
-1.225 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6157 | 2.4352 |
C2 | 3.6157 | | 1.1804 | N3 | 2.4352 | 1.1804 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -692.622106 |
| Energy at 298.15K | -692.621996 |
| HF Energy | -692.425940 |
| Nuclear repulsion energy | 68.664029 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.040 |
0.000 |
| C2 |
0.629 |
-1.613 |
0.000 |
| N3 |
-0.539 |
-1.441 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7266 | 2.5386 |
C2 | 2.7266 | | 1.1813 | N3 | 2.5386 | 1.1813 | |
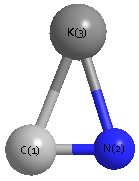 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
68.266 |
|
K1 |
N3 |
C2 |
86.124 |
| C2 |
K1 |
N3 |
25.610 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
