Jump to
S1C2
S1C3
Energy calculated at wB97X-D/6-311G*
| | hartrees |
|---|
| Energy at 0K | -692.770340 |
| Energy at 298.15K | -692.769696 |
| HF Energy | -692.770340 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at wB97X-D/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.307 |
| C2 |
0.000 |
0.000 |
-1.284 |
| N3 |
0.000 |
0.000 |
-2.447 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.5905 | 3.7533 |
C2 | 2.5905 | | 1.1628 | N3 | 3.7533 | 1.1628 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.769 |
|
|
|
| 2 |
C |
-0.453 |
|
|
|
| 3 |
N |
-0.316 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
12.212 |
12.212 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.844 |
0.000 |
0.000 |
| y |
0.000 |
-22.844 |
0.000 |
| z |
0.000 |
0.000 |
-30.484 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.820 |
0.000 |
0.000 |
| y |
0.000 |
3.820 |
0.000 |
| z |
0.000 |
0.000 |
-7.640 |
|
| Polar |
|---|
| 3z2-r2 | -15.279 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.995 |
0.000 |
0.000 |
| y |
0.000 |
2.995 |
0.000 |
| z |
0.000 |
0.000 |
5.823 |
<r2> (average value of r
2) Å
2
| <r2> |
100.092 |
| (<r2>)1/2 |
10.005 |
Jump to
S1C1
S1C3
Energy calculated at wB97X-D/6-311G*
| | hartrees |
|---|
| Energy at 0K | -692.773688 |
| Energy at 298.15K | |
| HF Energy | -692.773688 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at wB97X-D/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.209 |
| C2 |
0.000 |
0.000 |
-2.397 |
| N3 |
0.000 |
0.000 |
-1.225 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6060 | 2.4338 |
C2 | 3.6060 | | 1.1721 | N3 | 2.4338 | 1.1721 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.779 |
|
|
|
| 2 |
C |
-0.237 |
|
|
|
| 3 |
N |
-0.543 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
12.106 |
12.106 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.734 |
0.000 |
0.000 |
| y |
0.000 |
-22.734 |
0.000 |
| z |
0.000 |
0.000 |
-32.792 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.029 |
0.000 |
0.000 |
| y |
0.000 |
5.029 |
0.000 |
| z |
0.000 |
0.000 |
-10.058 |
|
| Polar |
|---|
| 3z2-r2 | -20.116 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.952 |
0.000 |
0.000 |
| y |
0.000 |
2.952 |
0.000 |
| z |
0.000 |
0.000 |
5.738 |
<r2> (average value of r
2) Å
2
| <r2> |
89.039 |
| (<r2>)1/2 |
9.436 |
Jump to
S1C1
S1C2
Energy calculated at wB97X-D/6-311G*
| | hartrees |
|---|
| Energy at 0K | -692.777344 |
| Energy at 298.15K | -692.777145 |
| HF Energy | -692.777344 |
| Nuclear repulsion energy | 68.592978 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at wB97X-D/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.047 |
0.000 |
| C2 |
0.619 |
-1.652 |
0.000 |
| N3 |
-0.531 |
-1.425 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7692 | 2.5276 |
C2 | 2.7692 | | 1.1720 | N3 | 2.5276 | 1.1720 | |
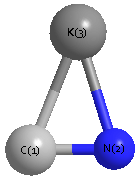 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
65.874 |
|
K1 |
N3 |
C2 |
89.091 |
| C2 |
K1 |
N3 |
25.035 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.721 |
|
|
|
| 2 |
C |
-0.279 |
|
|
|
| 3 |
N |
-0.442 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.410 |
10.253 |
0.000 |
10.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.078 |
1.159 |
0.000 |
| y |
1.159 |
-26.448 |
0.000 |
| z |
0.000 |
0.000 |
-22.975 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.366 |
1.159 |
0.000 |
| y |
1.159 |
-1.421 |
0.000 |
| z |
0.000 |
0.000 |
3.787 |
|
| Polar |
|---|
| 3z2-r2 | 7.575 |
|---|
| x2-y2 | -0.630 |
|---|
| xy | 1.159 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.282 |
-0.264 |
0.000 |
| y |
-0.264 |
4.259 |
0.000 |
| z |
0.000 |
0.000 |
3.131 |
<r2> (average value of r
2) Å
2
| <r2> |
71.603 |
| (<r2>)1/2 |
8.462 |
