return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (3-Penten-1-yne, (Z)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-193.538607
Energy at 298.15K-193.542826
HF Energy-192.781589
Nuclear repulsion energy141.005340
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3493 3308 56.36      
2 A' 3212 3042 21.12      
3 A' 3194 3025 5.31      
4 A' 3182 3014 1.52      
5 A' 3075 2912 15.93      
6 A' 2125 2013 0.54      
7 A' 1690 1601 1.73      
8 A' 1523 1442 10.34      
9 A' 1457 1380 6.80      
10 A' 1419 1344 6.13      
11 A' 1278 1210 0.16      
12 A' 1138 1078 0.44      
13 A' 985 933 17.38      
14 A' 919 871 1.68      
15 A' 661 626 4.12      
16 A' 589 558 45.35      
17 A' 367 348 4.34      
18 A' 156 147 0.38      
19 A" 3145 2979 14.93      
20 A" 1522 1442 9.56      
21 A" 1076 1019 1.40      
22 A" 949 899 0.01      
23 A" 735 696 25.19      
24 A" 530 502 66.41      
25 A" 412 390 4.19      
26 A" 240 228 0.19      
27 A" 125 118 0.52      

Unscaled Zero Point Vibrational Energy (zpe) 19597.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18560.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.39482 0.11157 0.08841

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 3.146 -0.971 0.000
C2 2.241 -0.410 0.000
C3 1.197 0.223 0.000
H4 0.120 2.082 0.000
C5 0.000 1.001 0.000
H6 -2.080 1.168 0.000
C7 -1.241 0.474 0.000
H8 -0.659 -1.593 0.000
H9 -2.160 -1.250 0.879
H10 -2.160 -1.250 -0.879
C11 -1.565 -0.986 0.000

Atom - Atom Distances (Å)
  H1 C2 C3 H4 C5 H6 C7 H8 H9 H10 C11
H11.06472.28584.29913.71325.64684.61913.85505.38585.38584.7106
C21.06471.22123.27282.64854.60033.59303.13214.56664.56663.8494
C32.28581.22122.14871.42773.41032.45102.59613.76993.76993.0145
H44.29913.27282.14871.08742.38172.10663.75634.13164.13163.4998
C53.71322.64851.42771.08742.08651.34862.67633.24133.24132.5291
H65.64684.60033.41032.38172.08651.08873.10532.57392.57392.2148
C74.61913.59302.45102.10661.34861.08872.14732.14202.14201.4952
H83.85503.13212.59613.75632.67633.10532.14731.77371.77371.0906
H95.38584.56663.76994.13163.24132.57392.14201.77371.75851.0943
H105.38584.56663.76994.13163.24132.57392.14201.77371.75851.0943
C114.71063.84943.01453.49982.52912.21481.49521.09061.09431.0943

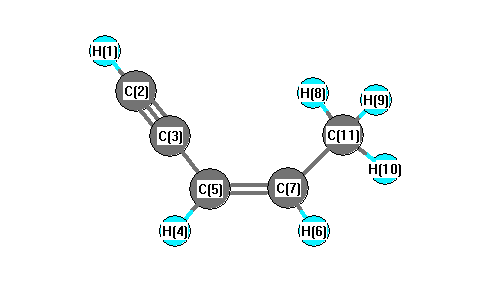
picture of 3-Penten-1-yne, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 179.367 C2 C3 C5 178.164
C3 C5 H4 116.727 C3 C5 C7 123.950
H4 C5 C7 119.323 C5 C7 H6 117.366
C5 C7 C11 125.504 H6 C7 C11 117.130
C7 C11 H8 111.329 C7 C11 H9 110.672
C7 C11 H10 110.672 H8 C11 H9 108.550
H8 C11 H10 108.550 H9 C11 H10 106.929
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability