return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-194.753564
Energy at 298.15K-194.760883
HF Energy-193.990327
Nuclear repulsion energy155.389087
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3281 3108 13.89      
2 A' 3214 3044 19.80      
3 A' 3197 3028 25.84      
4 A' 3187 3019 4.84      
5 A' 3181 3013 6.75      
6 A' 3170 3002 0.17      
7 A' 3076 2914 21.65      
8 A' 1726 1635 4.25      
9 A' 1657 1570 1.73      
10 A' 1531 1450 10.46      
11 A' 1491 1412 12.55      
12 A' 1451 1374 5.90      
13 A' 1408 1333 0.32      
14 A' 1339 1268 0.89      
15 A' 1306 1237 1.09      
16 A' 1208 1144 1.45      
17 A' 1082 1024 0.01      
18 A' 984 932 9.84      
19 A' 910 862 1.34      
20 A' 619 586 4.87      
21 A' 392 371 3.37      
22 A' 226 214 0.20      
23 A" 3142 2976 16.78      
24 A" 1521 1441 9.44      
25 A" 1073 1016 1.52      
26 A" 1008 955 23.85      
27 A" 946 896 2.76      
28 A" 884 837 44.26      
29 A" 785 743 11.82      
30 A" 624 591 34.26      
31 A" 341 323 0.39      
32 A" 114 108 0.11      
33 A" 95 90 0.47      

Unscaled Zero Point Vibrational Energy (zpe) 25085.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23758.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.52962 0.08793 0.07647

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.394 0.159 0.000
C2 1.102 -0.215 0.000
C3 0.000 0.740 0.000
C4 -1.318 0.450 0.000
C5 -1.949 -0.910 0.000
H6 2.679 1.207 0.000
H7 3.196 -0.570 0.000
H8 0.869 -1.276 0.000
H9 0.287 1.791 0.000
H10 -2.012 1.289 0.000
H11 -1.217 -1.717 0.000
H12 -2.589 -1.036 0.879
H13 -2.589 -1.036 -0.879

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.34552.46373.72324.47281.08621.08422.09482.66564.54874.07005.19935.1993
C21.34551.45782.50893.12882.12372.12441.08702.16523.45772.76343.88173.8817
C32.46371.45781.34932.55362.71963.45442.19561.08972.08552.74253.26013.2601
C43.72322.50891.34931.49904.06804.62762.78562.09151.08932.16942.14352.1435
C54.47283.12882.55361.49905.08945.15622.84143.50642.19981.08961.09441.0944
H61.08622.12372.71964.06805.08941.85153.07352.46274.69184.87205.79285.7928
H71.08422.12443.45444.62765.15621.85152.43203.74745.53004.56005.87005.8700
H82.09481.08702.19562.78562.84143.07352.43203.12253.85742.13223.57593.5759
H92.66562.16521.08972.09153.50642.46273.74743.12252.35293.81764.12724.1272
H104.54873.45772.08551.08932.19984.69185.53003.85742.35293.10962.55122.5512
H114.07002.76342.74252.16941.08964.87204.56002.13223.81763.10961.76591.7659
H125.19933.88173.26012.14351.09445.79285.87003.57594.12722.55121.76591.7572
H135.19933.88173.26012.14351.09445.79285.87003.57594.12722.55121.76591.7572

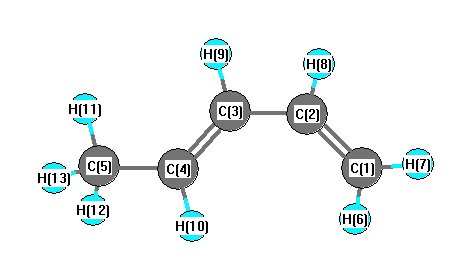
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 122.959 C1 C2 H8 118.502
C2 C1 H6 121.337 C2 C1 H7 121.567
C2 C3 C4 126.661 C2 C3 H9 115.654
C3 C2 H8 118.539 C3 C4 C5 127.333
C3 C4 H10 117.171 C4 C3 H9 117.685
C4 C5 H11 112.919 C4 C5 H12 110.518
C4 C5 H13 110.518 C5 C4 H10 115.496
H6 C1 H7 117.097 H11 C5 H12 107.919
H11 C5 H13 107.919 H12 C5 H13 106.810
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability