return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (2H-Thiopyran, tetrahydro-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-593.774583
Energy at 298.15K-593.787032
HF Energy-592.736482
Nuclear repulsion energy312.839959
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3148 2981 29.63      
2 A 3134 2968 37.98      
3 A 3120 2955 29.34      
4 A 3076 2913 20.78      
5 A 3071 2908 28.76      
6 A 3063 2901 19.94      
7 A 1535 1453 4.24      
8 A 1522 1442 14.79      
9 A 1504 1425 11.73      
10 A 1410 1336 0.06      
11 A 1381 1308 5.30      
12 A 1304 1235 0.82      
13 A 1277 1210 7.75      
14 A 1118 1059 3.77      
15 A 1060 1004 0.94      
16 A 1001 949 12.19      
17 A 852 807 5.32      
18 A 845 800 0.53      
19 A 693 656 3.20      
20 A 515 488 0.94      
21 A 373 354 0.16      
22 A 350 331 0.08      
23 A 190 180 1.81      
24 A 3149 2982 5.87      
25 A 3128 2962 17.10      
26 A 3078 2915 25.24      
27 A 3071 2909 37.77      
28 A 1513 1433 5.06      
29 A 1497 1418 2.36      
30 A 1416 1341 0.02      
31 A 1395 1321 0.29      
32 A 1343 1272 17.13      
33 A 1320 1250 1.42      
34 A 1198 1135 0.03      
35 A 1136 1076 0.61      
36 A 1125 1066 0.29      
37 A 975 923 0.40      
38 A 930 881 4.48      
39 A 819 776 0.07      
40 A 720 682 0.09      
41 A 396 375 0.30      
42 A 247 234 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 31998.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 30305.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.13429 0.10033 0.06420

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 1.505 0.000
C2 0.080 -1.720 0.000
C3 0.388 -0.934 1.274
C4 0.388 -0.934 -1.274
C5 -0.396 0.371 1.356
C6 -0.396 0.371 -1.356
H7 0.652 -2.654 0.000
H8 -0.981 -2.003 0.000
H9 -1.474 0.170 1.355
H10 -1.474 0.170 -1.355
H11 0.150 -1.549 2.151
H12 0.150 -1.549 -2.151
H13 1.459 -0.709 1.317
H14 1.459 -0.709 -1.317
H15 -0.162 0.908 -2.279
H16 -0.162 0.908 2.279

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S13.22602.77902.77901.81171.81174.20983.64342.40632.40633.73823.73822.96082.96082.36142.3614
C23.22601.52851.52852.53652.53651.09511.09812.79672.79672.15862.15862.15862.15863.48613.4861
C32.77901.52852.54831.52423.03842.15682.15462.16603.40551.09693.48771.09552.81294.03912.1682
C42.77901.52852.54833.03841.52422.15682.15463.40552.16603.48771.09692.81291.09552.16824.0391
C51.81172.53651.52423.03842.71113.47602.79571.09582.92372.14814.03432.14713.42803.68121.0933
C61.81172.53653.03841.52422.71113.47602.79572.92371.09584.03432.14813.42802.14711.09333.6812
H74.20981.09512.15682.15683.47603.47601.75723.78523.78522.46942.46942.48402.48404.30544.3054
H83.64341.09812.15462.15462.79572.79571.75722.60842.60842.47212.47213.06043.06043.78653.7865
H92.40632.79672.16603.40551.09582.92373.78522.60842.70992.49484.22853.06224.06393.93301.7658
H102.40632.79673.40552.16602.92371.09583.78522.60842.70994.22852.49484.06393.06221.76583.9330
H113.73822.15861.09693.48772.14814.03432.46942.47212.49484.22854.30131.76493.80065.07432.4791
H123.73822.15863.48771.09694.03432.14812.46942.47214.22852.49484.30133.80061.76492.47915.0743
H132.96082.15861.09552.81292.14713.42802.48403.06043.06224.06391.76493.80062.63404.26252.4829
H142.96082.15862.81291.09553.42802.14712.48403.06044.06393.06223.80061.76492.63402.48294.2625
H152.36143.48614.03912.16823.68121.09334.30543.78653.93301.76585.07432.47914.26252.48294.5574
H162.36143.48612.16824.03911.09333.68124.30543.78651.76583.93302.47915.07432.48294.26254.5574

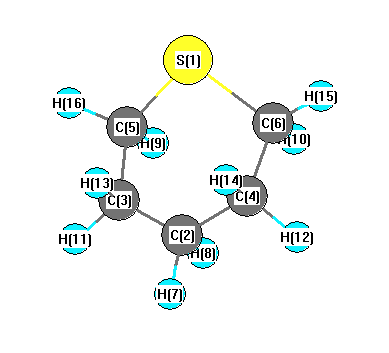
picture of 2H-Thiopyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C5 C3 112.543 S1 C5 H9 109.220
S1 C5 H16 106.096 S1 C6 C4 112.543
S1 C6 H10 109.220 S1 C6 H15 106.096
C2 C3 C5 112.380 C2 C3 H11 109.514
C2 C3 H13 109.598 C2 C4 C6 112.380
C2 C4 H12 109.514 C2 C4 H14 109.598
C3 C2 C4 112.940 C3 C2 H7 109.472
C3 C2 H8 109.132 C3 C5 H9 110.458
C3 C5 H16 110.780 C4 C2 H7 109.472
C4 C2 H8 109.132 C4 C6 H10 110.458
C4 C6 H15 110.780 C5 S1 C6 96.875
C5 C3 H11 108.987 C5 C3 H13 108.996
C6 C4 H12 108.987 C6 C4 H14 108.996
H7 C2 H8 106.487 H9 C5 H16 107.532
H10 C6 H15 107.532 H11 C3 H13 107.228
H12 C4 H14 107.228
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability