return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H5Cl (1-chloro-1-propene(E))

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-576.667487
Energy at 298.15K-576.672294
HF Energy-576.018193
Nuclear repulsion energy141.623253
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3243 3071 10.86      
2 A' 3209 3040 7.13      
3 A' 3175 3007 13.48      
4 A' 3077 2915 21.59      
5 A' 1706 1616 20.16      
6 A' 1531 1450 13.15      
7 A' 1454 1377 3.60      
8 A' 1341 1270 3.66      
9 A' 1306 1237 12.04      
10 A' 1142 1082 1.34      
11 A' 994 941 33.51      
12 A' 840 796 31.34      
13 A' 433 410 3.34      
14 A' 263 250 0.57      
15 A" 3154 2987 14.66      
16 A" 1519 1438 10.41      
17 A" 1081 1024 0.15      
18 A" 960 909 62.01      
19 A" 751 711 0.55      
20 A" 236 224 0.66      
21 A" 192 182 0.81      

Unscaled Zero Point Vibrational Energy (zpe) 15804.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14968.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.38628 0.08148 0.07807

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.454 0.000
C2 0.915 -0.517 0.000
C3 2.389 -0.247 0.000
Cl4 -1.705 0.144 0.000
H5 0.244 1.511 0.000
H6 0.583 -1.552 0.000
H7 2.599 0.824 0.000
H8 2.865 -0.688 0.880
H9 2.865 -0.688 -0.880

Atom - Atom Distances (Å)
  C1 C2 C3 Cl4 H5 H6 H7 H8 H9
C11.33452.48991.73281.08412.08962.62563.20743.2074
C21.33451.49852.70212.13551.08762.15302.14612.1461
C32.48991.49854.11262.77282.22901.09171.09351.0935
Cl41.73282.70214.11262.38042.84784.35774.72754.7275
H51.08412.13552.77282.38043.08152.45293.53213.5321
H62.08961.08762.22902.84783.08153.11722.59452.5945
H72.62562.15301.09174.35772.45293.11721.76991.7699
H83.20742.14611.09354.72753.53212.59451.76991.7604
H93.20742.14611.09354.72753.53212.59451.76991.7604

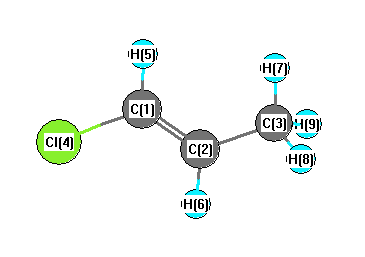
picture of 1-chloro-1-propene(E) state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 122.917 C1 C2 H6 118.892
C2 C1 Cl4 122.984 C2 C1 H5 123.675
C2 C3 H7 111.488 C2 C3 H8 110.812
C2 C3 H9 110.812 C3 C2 H6 118.191
Cl4 C1 H5 113.341 H7 C3 H8 108.183
H7 C3 H9 108.183 H8 C3 H9 107.207
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability