return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (Cyclopropane, 1,1-dimethyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-195.953991
Energy at 298.15K-195.964861
HF Energy-195.164907
Nuclear repulsion energy184.726786
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3173 3005 12.98      
2 A 3159 2992 32.47      
3 A 3063 2901 33.25      
4 A 1557 1475 0.08      
5 A 1540 1458 9.93      
6 A 1457 1379 0.48      
7 A 1395 1321 0.34      
8 A 1110 1051 0.57      
9 A 1058 1002 16.61      
10 A 978 927 12.93      
11 A 705 668 1.59      
12 A 359 340 0.03      
13 A 3259 3087 0.00      
14 A 3143 2977 0.00      
15 A 1517 1437 0.00      
16 A 1208 1144 0.00      
17 A 1087 1030 0.00      
18 A 921 872 0.00      
19 A 340 322 0.00      
20 A 224 212 0.00      
21 A 3167 3000 17.34      
22 A 3145 2979 56.18      
23 A 1538 1457 9.21      
24 A 1498 1419 7.82      
25 A 1185 1123 4.24      
26 A 1073 1017 1.87      
27 A 886 839 6.21      
28 A 397 376 1.27      
29 A 254 241 0.01      
30 A 3272 3099 21.20      
31 A 3158 2991 14.13      
32 A 3060 2898 31.95      
33 A 1531 1450 6.68      
34 A 1447 1370 14.19      
35 A 1357 1285 0.06      
36 A 993 941 0.66      
37 A 978 926 0.32      
38 A 806 764 0.03      
39 A 365 346 0.85      

Unscaled Zero Point Vibrational Energy (zpe) 30680.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29057.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.20589 0.17403 0.12763

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.152
C2 0.757 0.000 -1.150
C3 -0.757 0.000 -1.150
C4 0.000 1.271 0.967
C5 0.000 -1.271 0.967
H6 1.261 0.913 -1.448
H7 1.261 -0.913 -1.448
H8 -1.261 -0.913 -1.448
H9 -1.261 0.913 -1.448
H10 -0.886 1.323 1.609
H11 -0.886 -1.323 1.609
H12 0.886 -1.323 1.609
H13 0.886 1.323 1.609
H14 0.000 2.154 0.322
H15 0.000 -2.154 0.322

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.50621.50621.51001.51002.23262.23262.23262.23262.15782.15782.15782.15782.16032.1603
C21.50621.51432.58292.58291.08481.08482.23532.23533.47273.47273.06233.06232.71612.7161
C31.50621.51432.58292.58292.23532.23531.08481.08483.06233.06233.47273.47272.71612.7161
C41.51002.58292.58292.54232.74783.49203.49202.74781.09492.81502.81501.09491.09333.4851
C51.51002.58292.58292.54233.49202.74782.74783.49202.81501.09491.09492.81503.48511.0933
H62.23261.08482.23532.74783.49201.82663.11432.52243.75764.35343.80583.10682.50203.7587
H72.23261.08482.23533.49202.74781.82662.52243.11434.35343.75763.10683.80583.75872.5020
H82.23262.23531.08483.49202.74783.11432.52241.82663.80583.10683.75764.35343.75872.5020
H92.23262.23531.08482.74783.49202.52243.11431.82663.10683.80584.35343.75762.50203.7587
H102.15783.47273.06231.09492.81503.75764.35343.80583.10682.64543.18361.77121.76963.8113
H112.15783.47273.06232.81501.09494.35343.75763.10683.80582.64541.77123.18363.81131.7696
H122.15783.06233.47272.81501.09493.80583.10683.75764.35343.18361.77122.64543.81131.7696
H132.15783.06233.47271.09492.81503.10683.80584.35343.75761.77123.18362.64541.76963.8113
H142.16032.71612.71611.09333.48512.50203.75873.75872.50201.76963.81133.81131.76964.3073
H152.16032.71612.71613.48511.09333.75872.50202.50203.75873.81131.76961.76963.81134.3073

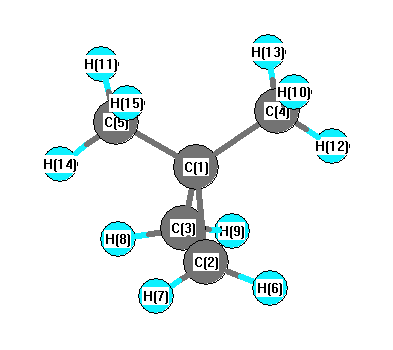
picture of Cyclopropane, 1,1-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 59.822 C1 C2 H6 118.092
C1 C2 H7 118.092 C1 C3 C2 59.822
C1 C3 H8 118.092 C1 C3 H9 118.092
C1 C4 H10 110.855 C1 C4 H13 110.855
C1 C4 H14 111.152 C1 C5 H11 110.855
C1 C5 H12 110.855 C1 C5 H15 111.152
C2 C1 C3 60.356 C2 C1 C4 117.814
C2 C1 C5 117.814 C2 C3 H8 117.687
C2 C3 H9 117.687 C3 C1 C4 117.814
C3 C1 C5 117.814 C3 C2 H6 117.687
C3 C2 H7 117.687 C4 C1 C5 114.666
H6 C2 H7 114.676 H8 C3 H9 114.676
H10 C4 H13 107.965 H10 C4 H14 107.940
H11 C5 H12 107.965 H11 C5 H15 107.940
H12 C5 H15 107.940 H13 C4 H14 107.940
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability