return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7N ((E)-3-Pentenenitrile)

using model chemistry: HF/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at HF/6-311G*
 hartrees
Energy at 0K-247.890272
Energy at 298.15K 
Nuclear repulsion energy195.337021
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
Rotational Constants (cm-1) from geometry optimized at HF/6-311G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at HF/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 3.667 0.198 -0.078
H2 2.951 -1.203 -0.815
H3 2.813 -0.983 0.918
C4 2.808 -0.459 -0.038
H5 1.545 1.013 -1.064
C6 1.531 0.289 -0.263
H7 0.374 -0.634 1.199
C8 0.406 0.080 0.387
C9 -0.868 0.811 0.125
N10 -2.939 -0.733 -0.188
C11 -2.029 -0.083 -0.056
H12 -0.771 1.451 -0.737
H13 -1.089 1.460 0.966

Atom - Atom Distances (Å)
  H1 H2 H3 C4 H5 C6 H7 C8 C9 N10 C11 H12 H13
H11.73751.76531.08252.47832.14623.62843.29674.58096.67285.70284.65875.0307
H21.73751.75201.08452.63662.13193.31873.09324.41845.94205.15984.57195.1561
H31.76531.75201.09003.08602.15832.47972.68524.17155.86355.02064.63824.6049
C41.08251.08451.09002.19521.49752.73562.49903.89315.75614.85164.11724.4593
H52.47832.63663.08602.19521.07983.03372.06722.69714.89113.87112.37943.3550
C62.14622.13192.15831.49751.07982.08021.31642.48564.58623.58492.62203.1223
H73.62843.31872.47972.73563.03372.08021.08192.18783.59362.76673.06722.5663
C83.29673.09322.68522.49902.06721.31641.08191.49173.49012.47972.12772.1154
C94.58094.41844.17153.89312.69712.48562.18781.49172.60261.47641.07761.0849
N106.67285.94205.86355.75614.89114.58623.59363.49012.60261.12663.12593.0926
C115.70285.15985.02064.85163.87113.58492.76672.47971.47641.12662.09682.0757
H124.65874.57194.63824.11722.37942.62203.06722.12771.07763.12592.09681.7319
H135.03075.15614.60494.45933.35503.12232.56632.11541.08493.09262.07571.7319

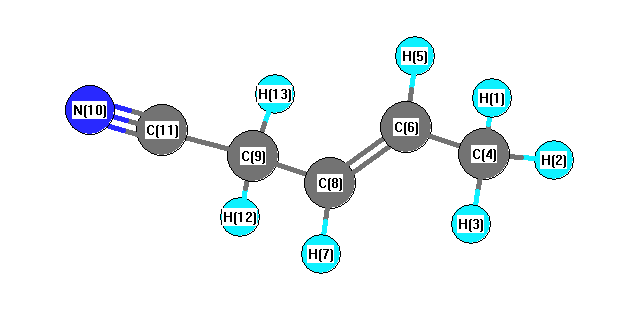
picture of (E)-3-Pentenenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H2 106.606 H1 C4 H3 108.690
H1 C4 C6 111.574 H2 C4 H3 107.349
H2 C4 C6 110.293 H3 C4 C6 112.088
C4 C6 H5 115.854 C4 C6 C8 125.146
H5 C6 C8 118.911 C6 C8 H7 119.998
C6 C8 C9 124.429 H7 C8 C9 115.528
C8 C9 C11 113.326 C8 C9 H12 110.783
C8 C9 H13 109.358 C9 C11 N10 177.894
C11 C9 H12 109.375 C11 C9 H13 107.284
H12 C9 H13 106.426
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.220 0.022   0.103
2 H 0.230 0.030   0.113
3 H 0.219 0.027   0.099
4 C -0.635 -0.002   -0.309
5 H 0.210 0.137   0.146
6 C -0.190 -0.139   -0.063
7 H 0.226 0.164   0.190
8 C -0.201 -0.278   -0.247
9 C -0.466 0.171   -0.189
10 N -0.338 -0.500   -0.509
11 C 0.180 0.325   0.418
12 H 0.270 0.021   0.123
13 H 0.275 0.022   0.127


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  3.831 2.229 0.371 4.448
CHELPG        
AIM        
ESP 3.849 2.246 0.369 4.471


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -47.750 -5.982 -2.196
y -5.982 -36.918 -1.596
z -2.196 -1.596 -35.861
Traceless
 xyz
x -11.361 -5.982 -2.196
y -5.982 4.887 -1.596
z -2.196 -1.596 6.474
Polar
3z2-r212.947
x2-y2-10.832
xy-5.982
xz-2.196
yz-1.596


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 236.449
(<r2>)1/2 15.377