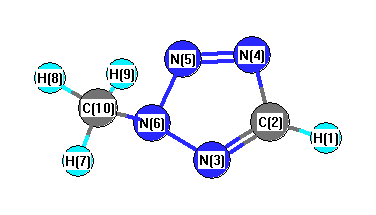
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3466 |
3135 |
3.10 |
|
|
|
| 2 |
A |
3346 |
3026 |
5.50 |
|
|
|
| 3 |
A |
3311 |
2995 |
18.31 |
|
|
|
| 4 |
A |
3236 |
2927 |
30.76 |
|
|
|
| 5 |
A |
1696 |
1534 |
13.11 |
|
|
|
| 6 |
A |
1642 |
1485 |
11.43 |
|
|
|
| 7 |
A |
1617 |
1462 |
10.03 |
|
|
|
| 8 |
A |
1595 |
1442 |
12.60 |
|
|
|
| 9 |
A |
1586 |
1434 |
29.90 |
|
|
|
| 10 |
A |
1491 |
1349 |
10.86 |
|
|
|
| 11 |
A |
1458 |
1319 |
25.13 |
|
|
|
| 12 |
A |
1413 |
1278 |
31.18 |
|
|
|
| 13 |
A |
1264 |
1143 |
0.30 |
|
|
|
| 14 |
A |
1259 |
1138 |
11.35 |
|
|
|
| 15 |
A |
1169 |
1057 |
32.36 |
|
|
|
| 16 |
A |
1150 |
1040 |
28.13 |
|
|
|
| 17 |
A |
1135 |
1026 |
8.79 |
|
|
|
| 18 |
A |
1014 |
917 |
10.09 |
|
|
|
| 19 |
A |
800 |
724 |
7.07 |
|
|
|
| 20 |
A |
771 |
698 |
23.33 |
|
|
|
| 21 |
A |
767 |
694 |
9.65 |
|
|
|
| 22 |
A |
406 |
367 |
5.63 |
|
|
|
| 23 |
A |
255 |
231 |
6.00 |
|
|
|
| 24 |
A |
83 |
75 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17964.8 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 16247.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.252 |
0.032 |
|
0.051 |
| 2 |
C |
0.091 |
0.574 |
|
0.560 |
| 3 |
N |
-0.260 |
-0.681 |
|
-0.690 |
| 4 |
N |
-0.285 |
-0.433 |
|
-0.444 |
| 5 |
N |
-0.019 |
-0.262 |
|
-0.262 |
| 6 |
N |
-0.103 |
0.733 |
|
0.769 |
| 7 |
H |
0.248 |
0.149 |
|
0.181 |
| 8 |
H |
0.252 |
0.108 |
|
0.142 |
| 9 |
H |
0.248 |
0.149 |
|
0.181 |
| 10 |
C |
-0.423 |
-0.369 |
|
-0.487 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.528 |
-1.459 |
-0.000 |
2.919 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-2.473 |
-1.456 |
-0.000 |
2.869 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.382 |
-4.368 |
0.001 |
| y |
-4.368 |
-40.230 |
-0.000 |
| z |
0.001 |
-0.000 |
-34.496 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.981 |
-4.368 |
0.001 |
| y |
-4.368 |
-8.791 |
-0.000 |
| z |
0.001 |
-0.000 |
-0.190 |
|
| Polar |
|---|
| 3z2-r2 | -0.381 |
|---|
| x2-y2 | 11.848 |
|---|
| xy | -4.368 |
|---|
| xz | 0.001 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.987 |
-0.057 |
-0.001 |
| y |
-0.057 |
6.182 |
0.000 |
| z |
-0.001 |
0.000 |
3.878 |
<r2> (average value of r
2) Å
2
| <r2> |
121.215 |
| (<r2>)1/2 |
11.010 |