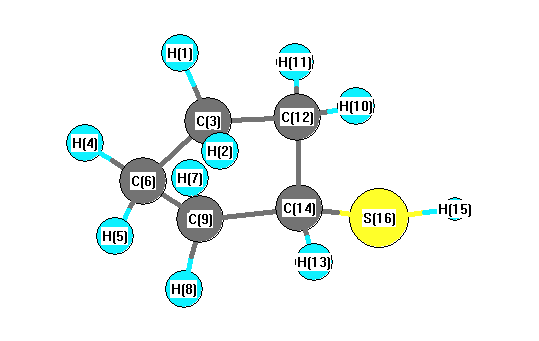
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3139 |
2996 |
79.30 |
|
|
|
| 2 |
A |
3127 |
2984 |
22.32 |
|
|
|
| 3 |
A |
3121 |
2978 |
8.46 |
|
|
|
| 4 |
A |
3107 |
2965 |
24.52 |
|
|
|
| 5 |
A |
3086 |
2945 |
41.98 |
|
|
|
| 6 |
A |
3077 |
2937 |
36.55 |
|
|
|
| 7 |
A |
3070 |
2930 |
27.81 |
|
|
|
| 8 |
A |
3067 |
2927 |
11.01 |
|
|
|
| 9 |
A |
3061 |
2922 |
10.74 |
|
|
|
| 10 |
A |
2663 |
2541 |
13.58 |
|
|
|
| 11 |
A |
1539 |
1469 |
1.43 |
|
|
|
| 12 |
A |
1515 |
1446 |
7.70 |
|
|
|
| 13 |
A |
1511 |
1442 |
5.39 |
|
|
|
| 14 |
A |
1501 |
1432 |
1.22 |
|
|
|
| 15 |
A |
1373 |
1310 |
8.46 |
|
|
|
| 16 |
A |
1362 |
1300 |
0.79 |
|
|
|
| 17 |
A |
1350 |
1288 |
0.95 |
|
|
|
| 18 |
A |
1344 |
1283 |
1.50 |
|
|
|
| 19 |
A |
1311 |
1251 |
0.38 |
|
|
|
| 20 |
A |
1297 |
1238 |
17.28 |
|
|
|
| 21 |
A |
1276 |
1218 |
2.22 |
|
|
|
| 22 |
A |
1249 |
1192 |
5.75 |
|
|
|
| 23 |
A |
1216 |
1161 |
0.65 |
|
|
|
| 24 |
A |
1190 |
1136 |
5.92 |
|
|
|
| 25 |
A |
1136 |
1084 |
0.49 |
|
|
|
| 26 |
A |
1075 |
1026 |
1.16 |
|
|
|
| 27 |
A |
1005 |
960 |
0.50 |
|
|
|
| 28 |
A |
992 |
947 |
0.27 |
|
|
|
| 29 |
A |
976 |
932 |
3.11 |
|
|
|
| 30 |
A |
950 |
907 |
0.70 |
|
|
|
| 31 |
A |
911 |
870 |
2.13 |
|
|
|
| 32 |
A |
884 |
844 |
5.39 |
|
|
|
| 33 |
A |
835 |
797 |
5.97 |
|
|
|
| 34 |
A |
822 |
785 |
1.76 |
|
|
|
| 35 |
A |
753 |
719 |
1.88 |
|
|
|
| 36 |
A |
630 |
601 |
0.61 |
|
|
|
| 37 |
A |
465 |
444 |
0.52 |
|
|
|
| 38 |
A |
371 |
354 |
1.08 |
|
|
|
| 39 |
A |
249 |
237 |
3.83 |
|
|
|
| 40 |
A |
205 |
195 |
20.75 |
|
|
|
| 41 |
A |
166 |
158 |
0.59 |
|
|
|
| 42 |
A |
44 |
42 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31009.0 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 29595.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.232 |
|
|
|
| 2 |
H |
0.227 |
|
|
|
| 3 |
C |
-0.451 |
|
|
|
| 4 |
H |
0.231 |
|
|
|
| 5 |
H |
0.229 |
|
|
|
| 6 |
C |
-0.432 |
|
|
|
| 7 |
H |
0.242 |
|
|
|
| 8 |
H |
0.241 |
|
|
|
| 9 |
C |
-0.450 |
|
|
|
| 10 |
H |
0.227 |
|
|
|
| 11 |
H |
0.245 |
|
|
|
| 12 |
C |
-0.414 |
|
|
|
| 13 |
H |
0.259 |
|
|
|
| 14 |
C |
-0.437 |
|
|
|
| 15 |
H |
0.153 |
|
|
|
| 16 |
S |
-0.102 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.617 |
0.843 |
0.713 |
1.958 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.813 |
2.954 |
1.386 |
| y |
2.954 |
-45.151 |
1.170 |
| z |
1.386 |
1.170 |
-46.066 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.204 |
2.954 |
1.386 |
| y |
2.954 |
2.288 |
1.170 |
| z |
1.386 |
1.170 |
0.916 |
|
| Polar |
|---|
| 3z2-r2 | 1.832 |
|---|
| x2-y2 | -3.662 |
|---|
| xy | 2.954 |
|---|
| xz | 1.386 |
|---|
| yz | 1.170 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
229.544 |
| (<r2>)1/2 |
15.151 |