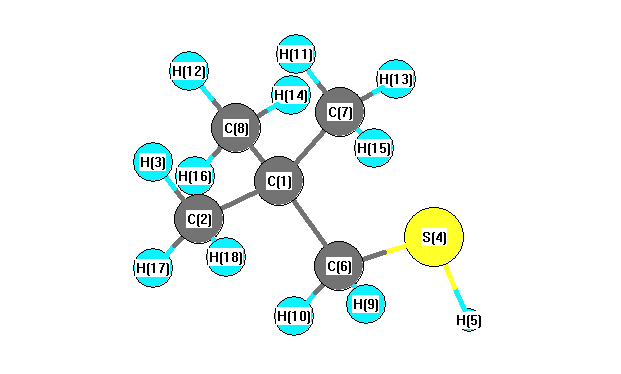
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3137 |
2994 |
54.99 |
|
|
|
| 2 |
A' |
3127 |
2984 |
79.34 |
|
|
|
| 3 |
A' |
3121 |
2979 |
0.02 |
|
|
|
| 4 |
A' |
3072 |
2932 |
30.60 |
|
|
|
| 5 |
A' |
3056 |
2917 |
13.42 |
|
|
|
| 6 |
A' |
3046 |
2907 |
26.06 |
|
|
|
| 7 |
A' |
2671 |
2549 |
11.52 |
|
|
|
| 8 |
A' |
1540 |
1469 |
16.67 |
|
|
|
| 9 |
A' |
1529 |
1459 |
11.67 |
|
|
|
| 10 |
A' |
1505 |
1436 |
0.29 |
|
|
|
| 11 |
A' |
1494 |
1426 |
4.53 |
|
|
|
| 12 |
A' |
1448 |
1382 |
10.77 |
|
|
|
| 13 |
A' |
1418 |
1353 |
11.75 |
|
|
|
| 14 |
A' |
1337 |
1276 |
18.97 |
|
|
|
| 15 |
A' |
1293 |
1234 |
11.55 |
|
|
|
| 16 |
A' |
1239 |
1182 |
16.59 |
|
|
|
| 17 |
A' |
1063 |
1014 |
0.17 |
|
|
|
| 18 |
A' |
977 |
932 |
1.02 |
|
|
|
| 19 |
A' |
952 |
908 |
1.15 |
|
|
|
| 20 |
A' |
892 |
851 |
6.74 |
|
|
|
| 21 |
A' |
788 |
752 |
3.53 |
|
|
|
| 22 |
A' |
709 |
677 |
0.22 |
|
|
|
| 23 |
A' |
457 |
436 |
1.71 |
|
|
|
| 24 |
A' |
394 |
376 |
0.28 |
|
|
|
| 25 |
A' |
321 |
306 |
0.31 |
|
|
|
| 26 |
A' |
268 |
256 |
0.01 |
|
|
|
| 27 |
A' |
201 |
192 |
1.42 |
|
|
|
| 28 |
A" |
3133 |
2990 |
14.95 |
|
|
|
| 29 |
A" |
3128 |
2985 |
39.04 |
|
|
|
| 30 |
A" |
3121 |
2979 |
7.89 |
|
|
|
| 31 |
A" |
3118 |
2976 |
9.58 |
|
|
|
| 32 |
A" |
3051 |
2912 |
35.64 |
|
|
|
| 33 |
A" |
1528 |
1459 |
8.54 |
|
|
|
| 34 |
A" |
1503 |
1435 |
0.00 |
|
|
|
| 35 |
A" |
1497 |
1429 |
0.18 |
|
|
|
| 36 |
A" |
1422 |
1357 |
15.51 |
|
|
|
| 37 |
A" |
1324 |
1263 |
0.26 |
|
|
|
| 38 |
A" |
1169 |
1116 |
5.20 |
|
|
|
| 39 |
A" |
1062 |
1013 |
1.96 |
|
|
|
| 40 |
A" |
971 |
927 |
0.03 |
|
|
|
| 41 |
A" |
957 |
913 |
0.34 |
|
|
|
| 42 |
A" |
897 |
856 |
3.93 |
|
|
|
| 43 |
A" |
416 |
397 |
0.01 |
|
|
|
| 44 |
A" |
340 |
324 |
0.16 |
|
|
|
| 45 |
A" |
278 |
265 |
0.00 |
|
|
|
| 46 |
A" |
225 |
215 |
0.42 |
|
|
|
| 47 |
A" |
145 |
139 |
10.61 |
|
|
|
| 48 |
A" |
32 |
30 |
18.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35183.0 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 33578.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.203 |
|
|
|
| 2 |
C |
-0.616 |
|
|
|
| 3 |
H |
0.229 |
|
|
|
| 4 |
S |
-0.119 |
|
|
|
| 5 |
H |
0.154 |
|
|
|
| 6 |
C |
-0.562 |
|
|
|
| 7 |
C |
-0.616 |
|
|
|
| 8 |
C |
-0.616 |
|
|
|
| 9 |
H |
0.256 |
|
|
|
| 10 |
H |
0.256 |
|
|
|
| 11 |
H |
0.223 |
|
|
|
| 12 |
H |
0.223 |
|
|
|
| 13 |
H |
0.248 |
|
|
|
| 14 |
H |
0.248 |
|
|
|
| 15 |
H |
0.223 |
|
|
|
| 16 |
H |
0.223 |
|
|
|
| 17 |
H |
0.224 |
|
|
|
| 18 |
H |
0.224 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.366 |
1.034 |
0.000 |
1.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.447 |
3.343 |
0.000 |
| y |
3.343 |
-48.007 |
0.000 |
| z |
0.000 |
0.000 |
-49.501 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.307 |
3.343 |
0.000 |
| y |
3.343 |
-0.033 |
0.000 |
| z |
0.000 |
0.000 |
-2.274 |
|
| Polar |
|---|
| 3z2-r2 | -4.547 |
|---|
| x2-y2 | 1.560 |
|---|
| xy | 3.343 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
242.732 |
| (<r2>)1/2 |
15.580 |