return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NH3 (methyl ammonium radical)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3V 2A1
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-96.252323
Energy at 298.15K-96.259453
HF Energy-96.131091
Nuclear repulsion energy47.404621
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3091 3091 80.06      
2 A1 2801 2801 58.53      
3 A1 1385 1385 82.98      
4 A1 1300 1300 183.53      
5 A1 919 919 37.17      
6 A2 291 291 0.00      
7 E 3190 3190 9.16      
7 E 3190 3190 9.16      
8 E 2381 2381 3944.91      
8 E 2381 2381 3945.79      
9 E 1509 1509 6.22      
9 E 1509 1509 6.23      
10 E 1417 1417 1.63      
10 E 1417 1417 1.62      
11 E 1230 1230 79.88      
11 E 1230 1230 79.94      
12 E 879 879 101.15      
12 E 879 879 101.17      

Unscaled Zero Point Vibrational Energy (zpe) 15500.1 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15500.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
2.70876 0.67089 0.67089

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C3v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.798
N2 0.000 0.000 0.706
H3 0.000 -1.035 -1.131
H4 -0.896 0.517 -1.131
H5 0.896 0.517 -1.131
H6 0.000 0.994 1.079
H7 -0.861 -0.497 1.079
H8 0.861 -0.497 1.079

Atom - Atom Distances (Å)
  C1 N2 H3 H4 H5 H6 H7 H8
C11.50441.08701.08701.08702.12392.12392.1239
N21.50442.10862.10862.10861.06131.06131.0613
H31.08702.10861.79241.79242.99982.43182.4318
H41.08702.10861.79241.79242.43182.43182.9998
H51.08702.10861.79241.79242.43182.99982.4318
H62.12391.06132.99982.43182.43181.72121.7212
H72.12391.06132.43182.43182.99981.72121.7212
H82.12391.06132.43182.99982.43181.72121.7212

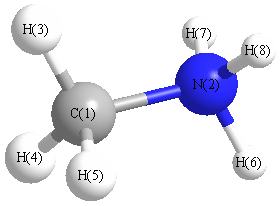
picture of methyl ammonium radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 110.558 C1 N2 H7 110.558
C1 N2 H8 110.558 N2 C1 H3 107.827
N2 C1 H4 107.827 N2 C1 H5 107.827
H3 C1 H4 111.064 H3 C1 H5 111.064
H4 C1 H5 111.064 H6 N2 H7 108.363
H6 N2 H8 108.363 H7 N2 H8 108.363
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability