return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Furan, 2,5-dihydro-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-230.650595
Energy at 298.15K-230.657612
HF Energy-229.832485
Nuclear repulsion energy174.114265
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3264 3091 7.19      
2 A' 3096 2932 79.33      
3 A' 3049 2887 24.46      
4 A' 1678 1589 0.29      
5 A' 1568 1485 0.94      
6 A' 1422 1347 3.63      
7 A' 1204 1140 2.04      
8 A' 1128 1069 1.86      
9 A' 1045 990 19.96      
10 A' 1017 963 10.01      
11 A' 937 887 19.11      
12 A' 759 718 6.52      
13 A' 669 634 42.99      
14 A' 36 34 8.38      
15 A" 3240 3069 2.39      
16 A" 3094 2930 11.19      
17 A" 3045 2884 117.82      
18 A" 1556 1474 0.82      
19 A" 1391 1317 1.59      
20 A" 1346 1275 0.71      
21 A" 1250 1184 0.48      
22 A" 1154 1093 102.35      
23 A" 1078 1021 0.23      
24 A" 945 895 0.00      
25 A" 916 867 0.02      
26 A" 812 769 1.72      
27 A" 370 350 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 20533.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19447.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.26832 0.26311 0.13995

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.065 -1.187 0.000
C2 -0.013 -0.372 1.168
C3 -0.013 1.044 0.668
C4 -0.013 1.044 -0.668
C5 -0.013 -0.372 -1.168
H6 0.845 -0.590 1.815
H7 -0.929 -0.607 1.729
H8 -0.929 -0.607 -1.729
H9 0.845 -0.590 -1.815
H10 -0.023 1.914 1.316
H11 -0.023 1.914 -1.316

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.42642.33022.33021.42642.06372.07652.07652.06373.36933.3693
C21.42641.50182.31872.33621.09661.09903.04713.11162.29043.3754
C32.33021.50181.33592.31872.17322.16553.05093.09381.08432.1658
C42.33022.31871.33591.50183.09383.05092.16552.17322.16581.0843
C51.42642.33622.31871.50183.11163.04711.09901.09663.37542.2904
H62.06371.09662.17323.09383.11161.77613.96273.62972.69664.1015
H72.07651.09902.16553.05093.04711.77613.45723.96272.71004.0547
H82.07653.04713.05092.16551.09903.96273.45721.77614.05472.7100
H92.06373.11163.09382.17321.09663.62973.96271.77614.10152.6966
H103.36932.29041.08432.16583.37542.69662.71004.05474.10152.6312
H113.36933.37542.16581.08432.29044.10154.05472.71002.69662.6312

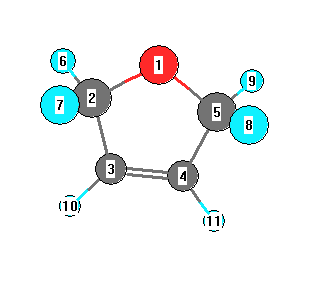
picture of Furan, 2,5-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 105.426 O1 C2 H6 109.060
O1 C2 H7 109.940 O1 C5 C4 105.426
O1 C5 H8 109.940 O1 C5 H9 109.060
C2 O1 C5 109.950 C2 C3 C4 109.453
C2 C3 H10 123.867 C3 C2 H6 112.581
C3 C2 H7 111.805 C3 C4 C5 109.453
C3 C4 H11 126.677 C4 C3 H10 126.677
C4 C5 H8 111.805 C4 C5 H9 112.581
C5 C4 H11 123.867 H6 C2 H7 107.984
H8 C5 H9 107.984
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability