Jump to
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -245.848878 |
| Energy at 298.15K | -245.854683 |
| HF Energy | -245.848878 |
| Nuclear repulsion energy | 158.781836 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3308 |
2992 |
35.14 |
|
|
|
| 2 |
A' |
3191 |
2886 |
40.57 |
|
|
|
| 3 |
A' |
3164 |
2862 |
50.58 |
|
|
|
| 4 |
A' |
2612 |
2362 |
2.34 |
|
|
|
| 5 |
A' |
1658 |
1500 |
2.97 |
|
|
|
| 6 |
A' |
1641 |
1484 |
7.68 |
|
|
|
| 7 |
A' |
1623 |
1468 |
0.09 |
|
|
|
| 8 |
A' |
1563 |
1414 |
69.16 |
|
|
|
| 9 |
A' |
1368 |
1237 |
94.49 |
|
|
|
| 10 |
A' |
1280 |
1158 |
159.49 |
|
|
|
| 11 |
A' |
1102 |
997 |
46.90 |
|
|
|
| 12 |
A' |
1010 |
913 |
11.63 |
|
|
|
| 13 |
A' |
600 |
542 |
3.03 |
|
|
|
| 14 |
A' |
408 |
369 |
3.00 |
|
|
|
| 15 |
A' |
200 |
181 |
3.01 |
|
|
|
| 16 |
A" |
3222 |
2914 |
37.74 |
|
|
|
| 17 |
A" |
3215 |
2908 |
76.37 |
|
|
|
| 18 |
A" |
1634 |
1477 |
8.01 |
|
|
|
| 19 |
A" |
1399 |
1265 |
4.72 |
|
|
|
| 20 |
A" |
1298 |
1174 |
6.97 |
|
|
|
| 21 |
A" |
1141 |
1032 |
4.17 |
|
|
|
| 22 |
A" |
422 |
382 |
3.27 |
|
|
|
| 23 |
A" |
231 |
209 |
6.01 |
|
|
|
| 24 |
A" |
91 |
82 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18690.5 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 16903.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.115 |
-0.374 |
0.000 |
| O2 |
-0.733 |
-0.578 |
0.000 |
| C3 |
0.000 |
0.594 |
0.000 |
| C4 |
1.436 |
0.264 |
0.000 |
| N5 |
2.543 |
0.047 |
0.000 |
| H6 |
-2.579 |
-1.349 |
0.000 |
| H7 |
-2.433 |
0.171 |
0.885 |
| H8 |
-2.433 |
0.171 |
-0.885 |
| H9 |
-0.207 |
1.198 |
-0.879 |
| H10 |
-0.207 |
1.198 |
0.879 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3973 | 2.3264 | 3.6083 | 4.6774 | 1.0786 | 1.0871 | 1.0871 | 2.6240 | 2.6240 |
O2 | 1.3973 | | 1.3817 | 2.3265 | 3.3351 | 2.0001 | 2.0579 | 2.0579 | 2.0498 | 2.0498 | C3 | 2.3264 | 1.3817 | | 1.4732 | 2.6009 | 3.2281 | 2.6233 | 2.6233 | 1.0866 | 1.0866 | C4 | 3.6083 | 2.3265 | 1.4732 | | 1.1281 | 4.3263 | 3.9701 | 3.9701 | 2.0844 | 2.0844 | N5 | 4.6774 | 3.3351 | 2.6009 | 1.1281 | | 5.3084 | 5.0558 | 5.0558 | 3.1081 | 3.1081 | H6 | 1.0786 | 2.0001 | 3.2281 | 4.3263 | 5.3084 | | 1.7647 | 1.7647 | 3.5888 | 3.5888 | H7 | 1.0871 | 2.0579 | 2.6233 | 3.9701 | 5.0558 | 1.7647 | | 1.7695 | 3.0198 | 2.4512 | H8 | 1.0871 | 2.0579 | 2.6233 | 3.9701 | 5.0558 | 1.7647 | 1.7695 | | 2.4512 | 3.0198 | H9 | 2.6240 | 2.0498 | 1.0866 | 2.0844 | 3.1081 | 3.5888 | 3.0198 | 2.4512 | | 1.7579 | H10 | 2.6240 | 2.0498 | 1.0866 | 2.0844 | 3.1081 | 3.5888 | 2.4512 | 3.0198 | 1.7579 | |
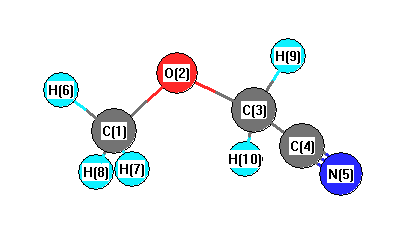 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.674 |
|
O2 |
C1 |
H6 |
107.063 |
| O2 |
C1 |
H7 |
111.242 |
|
O2 |
C1 |
H8 |
111.242 |
| O2 |
C3 |
C4 |
109.118 |
|
O2 |
C3 |
H9 |
111.729 |
| O2 |
C3 |
H10 |
111.729 |
|
C3 |
C4 |
N5 |
178.155 |
| C4 |
C3 |
H9 |
108.081 |
|
C4 |
C3 |
H10 |
108.081 |
| H6 |
C1 |
H7 |
109.141 |
|
H6 |
C1 |
H8 |
109.141 |
| H7 |
C1 |
H8 |
108.958 |
|
H9 |
C3 |
H10 |
107.972 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.381 |
|
|
|
| 2 |
O |
-0.423 |
|
|
|
| 3 |
C |
-0.091 |
|
|
|
| 4 |
C |
0.132 |
|
|
|
| 5 |
N |
-0.332 |
|
|
|
| 6 |
H |
0.232 |
|
|
|
| 7 |
H |
0.198 |
|
|
|
| 8 |
H |
0.198 |
|
|
|
| 9 |
H |
0.234 |
|
|
|
| 10 |
H |
0.234 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.709 |
1.952 |
0.000 |
5.097 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.802 |
0.783 |
0.000 |
| y |
0.783 |
-29.333 |
0.000 |
| z |
0.000 |
0.000 |
-29.226 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.522 |
0.783 |
0.000 |
| y |
0.783 |
3.681 |
0.000 |
| z |
0.000 |
0.000 |
3.841 |
|
| Polar |
|---|
| 3z2-r2 | 7.683 |
|---|
| x2-y2 | -7.469 |
|---|
| xy | 0.783 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.360 |
-0.034 |
0.000 |
| y |
-0.034 |
4.529 |
0.000 |
| z |
0.000 |
0.000 |
4.280 |
<r2> (average value of r
2) Å
2
| <r2> |
141.133 |
| (<r2>)1/2 |
11.880 |
Jump to
S1C1
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -245.850659 |
| Energy at 298.15K | -245.856570 |
| HF Energy | -245.850659 |
| Nuclear repulsion energy | 161.273535 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3309 |
2992 |
37.28 |
|
|
|
| 2 |
A |
3297 |
2982 |
13.33 |
|
|
|
| 3 |
A |
3235 |
2926 |
57.86 |
|
|
|
| 4 |
A |
3206 |
2899 |
39.45 |
|
|
|
| 5 |
A |
3174 |
2871 |
47.16 |
|
|
|
| 6 |
A |
2597 |
2349 |
1.02 |
|
|
|
| 7 |
A |
1648 |
1490 |
10.50 |
|
|
|
| 8 |
A |
1637 |
1481 |
4.39 |
|
|
|
| 9 |
A |
1634 |
1478 |
4.80 |
|
|
|
| 10 |
A |
1618 |
1463 |
0.61 |
|
|
|
| 11 |
A |
1534 |
1388 |
31.71 |
|
|
|
| 12 |
A |
1447 |
1309 |
15.35 |
|
|
|
| 13 |
A |
1360 |
1230 |
101.87 |
|
|
|
| 14 |
A |
1300 |
1175 |
15.92 |
|
|
|
| 15 |
A |
1282 |
1159 |
106.22 |
|
|
|
| 16 |
A |
1143 |
1033 |
10.25 |
|
|
|
| 17 |
A |
1036 |
937 |
40.95 |
|
|
|
| 18 |
A |
953 |
862 |
37.81 |
|
|
|
| 19 |
A |
653 |
590 |
1.19 |
|
|
|
| 20 |
A |
424 |
383 |
2.03 |
|
|
|
| 21 |
A |
410 |
371 |
3.09 |
|
|
|
| 22 |
A |
283 |
256 |
16.19 |
|
|
|
| 23 |
A |
185 |
167 |
3.45 |
|
|
|
| 24 |
A |
118 |
106 |
9.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18740.4 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 16948.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.610 |
-0.733 |
0.133 |
| O2 |
1.049 |
0.420 |
-0.433 |
| C3 |
-0.071 |
0.907 |
0.204 |
| C4 |
-1.260 |
0.033 |
0.033 |
| N5 |
-2.166 |
-0.631 |
-0.087 |
| H6 |
2.495 |
-0.963 |
-0.439 |
| H7 |
0.924 |
-1.572 |
0.082 |
| H8 |
1.888 |
-0.561 |
1.169 |
| H9 |
-0.300 |
1.871 |
-0.225 |
| H10 |
0.092 |
1.038 |
1.270 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4011 | 2.3492 | 2.9718 | 3.7831 | 1.0787 | 1.0850 | 1.0869 | 3.2482 | 2.5950 |
O2 | 1.4011 | | 1.3782 | 2.3880 | 3.4004 | 2.0009 | 2.0617 | 2.0574 | 1.9918 | 2.0497 | C3 | 2.3492 | 1.3782 | | 1.4857 | 2.6145 | 3.2399 | 2.6739 | 2.6311 | 1.0796 | 1.0862 | C4 | 2.9718 | 2.3880 | 1.4857 | | 1.1288 | 3.9134 | 2.7106 | 3.3991 | 2.0897 | 2.0906 | N5 | 3.7831 | 3.4004 | 2.6145 | 1.1288 | | 4.6857 | 3.2343 | 4.2443 | 3.1236 | 3.1181 | H6 | 1.0787 | 2.0009 | 3.2399 | 3.9134 | 4.6857 | | 1.7634 | 1.7654 | 3.9860 | 3.5642 | H7 | 1.0850 | 2.0617 | 2.6739 | 2.7106 | 3.2343 | 1.7634 | | 1.7700 | 3.6665 | 2.9861 | H8 | 1.0869 | 2.0574 | 2.6311 | 3.3991 | 4.2443 | 1.7654 | 1.7700 | | 3.5555 | 2.4069 | H9 | 3.2482 | 1.9918 | 1.0796 | 2.0897 | 3.1236 | 3.9860 | 3.6665 | 3.5555 | | 1.7557 | H10 | 2.5950 | 2.0497 | 1.0862 | 2.0906 | 3.1181 | 3.5642 | 2.9861 | 2.4069 | 1.7557 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
115.389 |
|
O2 |
C1 |
H6 |
106.866 |
| O2 |
C1 |
H7 |
111.417 |
|
O2 |
C1 |
H8 |
110.937 |
| O2 |
C3 |
C4 |
112.928 |
|
O2 |
C3 |
H9 |
107.654 |
| O2 |
C3 |
H10 |
112.003 |
|
C3 |
C4 |
N5 |
179.430 |
| C4 |
C3 |
H9 |
108.049 |
|
C4 |
C3 |
H10 |
107.733 |
| H6 |
C1 |
H7 |
109.175 |
|
H6 |
C1 |
H8 |
109.214 |
| H7 |
C1 |
H8 |
109.173 |
|
H9 |
C3 |
H10 |
108.319 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.387 |
|
|
|
| 2 |
O |
-0.419 |
|
|
|
| 3 |
C |
-0.153 |
|
|
|
| 4 |
C |
0.145 |
|
|
|
| 5 |
N |
-0.335 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
| 7 |
H |
0.221 |
|
|
|
| 8 |
H |
0.195 |
|
|
|
| 9 |
H |
0.263 |
|
|
|
| 10 |
H |
0.243 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.788 |
1.315 |
1.575 |
3.462 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
4.732 |
1.997 |
0.000 |
5.136 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.849 |
-5.910 |
0.758 |
| y |
-5.910 |
-28.205 |
0.676 |
| z |
0.758 |
0.676 |
-29.895 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.799 |
-5.910 |
0.758 |
| y |
-5.910 |
5.167 |
0.676 |
| z |
0.758 |
0.676 |
2.632 |
|
| Polar |
|---|
| 3z2-r2 | 5.265 |
|---|
| x2-y2 | -8.644 |
|---|
| xy | -5.910 |
|---|
| xz | 0.758 |
|---|
| yz | 0.676 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.394 |
0.473 |
0.153 |
| y |
0.473 |
5.185 |
0.197 |
| z |
0.153 |
0.197 |
4.275 |
<r2> (average value of r
2) Å
2
| <r2> |
123.056 |
| (<r2>)1/2 |
11.093 |
