Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -246.697880 |
| Energy at 298.15K | -246.703473 |
| HF Energy | -245.842568 |
| Nuclear repulsion energy | 157.038939 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3213 |
3043 |
15.55 |
|
|
|
| 2 |
A' |
3046 |
2885 |
52.75 |
|
|
|
| 3 |
A' |
3039 |
2878 |
20.11 |
|
|
|
| 4 |
A' |
2217 |
2100 |
9.24 |
|
|
|
| 5 |
A' |
1560 |
1478 |
6.11 |
|
|
|
| 6 |
A' |
1542 |
1460 |
9.14 |
|
|
|
| 7 |
A' |
1518 |
1437 |
0.00 |
|
|
|
| 8 |
A' |
1447 |
1370 |
32.16 |
|
|
|
| 9 |
A' |
1254 |
1188 |
44.50 |
|
|
|
| 10 |
A' |
1186 |
1123 |
122.09 |
|
|
|
| 11 |
A' |
1021 |
967 |
36.02 |
|
|
|
| 12 |
A' |
969 |
918 |
6.16 |
|
|
|
| 13 |
A' |
548 |
519 |
1.06 |
|
|
|
| 14 |
A' |
387 |
367 |
1.74 |
|
|
|
| 15 |
A' |
180 |
171 |
2.01 |
|
|
|
| 16 |
A" |
3116 |
2951 |
45.49 |
|
|
|
| 17 |
A" |
3082 |
2919 |
31.42 |
|
|
|
| 18 |
A" |
1531 |
1450 |
10.89 |
|
|
|
| 19 |
A" |
1296 |
1228 |
3.57 |
|
|
|
| 20 |
A" |
1209 |
1145 |
3.50 |
|
|
|
| 21 |
A" |
1055 |
999 |
2.55 |
|
|
|
| 22 |
A" |
363 |
344 |
1.17 |
|
|
|
| 23 |
A" |
237 |
225 |
4.66 |
|
|
|
| 24 |
A" |
88 |
83 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17552.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16623.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.127 |
-0.326 |
0.000 |
| O2 |
-0.740 |
-0.604 |
0.000 |
| C3 |
0.000 |
0.590 |
0.000 |
| C4 |
1.428 |
0.258 |
0.000 |
| N5 |
2.575 |
0.015 |
0.000 |
| H6 |
-2.635 |
-1.287 |
0.000 |
| H7 |
-2.420 |
0.240 |
0.893 |
| H8 |
-2.420 |
0.240 |
-0.893 |
| H9 |
-0.219 |
1.200 |
-0.888 |
| H10 |
-0.219 |
1.200 |
0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4152 | 2.3160 | 3.6031 | 4.7147 | 1.0876 | 1.0967 | 1.0967 | 2.5995 | 2.5995 |
O2 | 1.4152 | | 1.4048 | 2.3331 | 3.3722 | 2.0148 | 2.0815 | 2.0815 | 2.0772 | 2.0772 | C3 | 2.3160 | 1.4048 | | 1.4663 | 2.6384 | 3.2356 | 2.6030 | 2.6030 | 1.0995 | 1.0995 | C4 | 3.6031 | 2.3331 | 1.4663 | | 1.1722 | 4.3473 | 3.9503 | 3.9503 | 2.0951 | 2.0951 | N5 | 4.7147 | 3.3722 | 2.6384 | 1.1722 | | 5.3707 | 5.0791 | 5.0791 | 3.1621 | 3.1621 | H6 | 1.0876 | 2.0148 | 3.2356 | 4.3473 | 5.3707 | | 1.7823 | 1.7823 | 3.5796 | 3.5796 | H7 | 1.0967 | 2.0815 | 2.6030 | 3.9503 | 5.0791 | 1.7823 | | 1.7859 | 2.9896 | 2.4010 | H8 | 1.0967 | 2.0815 | 2.6030 | 3.9503 | 5.0791 | 1.7823 | 1.7859 | | 2.4010 | 2.9896 | H9 | 2.5995 | 2.0772 | 1.0995 | 2.0951 | 3.1621 | 3.5796 | 2.9896 | 2.4010 | | 1.7766 | H10 | 2.5995 | 2.0772 | 1.0995 | 2.0951 | 3.1621 | 3.5796 | 2.4010 | 2.9896 | 1.7766 | |
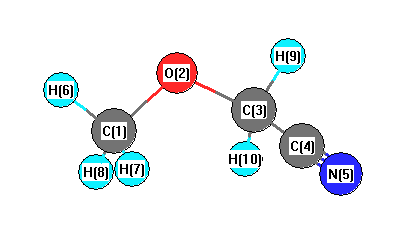 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.425 |
|
O2 |
C1 |
H6 |
106.490 |
| O2 |
C1 |
H7 |
111.286 |
|
O2 |
C1 |
H8 |
111.286 |
| O2 |
C3 |
C4 |
108.682 |
|
O2 |
C3 |
H9 |
111.503 |
| O2 |
C3 |
H10 |
111.503 |
|
C3 |
C4 |
N5 |
178.843 |
| C4 |
C3 |
H9 |
108.642 |
|
C4 |
C3 |
H10 |
108.642 |
| H6 |
C1 |
H7 |
109.357 |
|
H6 |
C1 |
H8 |
109.357 |
| H7 |
C1 |
H8 |
109.012 |
|
H9 |
C3 |
H10 |
107.793 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -246.700801 |
| Energy at 298.15K | -246.706527 |
| HF Energy | -245.844147 |
| Nuclear repulsion energy | 160.043335 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3213 |
3043 |
16.56 |
|
|
|
| 2 |
A |
3174 |
3006 |
4.35 |
|
|
|
| 3 |
A |
3130 |
2965 |
35.61 |
|
|
|
| 4 |
A |
3060 |
2898 |
31.34 |
|
|
|
| 5 |
A |
3052 |
2891 |
34.80 |
|
|
|
| 6 |
A |
2198 |
2082 |
9.56 |
|
|
|
| 7 |
A |
1552 |
1470 |
12.03 |
|
|
|
| 8 |
A |
1537 |
1455 |
9.38 |
|
|
|
| 9 |
A |
1527 |
1446 |
4.10 |
|
|
|
| 10 |
A |
1515 |
1435 |
2.00 |
|
|
|
| 11 |
A |
1417 |
1342 |
13.58 |
|
|
|
| 12 |
A |
1342 |
1271 |
6.48 |
|
|
|
| 13 |
A |
1246 |
1180 |
40.83 |
|
|
|
| 14 |
A |
1212 |
1148 |
8.69 |
|
|
|
| 15 |
A |
1184 |
1122 |
95.53 |
|
|
|
| 16 |
A |
1053 |
998 |
7.32 |
|
|
|
| 17 |
A |
971 |
920 |
33.56 |
|
|
|
| 18 |
A |
917 |
868 |
18.59 |
|
|
|
| 19 |
A |
605 |
573 |
1.91 |
|
|
|
| 20 |
A |
407 |
386 |
3.13 |
|
|
|
| 21 |
A |
355 |
336 |
1.00 |
|
|
|
| 22 |
A |
267 |
253 |
10.15 |
|
|
|
| 23 |
A |
183 |
174 |
4.16 |
|
|
|
| 24 |
A |
123 |
117 |
8.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17620.8 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16688.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.532 |
-0.787 |
0.138 |
| O2 |
1.089 |
0.431 |
-0.440 |
| C3 |
-0.045 |
0.937 |
0.210 |
| C4 |
-1.231 |
0.070 |
0.039 |
| N5 |
-2.158 |
-0.635 |
-0.100 |
| H6 |
2.446 |
-1.060 |
-0.385 |
| H7 |
0.790 |
-1.583 |
0.014 |
| H8 |
1.746 |
-0.658 |
1.206 |
| H9 |
-0.255 |
1.913 |
-0.229 |
| H10 |
0.130 |
1.070 |
1.287 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4188 | 2.3372 | 2.8942 | 3.7003 | 1.0877 | 1.0950 | 1.0967 | 3.2581 | 2.5941 |
O2 | 1.4188 | | 1.4017 | 2.3957 | 3.4339 | 2.0163 | 2.0854 | 2.0800 | 2.0115 | 2.0756 | C3 | 2.3372 | 1.4017 | | 1.4781 | 2.6511 | 3.2471 | 2.6614 | 2.5969 | 1.0908 | 1.0989 | C4 | 2.8942 | 2.3957 | 1.4781 | | 1.1731 | 3.8693 | 2.6110 | 3.2795 | 2.1020 | 2.0996 | N5 | 3.7003 | 3.4339 | 2.6511 | 1.1731 | | 4.6316 | 3.0985 | 4.1166 | 3.1826 | 3.1724 | H6 | 1.0877 | 2.0163 | 3.2471 | 3.8693 | 4.6316 | | 1.7812 | 1.7836 | 4.0190 | 3.5621 | H7 | 1.0950 | 2.0854 | 2.6614 | 2.6110 | 3.0985 | 1.7812 | | 1.7864 | 3.6564 | 3.0151 | H8 | 1.0967 | 2.0800 | 2.5969 | 3.2795 | 4.1166 | 1.7836 | 1.7864 | | 3.5598 | 2.3669 | H9 | 3.2581 | 2.0115 | 1.0908 | 2.1020 | 3.1826 | 4.0190 | 3.6564 | 3.5598 | | 1.7767 | H10 | 2.5941 | 2.0756 | 1.0989 | 2.0996 | 3.1724 | 3.5621 | 3.0151 | 2.3669 | 1.7767 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.921 |
|
O2 |
C1 |
H6 |
106.358 |
| O2 |
C1 |
H7 |
111.461 |
|
O2 |
C1 |
H8 |
110.911 |
| O2 |
C3 |
C4 |
112.561 |
|
O2 |
C3 |
H9 |
106.955 |
| O2 |
C3 |
H10 |
111.635 |
|
C3 |
C4 |
N5 |
178.886 |
| C4 |
C3 |
H9 |
108.890 |
|
C4 |
C3 |
H10 |
108.224 |
| H6 |
C1 |
H7 |
109.387 |
|
H6 |
C1 |
H8 |
109.478 |
| H7 |
C1 |
H8 |
109.190 |
|
H9 |
C3 |
H10 |
108.471 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
