Jump to
S1C2
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -247.025748 |
| Energy at 298.15K | -247.031364 |
| HF Energy | -247.025748 |
| Nuclear repulsion energy | 157.856966 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3170 |
3039 |
19.22 |
|
|
|
| 2 |
A' |
3008 |
2883 |
58.99 |
|
|
|
| 3 |
A' |
2998 |
2874 |
26.00 |
|
|
|
| 4 |
A' |
2402 |
2303 |
0.58 |
|
|
|
| 5 |
A' |
1531 |
1468 |
7.49 |
|
|
|
| 6 |
A' |
1508 |
1446 |
11.11 |
|
|
|
| 7 |
A' |
1491 |
1429 |
0.14 |
|
|
|
| 8 |
A' |
1430 |
1370 |
35.37 |
|
|
|
| 9 |
A' |
1246 |
1194 |
68.21 |
|
|
|
| 10 |
A' |
1192 |
1143 |
118.26 |
|
|
|
| 11 |
A' |
1029 |
986 |
37.39 |
|
|
|
| 12 |
A' |
969 |
929 |
6.25 |
|
|
|
| 13 |
A' |
560 |
537 |
1.57 |
|
|
|
| 14 |
A' |
383 |
367 |
1.86 |
|
|
|
| 15 |
A' |
183 |
175 |
2.28 |
|
|
|
| 16 |
A" |
3062 |
2935 |
55.36 |
|
|
|
| 17 |
A" |
3028 |
2902 |
36.82 |
|
|
|
| 18 |
A" |
1503 |
1441 |
11.66 |
|
|
|
| 19 |
A" |
1282 |
1229 |
4.12 |
|
|
|
| 20 |
A" |
1195 |
1145 |
3.04 |
|
|
|
| 21 |
A" |
1038 |
995 |
1.88 |
|
|
|
| 22 |
A" |
385 |
369 |
1.46 |
|
|
|
| 23 |
A" |
234 |
224 |
4.61 |
|
|
|
| 24 |
A" |
90 |
87 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17458.5 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 16734.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.117 |
-0.359 |
0.000 |
| O2 |
-0.733 |
-0.608 |
0.000 |
| C3 |
0.000 |
0.580 |
0.000 |
| C4 |
1.427 |
0.273 |
0.000 |
| N5 |
2.559 |
0.063 |
0.000 |
| H6 |
-2.611 |
-1.330 |
0.000 |
| H7 |
-2.426 |
0.201 |
0.893 |
| H8 |
-2.426 |
0.201 |
-0.893 |
| H9 |
-0.222 |
1.195 |
-0.886 |
| H10 |
-0.222 |
1.195 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4056 | 2.3156 | 3.5999 | 4.6944 | 1.0890 | 1.0989 | 1.0989 | 2.6063 | 2.6063 |
O2 | 1.4056 | | 1.3956 | 2.3331 | 3.3597 | 2.0116 | 2.0779 | 2.0779 | 2.0729 | 2.0729 | C3 | 2.3156 | 1.3956 | | 1.4597 | 2.6102 | 3.2346 | 2.6129 | 2.6129 | 1.1015 | 1.1015 | C4 | 3.5999 | 2.3331 | 1.4597 | | 1.1508 | 4.3446 | 3.9561 | 3.9561 | 2.0866 | 2.0866 | N5 | 4.6944 | 3.3597 | 2.6102 | 1.1508 | | 5.3540 | 5.0660 | 5.0660 | 3.1298 | 3.1298 | H6 | 1.0890 | 2.0116 | 3.2346 | 4.3446 | 5.3540 | | 1.7820 | 1.7820 | 3.5872 | 3.5872 | H7 | 1.0989 | 2.0779 | 2.6129 | 3.9561 | 5.0660 | 1.7820 | | 1.7864 | 3.0024 | 2.4181 | H8 | 1.0989 | 2.0779 | 2.6129 | 3.9561 | 5.0660 | 1.7820 | 1.7864 | | 2.4181 | 3.0024 | H9 | 2.6063 | 2.0729 | 1.1015 | 2.0866 | 3.1298 | 3.5872 | 3.0024 | 2.4181 | | 1.7729 | H10 | 2.6063 | 2.0729 | 1.1015 | 2.0866 | 3.1298 | 3.5872 | 2.4181 | 3.0024 | 1.7729 | |
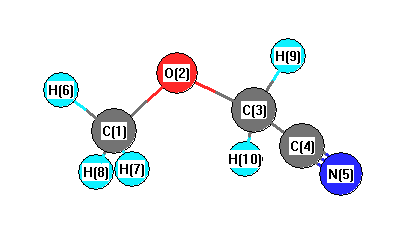 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.516 |
|
O2 |
C1 |
H6 |
106.806 |
| O2 |
C1 |
H7 |
111.546 |
|
O2 |
C1 |
H8 |
111.546 |
| O2 |
C3 |
C4 |
109.574 |
|
O2 |
C3 |
H9 |
111.683 |
| O2 |
C3 |
H10 |
111.683 |
|
C3 |
C4 |
N5 |
178.380 |
| C4 |
C3 |
H9 |
108.297 |
|
C4 |
C3 |
H10 |
108.297 |
| H6 |
C1 |
H7 |
109.069 |
|
H6 |
C1 |
H8 |
109.069 |
| H7 |
C1 |
H8 |
108.740 |
|
H9 |
C3 |
H10 |
107.177 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.478 |
|
|
|
| 2 |
O |
-0.312 |
|
|
|
| 3 |
C |
-0.231 |
|
|
|
| 4 |
C |
0.128 |
|
|
|
| 5 |
N |
-0.265 |
|
|
|
| 6 |
H |
0.245 |
|
|
|
| 7 |
H |
0.213 |
|
|
|
| 8 |
H |
0.213 |
|
|
|
| 9 |
H |
0.244 |
|
|
|
| 10 |
H |
0.244 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.364 |
1.751 |
0.000 |
4.702 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.055 |
0.696 |
0.000 |
| y |
0.696 |
-29.219 |
0.000 |
| z |
0.000 |
0.000 |
-29.183 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.854 |
0.696 |
0.000 |
| y |
0.696 |
3.400 |
0.000 |
| z |
0.000 |
0.000 |
3.453 |
|
| Polar |
|---|
| 3z2-r2 | 6.907 |
|---|
| x2-y2 | -6.836 |
|---|
| xy | 0.696 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.124 |
0.000 |
0.000 |
| y |
0.000 |
4.844 |
0.000 |
| z |
0.000 |
0.000 |
4.555 |
<r2> (average value of r
2) Å
2
| <r2> |
141.678 |
| (<r2>)1/2 |
11.903 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -247.028257 |
| Energy at 298.15K | -247.033959 |
| HF Energy | -247.028257 |
| Nuclear repulsion energy | 160.646875 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3040 |
20.46 |
|
|
|
| 2 |
A |
3132 |
3002 |
5.40 |
|
|
|
| 3 |
A |
3081 |
2953 |
41.77 |
|
|
|
| 4 |
A |
3019 |
2893 |
44.69 |
|
|
|
| 5 |
A |
3014 |
2889 |
33.35 |
|
|
|
| 6 |
A |
2384 |
2285 |
0.18 |
|
|
|
| 7 |
A |
1524 |
1460 |
11.73 |
|
|
|
| 8 |
A |
1506 |
1443 |
9.27 |
|
|
|
| 9 |
A |
1496 |
1434 |
4.17 |
|
|
|
| 10 |
A |
1480 |
1419 |
5.76 |
|
|
|
| 11 |
A |
1403 |
1345 |
13.88 |
|
|
|
| 12 |
A |
1330 |
1275 |
9.72 |
|
|
|
| 13 |
A |
1239 |
1187 |
62.53 |
|
|
|
| 14 |
A |
1196 |
1146 |
19.38 |
|
|
|
| 15 |
A |
1185 |
1135 |
74.79 |
|
|
|
| 16 |
A |
1041 |
998 |
8.53 |
|
|
|
| 17 |
A |
975 |
934 |
33.03 |
|
|
|
| 18 |
A |
916 |
878 |
18.59 |
|
|
|
| 19 |
A |
612 |
587 |
1.82 |
|
|
|
| 20 |
A |
395 |
378 |
2.41 |
|
|
|
| 21 |
A |
372 |
356 |
1.37 |
|
|
|
| 22 |
A |
259 |
248 |
11.09 |
|
|
|
| 23 |
A |
179 |
172 |
3.43 |
|
|
|
| 24 |
A |
119 |
114 |
8.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17512.3 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 16785.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.563 |
-0.762 |
0.135 |
| O2 |
1.074 |
0.432 |
-0.434 |
| C3 |
-0.063 |
0.919 |
0.207 |
| C4 |
-1.241 |
0.054 |
0.036 |
| N5 |
-2.154 |
-0.636 |
-0.096 |
| H6 |
2.467 |
-1.022 |
-0.414 |
| H7 |
0.838 |
-1.580 |
0.042 |
| H8 |
1.811 |
-0.624 |
1.196 |
| H9 |
-0.283 |
1.897 |
-0.228 |
| H10 |
0.103 |
1.058 |
1.286 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4102 | 2.3398 | 2.9221 | 3.7260 | 1.0890 | 1.0968 | 1.0985 | 3.2571 | 2.6019 |
O2 | 1.4102 | | 1.3929 | 2.3926 | 3.4168 | 2.0134 | 2.0813 | 2.0771 | 2.0074 | 2.0722 | C3 | 2.3398 | 1.3929 | | 1.4721 | 2.6237 | 3.2483 | 2.6620 | 2.6209 | 1.0922 | 1.1008 | C4 | 2.9221 | 2.3926 | 1.4721 | | 1.1517 | 3.8872 | 2.6447 | 3.3347 | 2.0940 | 2.0924 | N5 | 3.7260 | 3.4168 | 2.6237 | 1.1517 | | 4.6478 | 3.1404 | 4.1700 | 3.1519 | 3.1422 | H6 | 1.0890 | 2.0134 | 3.2483 | 3.8872 | 4.6478 | | 1.7813 | 1.7834 | 4.0143 | 3.5782 | H7 | 1.0968 | 2.0813 | 2.6620 | 2.6447 | 3.1404 | 1.7813 | | 1.7870 | 3.6633 | 3.0081 | H8 | 1.0985 | 2.0771 | 2.6209 | 3.3347 | 4.1700 | 1.7834 | 1.7870 | | 3.5725 | 2.3986 | H9 | 3.2571 | 2.0074 | 1.0922 | 2.0940 | 3.1519 | 4.0143 | 3.6633 | 3.5725 | | 1.7730 | H10 | 2.6019 | 2.0722 | 1.1008 | 2.0924 | 3.1422 | 3.5782 | 3.0081 | 2.3986 | 1.7730 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.170 |
|
O2 |
C1 |
H6 |
106.634 |
| O2 |
C1 |
H7 |
111.624 |
|
O2 |
C1 |
H8 |
111.163 |
| O2 |
C3 |
C4 |
113.227 |
|
O2 |
C3 |
H9 |
107.136 |
| O2 |
C3 |
H10 |
111.862 |
|
C3 |
C4 |
N5 |
179.195 |
| C4 |
C3 |
H9 |
108.585 |
|
C4 |
C3 |
H10 |
107.960 |
| H6 |
C1 |
H7 |
109.161 |
|
H6 |
C1 |
H8 |
109.220 |
| H7 |
C1 |
H8 |
108.977 |
|
H9 |
C3 |
H10 |
107.902 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.486 |
|
|
|
| 2 |
O |
-0.304 |
|
|
|
| 3 |
C |
-0.283 |
|
|
|
| 4 |
C |
0.134 |
|
|
|
| 5 |
N |
-0.270 |
|
|
|
| 6 |
H |
0.240 |
|
|
|
| 7 |
H |
0.237 |
|
|
|
| 8 |
H |
0.210 |
|
|
|
| 9 |
H |
0.272 |
|
|
|
| 10 |
H |
0.252 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.542 |
1.238 |
1.404 |
3.157 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.175 |
-5.537 |
0.601 |
| y |
-5.537 |
-27.976 |
0.592 |
| z |
0.601 |
0.592 |
-29.722 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.326 |
-5.537 |
0.601 |
| y |
-5.537 |
4.972 |
0.592 |
| z |
0.601 |
0.592 |
2.354 |
|
| Polar |
|---|
| 3z2-r2 | 4.708 |
|---|
| x2-y2 | -8.199 |
|---|
| xy | -5.537 |
|---|
| xz | 0.601 |
|---|
| yz | 0.592 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.935 |
0.451 |
0.174 |
| y |
0.451 |
5.627 |
0.228 |
| z |
0.174 |
0.228 |
4.564 |
<r2> (average value of r
2) Å
2
| <r2> |
122.128 |
| (<r2>)1/2 |
11.051 |
