Jump to
S1C2
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -247.215907 |
| Energy at 298.15K | -247.221580 |
| HF Energy | -247.215907 |
| Nuclear repulsion energy | 157.766819 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3169 |
3169 |
13.32 |
|
|
|
| 2 |
A' |
3046 |
3046 |
28.31 |
|
|
|
| 3 |
A' |
3037 |
3037 |
36.95 |
|
|
|
| 4 |
A' |
2438 |
2438 |
0.44 |
|
|
|
| 5 |
A' |
1543 |
1543 |
7.42 |
|
|
|
| 6 |
A' |
1521 |
1521 |
9.61 |
|
|
|
| 7 |
A' |
1503 |
1503 |
0.16 |
|
|
|
| 8 |
A' |
1441 |
1441 |
35.64 |
|
|
|
| 9 |
A' |
1260 |
1260 |
72.32 |
|
|
|
| 10 |
A' |
1202 |
1202 |
111.09 |
|
|
|
| 11 |
A' |
1039 |
1039 |
38.05 |
|
|
|
| 12 |
A' |
971 |
971 |
7.66 |
|
|
|
| 13 |
A' |
561 |
561 |
1.58 |
|
|
|
| 14 |
A' |
395 |
395 |
2.07 |
|
|
|
| 15 |
A' |
186 |
186 |
2.20 |
|
|
|
| 16 |
A" |
3096 |
3096 |
41.12 |
|
|
|
| 17 |
A" |
3076 |
3076 |
29.01 |
|
|
|
| 18 |
A" |
1515 |
1515 |
11.33 |
|
|
|
| 19 |
A" |
1298 |
1298 |
3.77 |
|
|
|
| 20 |
A" |
1205 |
1205 |
3.23 |
|
|
|
| 21 |
A" |
1048 |
1048 |
2.24 |
|
|
|
| 22 |
A" |
392 |
392 |
1.65 |
|
|
|
| 23 |
A" |
240 |
240 |
4.96 |
|
|
|
| 24 |
A" |
98 |
98 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17640.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17640.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.121 |
-0.350 |
0.000 |
| O2 |
-0.734 |
-0.604 |
0.000 |
| C3 |
0.000 |
0.585 |
0.000 |
| C4 |
1.433 |
0.267 |
0.000 |
| N5 |
2.558 |
0.046 |
0.000 |
| H6 |
-2.620 |
-1.316 |
0.000 |
| H7 |
-2.421 |
0.213 |
0.893 |
| H8 |
-2.421 |
0.213 |
-0.893 |
| H9 |
-0.220 |
1.195 |
-0.887 |
| H10 |
-0.220 |
1.195 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4098 | 2.3180 | 3.6066 | 4.6957 | 1.0876 | 1.0968 | 1.0968 | 2.6053 | 2.6053 |
O2 | 1.4098 | | 1.3980 | 2.3353 | 3.3559 | 2.0154 | 2.0756 | 2.0756 | 2.0708 | 2.0708 | C3 | 2.3180 | 1.3980 | | 1.4676 | 2.6141 | 3.2372 | 2.6067 | 2.6067 | 1.0987 | 1.0987 | C4 | 3.6066 | 2.3353 | 1.4676 | | 1.1468 | 4.3505 | 3.9556 | 3.9556 | 2.0926 | 2.0926 | N5 | 4.6957 | 3.3559 | 2.6141 | 1.1468 | | 5.3540 | 5.0607 | 5.0607 | 3.1341 | 3.1341 | H6 | 1.0876 | 2.0154 | 3.2372 | 4.3505 | 5.3540 | | 1.7817 | 1.7817 | 3.5849 | 3.5849 | H7 | 1.0968 | 2.0756 | 2.6067 | 3.9556 | 5.0607 | 1.7817 | | 1.7851 | 2.9960 | 2.4099 | H8 | 1.0968 | 2.0756 | 2.6067 | 3.9556 | 5.0607 | 1.7817 | 1.7851 | | 2.4099 | 2.9960 | H9 | 2.6053 | 2.0708 | 1.0987 | 2.0926 | 3.1341 | 3.5849 | 2.9960 | 2.4099 | | 1.7748 | H10 | 2.6053 | 2.0708 | 1.0987 | 2.0926 | 3.1341 | 3.5849 | 2.4099 | 2.9960 | 1.7748 | |
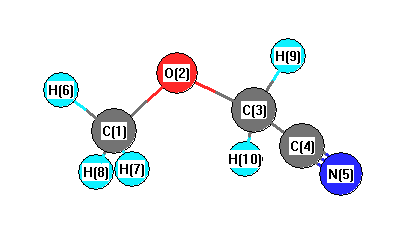 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.287 |
|
O2 |
C1 |
H6 |
106.899 |
| O2 |
C1 |
H7 |
111.179 |
|
O2 |
C1 |
H8 |
111.179 |
| O2 |
C3 |
C4 |
109.139 |
|
O2 |
C3 |
H9 |
111.518 |
| O2 |
C3 |
H10 |
111.518 |
|
C3 |
C4 |
N5 |
178.529 |
| C4 |
C3 |
H9 |
108.408 |
|
C4 |
C3 |
H10 |
108.408 |
| H6 |
C1 |
H7 |
109.303 |
|
H6 |
C1 |
H8 |
109.303 |
| H7 |
C1 |
H8 |
108.933 |
|
H9 |
C3 |
H10 |
107.749 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.474 |
|
|
|
| 2 |
O |
-0.337 |
|
|
|
| 3 |
C |
-0.240 |
|
|
|
| 4 |
C |
0.130 |
|
|
|
| 5 |
N |
-0.263 |
|
|
|
| 6 |
H |
0.248 |
|
|
|
| 7 |
H |
0.216 |
|
|
|
| 8 |
H |
0.216 |
|
|
|
| 9 |
H |
0.252 |
|
|
|
| 10 |
H |
0.252 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.399 |
1.854 |
0.000 |
4.774 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.980 |
0.727 |
0.000 |
| y |
0.727 |
-29.425 |
0.000 |
| z |
0.000 |
0.000 |
-29.331 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.602 |
0.727 |
0.000 |
| y |
0.727 |
3.231 |
0.000 |
| z |
0.000 |
0.000 |
3.371 |
|
| Polar |
|---|
| 3z2-r2 | 6.742 |
|---|
| x2-y2 | -6.556 |
|---|
| xy | 0.727 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.949 |
-0.011 |
0.000 |
| y |
-0.011 |
4.746 |
0.000 |
| z |
0.000 |
0.000 |
4.452 |
<r2> (average value of r
2) Å
2
| <r2> |
141.822 |
| (<r2>)1/2 |
11.909 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -247.219057 |
| Energy at 298.15K | -247.224811 |
| HF Energy | -247.219057 |
| Nuclear repulsion energy | 160.789903 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3185 |
3185 |
15.50 |
|
|
|
| 2 |
A |
3155 |
3155 |
3.35 |
|
|
|
| 3 |
A |
3108 |
3108 |
30.60 |
|
|
|
| 4 |
A |
3057 |
3057 |
25.95 |
|
|
|
| 5 |
A |
3039 |
3039 |
34.16 |
|
|
|
| 6 |
A |
2415 |
2415 |
0.14 |
|
|
|
| 7 |
A |
1536 |
1536 |
12.04 |
|
|
|
| 8 |
A |
1522 |
1522 |
8.58 |
|
|
|
| 9 |
A |
1511 |
1511 |
3.90 |
|
|
|
| 10 |
A |
1495 |
1495 |
3.35 |
|
|
|
| 11 |
A |
1408 |
1408 |
16.06 |
|
|
|
| 12 |
A |
1337 |
1337 |
9.37 |
|
|
|
| 13 |
A |
1253 |
1253 |
78.04 |
|
|
|
| 14 |
A |
1206 |
1206 |
41.43 |
|
|
|
| 15 |
A |
1196 |
1196 |
39.24 |
|
|
|
| 16 |
A |
1053 |
1053 |
8.86 |
|
|
|
| 17 |
A |
985 |
985 |
33.66 |
|
|
|
| 18 |
A |
915 |
915 |
22.96 |
|
|
|
| 19 |
A |
618 |
618 |
1.63 |
|
|
|
| 20 |
A |
409 |
409 |
3.15 |
|
|
|
| 21 |
A |
376 |
376 |
1.12 |
|
|
|
| 22 |
A |
265 |
265 |
11.15 |
|
|
|
| 23 |
A |
185 |
185 |
4.05 |
|
|
|
| 24 |
A |
122 |
122 |
8.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17674.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17674.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.538 |
-0.779 |
0.135 |
| O2 |
1.081 |
0.434 |
-0.430 |
| C3 |
-0.055 |
0.935 |
0.206 |
| C4 |
-1.235 |
0.058 |
0.033 |
| N5 |
-2.139 |
-0.638 |
-0.095 |
| H6 |
2.442 |
-1.053 |
-0.403 |
| H7 |
0.792 |
-1.573 |
0.022 |
| H8 |
1.769 |
-0.651 |
1.199 |
| H9 |
-0.275 |
1.907 |
-0.238 |
| H10 |
0.108 |
1.072 |
1.283 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4140 | 2.3408 | 2.8985 | 3.6863 | 1.0874 | 1.0952 | 1.0966 | 3.2613 | 2.6055 |
O2 | 1.4140 | | 1.3941 | 2.3918 | 3.4098 | 2.0163 | 2.0780 | 2.0749 | 2.0105 | 2.0701 | C3 | 2.3408 | 1.3941 | | 1.4810 | 2.6285 | 3.2492 | 2.6543 | 2.6138 | 1.0900 | 1.0982 | C4 | 2.8985 | 2.3918 | 1.4810 | | 1.1477 | 3.8665 | 2.6028 | 3.3003 | 2.1007 | 2.0966 | N5 | 3.6863 | 3.4098 | 2.6285 | 1.1477 | | 4.6100 | 3.0790 | 4.1169 | 3.1575 | 3.1422 | H6 | 1.0874 | 2.0163 | 3.2492 | 3.8665 | 4.6100 | | 1.7815 | 1.7833 | 4.0209 | 3.5785 | H7 | 1.0952 | 2.0780 | 2.6543 | 2.6028 | 3.0790 | 1.7815 | | 1.7862 | 3.6493 | 3.0098 | H8 | 1.0966 | 2.0749 | 2.6138 | 3.3003 | 4.1169 | 1.7833 | 1.7862 | | 3.5757 | 2.3954 | H9 | 3.2613 | 2.0105 | 1.0900 | 2.1007 | 3.1575 | 4.0209 | 3.6493 | 3.5757 | | 1.7760 | H10 | 2.6055 | 2.0701 | 1.0982 | 2.0966 | 3.1422 | 3.5785 | 3.0098 | 2.3954 | 1.7760 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.936 |
|
O2 |
C1 |
H6 |
106.702 |
| O2 |
C1 |
H7 |
111.187 |
|
O2 |
C1 |
H8 |
110.840 |
| O2 |
C3 |
C4 |
112.553 |
|
O2 |
C3 |
H9 |
107.430 |
| O2 |
C3 |
H10 |
111.774 |
|
C3 |
C4 |
N5 |
178.957 |
| C4 |
C3 |
H9 |
108.631 |
|
C4 |
C3 |
H10 |
107.837 |
| H6 |
C1 |
H7 |
109.417 |
|
H6 |
C1 |
H8 |
109.480 |
| H7 |
C1 |
H8 |
109.170 |
|
H9 |
C3 |
H10 |
108.509 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.482 |
|
|
|
| 2 |
O |
-0.330 |
|
|
|
| 3 |
C |
-0.285 |
|
|
|
| 4 |
C |
0.128 |
|
|
|
| 5 |
N |
-0.267 |
|
|
|
| 6 |
H |
0.242 |
|
|
|
| 7 |
H |
0.240 |
|
|
|
| 8 |
H |
0.214 |
|
|
|
| 9 |
H |
0.279 |
|
|
|
| 10 |
H |
0.260 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.468 |
1.213 |
1.447 |
3.107 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.319 |
-5.601 |
0.691 |
| y |
-5.601 |
-28.014 |
0.631 |
| z |
0.691 |
0.631 |
-29.905 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.359 |
-5.601 |
0.691 |
| y |
-5.601 |
5.098 |
0.631 |
| z |
0.691 |
0.631 |
2.262 |
|
| Polar |
|---|
| 3z2-r2 | 4.523 |
|---|
| x2-y2 | -8.305 |
|---|
| xy | -5.601 |
|---|
| xz | 0.691 |
|---|
| yz | 0.631 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.764 |
0.438 |
0.166 |
| y |
0.438 |
5.575 |
0.217 |
| z |
0.166 |
0.217 |
4.482 |
<r2> (average value of r
2) Å
2
| <r2> |
121.431 |
| (<r2>)1/2 |
11.020 |
