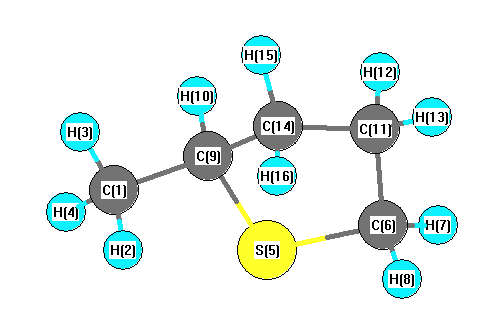
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3001 |
25.32 |
|
|
|
| 2 |
A |
3141 |
2998 |
29.64 |
|
|
|
| 3 |
A |
3124 |
2982 |
33.51 |
|
|
|
| 4 |
A |
3115 |
2973 |
41.50 |
|
|
|
| 5 |
A |
3104 |
2962 |
31.29 |
|
|
|
| 6 |
A |
3084 |
2944 |
31.27 |
|
|
|
| 7 |
A |
3069 |
2929 |
11.88 |
|
|
|
| 8 |
A |
3064 |
2924 |
27.40 |
|
|
|
| 9 |
A |
3054 |
2915 |
52.69 |
|
|
|
| 10 |
A |
3051 |
2912 |
7.91 |
|
|
|
| 11 |
A |
1522 |
1453 |
2.71 |
|
|
|
| 12 |
A |
1519 |
1450 |
6.68 |
|
|
|
| 13 |
A |
1508 |
1439 |
7.53 |
|
|
|
| 14 |
A |
1501 |
1433 |
6.28 |
|
|
|
| 15 |
A |
1496 |
1428 |
4.59 |
|
|
|
| 16 |
A |
1428 |
1362 |
6.13 |
|
|
|
| 17 |
A |
1396 |
1332 |
1.31 |
|
|
|
| 18 |
A |
1372 |
1310 |
0.61 |
|
|
|
| 19 |
A |
1333 |
1272 |
0.43 |
|
|
|
| 20 |
A |
1321 |
1260 |
5.35 |
|
|
|
| 21 |
A |
1305 |
1245 |
27.43 |
|
|
|
| 22 |
A |
1255 |
1198 |
5.41 |
|
|
|
| 23 |
A |
1218 |
1162 |
7.80 |
|
|
|
| 24 |
A |
1175 |
1122 |
0.82 |
|
|
|
| 25 |
A |
1125 |
1074 |
0.41 |
|
|
|
| 26 |
A |
1093 |
1043 |
3.88 |
|
|
|
| 27 |
A |
1053 |
1005 |
0.68 |
|
|
|
| 28 |
A |
1051 |
1003 |
4.96 |
|
|
|
| 29 |
A |
983 |
939 |
2.63 |
|
|
|
| 30 |
A |
967 |
923 |
2.14 |
|
|
|
| 31 |
A |
917 |
875 |
1.05 |
|
|
|
| 32 |
A |
876 |
836 |
3.04 |
|
|
|
| 33 |
A |
861 |
822 |
2.27 |
|
|
|
| 34 |
A |
702 |
670 |
1.95 |
|
|
|
| 35 |
A |
646 |
616 |
3.34 |
|
|
|
| 36 |
A |
531 |
507 |
0.88 |
|
|
|
| 37 |
A |
464 |
443 |
0.07 |
|
|
|
| 38 |
A |
415 |
396 |
0.87 |
|
|
|
| 39 |
A |
287 |
274 |
0.89 |
|
|
|
| 40 |
A |
259 |
248 |
0.10 |
|
|
|
| 41 |
A |
244 |
233 |
0.23 |
|
|
|
| 42 |
A |
93 |
89 |
1.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31432.0 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 29998.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.634 |
|
|
|
| 2 |
H |
0.235 |
|
|
|
| 3 |
H |
0.222 |
|
|
|
| 4 |
H |
0.237 |
|
|
|
| 5 |
S |
0.114 |
|
|
|
| 6 |
C |
-0.577 |
|
|
|
| 7 |
H |
0.255 |
|
|
|
| 8 |
H |
0.253 |
|
|
|
| 9 |
C |
-0.405 |
|
|
|
| 10 |
H |
0.252 |
|
|
|
| 11 |
C |
-0.456 |
|
|
|
| 12 |
H |
0.234 |
|
|
|
| 13 |
H |
0.232 |
|
|
|
| 14 |
C |
-0.425 |
|
|
|
| 15 |
H |
0.231 |
|
|
|
| 16 |
H |
0.231 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.390 |
2.143 |
0.006 |
2.178 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.613 |
-0.328 |
1.032 |
| y |
-0.328 |
-48.603 |
-0.359 |
| z |
1.032 |
-0.359 |
-46.294 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.836 |
-0.328 |
1.032 |
| y |
-0.328 |
-4.150 |
-0.359 |
| z |
1.032 |
-0.359 |
-0.686 |
|
| Polar |
|---|
| 3z2-r2 | -1.373 |
|---|
| x2-y2 | 5.990 |
|---|
| xy | -0.328 |
|---|
| xz | 1.032 |
|---|
| yz | -0.359 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
203.164 |
| (<r2>)1/2 |
14.254 |