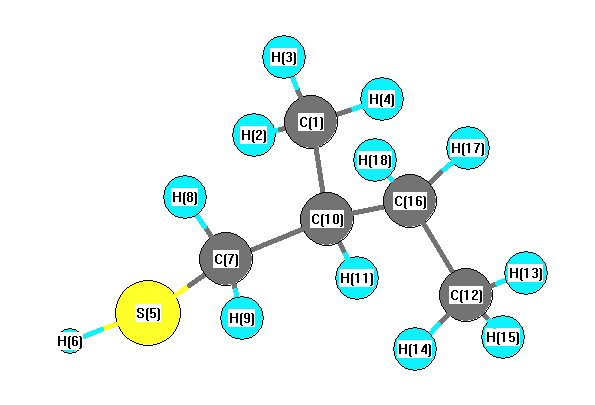
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3000 |
19.38 |
|
|
|
| 2 |
A |
3136 |
2993 |
33.87 |
|
|
|
| 3 |
A |
3129 |
2986 |
57.53 |
|
|
|
| 4 |
A |
3125 |
2982 |
62.01 |
|
|
|
| 5 |
A |
3121 |
2979 |
12.13 |
|
|
|
| 6 |
A |
3078 |
2937 |
24.42 |
|
|
|
| 7 |
A |
3073 |
2933 |
22.51 |
|
|
|
| 8 |
A |
3055 |
2916 |
28.11 |
|
|
|
| 9 |
A |
3053 |
2913 |
38.97 |
|
|
|
| 10 |
A |
3042 |
2903 |
17.72 |
|
|
|
| 11 |
A |
3034 |
2896 |
4.59 |
|
|
|
| 12 |
A |
2672 |
2550 |
13.54 |
|
|
|
| 13 |
A |
1531 |
1461 |
12.80 |
|
|
|
| 14 |
A |
1522 |
1452 |
16.04 |
|
|
|
| 15 |
A |
1520 |
1451 |
4.63 |
|
|
|
| 16 |
A |
1518 |
1449 |
0.87 |
|
|
|
| 17 |
A |
1502 |
1434 |
3.30 |
|
|
|
| 18 |
A |
1494 |
1426 |
6.99 |
|
|
|
| 19 |
A |
1432 |
1367 |
7.11 |
|
|
|
| 20 |
A |
1431 |
1366 |
11.10 |
|
|
|
| 21 |
A |
1403 |
1339 |
2.31 |
|
|
|
| 22 |
A |
1393 |
1329 |
2.10 |
|
|
|
| 23 |
A |
1342 |
1281 |
0.31 |
|
|
|
| 24 |
A |
1328 |
1267 |
16.18 |
|
|
|
| 25 |
A |
1294 |
1235 |
5.87 |
|
|
|
| 26 |
A |
1245 |
1188 |
8.52 |
|
|
|
| 27 |
A |
1196 |
1141 |
0.55 |
|
|
|
| 28 |
A |
1161 |
1108 |
3.52 |
|
|
|
| 29 |
A |
1106 |
1056 |
2.87 |
|
|
|
| 30 |
A |
1064 |
1015 |
1.09 |
|
|
|
| 31 |
A |
1042 |
994 |
2.56 |
|
|
|
| 32 |
A |
990 |
945 |
6.65 |
|
|
|
| 33 |
A |
957 |
913 |
1.30 |
|
|
|
| 34 |
A |
909 |
867 |
4.14 |
|
|
|
| 35 |
A |
882 |
841 |
2.45 |
|
|
|
| 36 |
A |
807 |
770 |
0.25 |
|
|
|
| 37 |
A |
795 |
759 |
8.81 |
|
|
|
| 38 |
A |
731 |
698 |
1.32 |
|
|
|
| 39 |
A |
459 |
438 |
0.25 |
|
|
|
| 40 |
A |
404 |
385 |
0.08 |
|
|
|
| 41 |
A |
382 |
365 |
1.31 |
|
|
|
| 42 |
A |
255 |
243 |
0.31 |
|
|
|
| 43 |
A |
247 |
235 |
0.10 |
|
|
|
| 44 |
A |
208 |
198 |
0.05 |
|
|
|
| 45 |
A |
202 |
192 |
1.86 |
|
|
|
| 46 |
A |
152 |
145 |
19.55 |
|
|
|
| 47 |
A |
93 |
89 |
1.18 |
|
|
|
| 48 |
A |
59 |
56 |
7.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35355.6 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 33743.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.647 |
|
|
|
| 2 |
H |
0.252 |
|
|
|
| 3 |
H |
0.220 |
|
|
|
| 4 |
H |
0.226 |
|
|
|
| 5 |
S |
-0.118 |
|
|
|
| 6 |
H |
0.154 |
|
|
|
| 7 |
C |
-0.587 |
|
|
|
| 8 |
H |
0.250 |
|
|
|
| 9 |
H |
0.267 |
|
|
|
| 10 |
C |
-0.286 |
|
|
|
| 11 |
H |
0.234 |
|
|
|
| 12 |
C |
-0.664 |
|
|
|
| 13 |
H |
0.229 |
|
|
|
| 14 |
H |
0.223 |
|
|
|
| 15 |
H |
0.224 |
|
|
|
| 16 |
C |
-0.422 |
|
|
|
| 17 |
H |
0.226 |
|
|
|
| 18 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.501 |
-0.714 |
0.806 |
1.847 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.122 |
2.570 |
-1.639 |
| y |
2.570 |
-46.343 |
-1.861 |
| z |
-1.639 |
-1.861 |
-48.492 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.705 |
2.570 |
-1.639 |
| y |
2.570 |
2.964 |
-1.861 |
| z |
-1.639 |
-1.861 |
-0.259 |
|
| Polar |
|---|
| 3z2-r2 | -0.519 |
|---|
| x2-y2 | -3.779 |
|---|
| xy | 2.570 |
|---|
| xz | -1.639 |
|---|
| yz | -1.861 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
286.064 |
| (<r2>)1/2 |
16.913 |