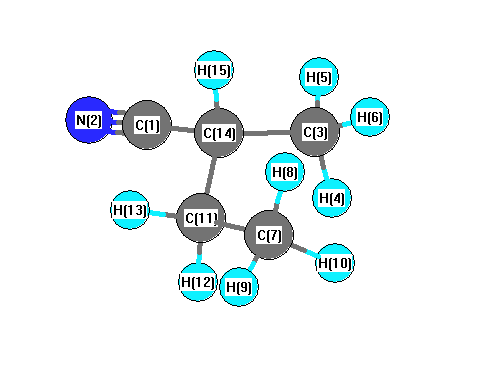
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3275 |
2962 |
37.01 |
|
|
|
| 2 |
A |
3265 |
2953 |
45.89 |
|
|
|
| 3 |
A |
3254 |
2943 |
81.28 |
|
|
|
| 4 |
A |
3243 |
2933 |
55.96 |
|
|
|
| 5 |
A |
3225 |
2916 |
3.02 |
|
|
|
| 6 |
A |
3206 |
2900 |
9.29 |
|
|
|
| 7 |
A |
3199 |
2893 |
11.74 |
|
|
|
| 8 |
A |
3194 |
2888 |
21.75 |
|
|
|
| 9 |
A |
3182 |
2877 |
32.85 |
|
|
|
| 10 |
A |
2587 |
2340 |
16.45 |
|
|
|
| 11 |
A |
1645 |
1488 |
1.99 |
|
|
|
| 12 |
A |
1641 |
1484 |
10.13 |
|
|
|
| 13 |
A |
1632 |
1476 |
5.88 |
|
|
|
| 14 |
A |
1630 |
1474 |
8.42 |
|
|
|
| 15 |
A |
1621 |
1466 |
0.14 |
|
|
|
| 16 |
A |
1557 |
1409 |
2.73 |
|
|
|
| 17 |
A |
1556 |
1407 |
2.36 |
|
|
|
| 18 |
A |
1520 |
1375 |
1.26 |
|
|
|
| 19 |
A |
1478 |
1337 |
1.72 |
|
|
|
| 20 |
A |
1446 |
1308 |
3.54 |
|
|
|
| 21 |
A |
1403 |
1269 |
1.16 |
|
|
|
| 22 |
A |
1293 |
1169 |
2.34 |
|
|
|
| 23 |
A |
1235 |
1117 |
0.19 |
|
|
|
| 24 |
A |
1216 |
1100 |
6.95 |
|
|
|
| 25 |
A |
1114 |
1008 |
1.74 |
|
|
|
| 26 |
A |
1074 |
972 |
0.14 |
|
|
|
| 27 |
A |
1046 |
946 |
3.02 |
|
|
|
| 28 |
A |
955 |
863 |
0.78 |
|
|
|
| 29 |
A |
871 |
788 |
1.16 |
|
|
|
| 30 |
A |
819 |
740 |
2.69 |
|
|
|
| 31 |
A |
628 |
568 |
0.15 |
|
|
|
| 32 |
A |
586 |
530 |
1.69 |
|
|
|
| 33 |
A |
425 |
384 |
0.41 |
|
|
|
| 34 |
A |
353 |
319 |
1.58 |
|
|
|
| 35 |
A |
304 |
275 |
0.30 |
|
|
|
| 36 |
A |
233 |
210 |
0.09 |
|
|
|
| 37 |
A |
226 |
205 |
2.81 |
|
|
|
| 38 |
A |
183 |
166 |
7.20 |
|
|
|
| 39 |
A |
96 |
87 |
1.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30706.9 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 27771.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.182 |
0.235 |
|
0.217 |
| 2 |
N |
-0.338 |
-0.479 |
|
-0.463 |
| 3 |
C |
-0.605 |
-0.198 |
|
-0.364 |
| 4 |
H |
0.226 |
0.055 |
|
0.110 |
| 5 |
H |
0.236 |
0.056 |
|
0.108 |
| 6 |
H |
0.231 |
0.047 |
|
0.093 |
| 7 |
C |
-0.629 |
-0.179 |
|
-0.214 |
| 8 |
H |
0.211 |
0.036 |
|
0.053 |
| 9 |
H |
0.225 |
0.056 |
|
0.070 |
| 10 |
H |
0.218 |
0.039 |
|
0.056 |
| 11 |
C |
-0.396 |
0.107 |
|
0.010 |
| 12 |
H |
0.222 |
-0.016 |
|
0.032 |
| 13 |
H |
0.232 |
-0.021 |
|
0.016 |
| 14 |
C |
-0.268 |
0.279 |
|
0.246 |
| 15 |
H |
0.254 |
-0.018 |
|
0.028 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.040 |
1.402 |
0.544 |
4.311 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
4.059 |
1.419 |
0.536 |
4.333 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.864 |
-4.092 |
-0.970 |
| y |
-4.092 |
-38.062 |
-0.127 |
| z |
-0.970 |
-0.127 |
-37.257 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.205 |
-4.092 |
-0.970 |
| y |
-4.092 |
5.499 |
-0.127 |
| z |
-0.970 |
-0.127 |
6.706 |
|
| Polar |
|---|
| 3z2-r2 | 13.411 |
|---|
| x2-y2 | -11.803 |
|---|
| xy | -4.092 |
|---|
| xz | -0.970 |
|---|
| yz | -0.127 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.775 |
0.451 |
0.239 |
| y |
0.451 |
7.697 |
0.112 |
| z |
0.239 |
0.112 |
6.824 |
<r2> (average value of r
2) Å
2
| <r2> |
198.641 |
| (<r2>)1/2 |
14.094 |