return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Butanenitrile, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-250.039856
Energy at 298.15K-250.049380
HF Energy-249.074227
Nuclear repulsion energy220.160568
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3193 3024 14.33      
2 A 3182 3013 15.95      
3 A 3180 3012 29.65      
4 A 3169 3001 25.41      
5 A 3140 2974 7.10      
6 A 3092 2928 10.86      
7 A 3090 2927 23.57      
8 A 3083 2920 9.51      
9 A 3082 2919 5.96      
10 A 2195 2079 0.12      
11 A 1555 1473 3.29      
12 A 1544 1462 18.99      
13 A 1538 1456 11.02      
14 A 1533 1452 7.33      
15 A 1524 1443 0.33      
16 A 1453 1376 6.80      
17 A 1448 1371 8.40      
18 A 1408 1334 0.73      
19 A 1378 1305 0.78      
20 A 1345 1274 0.79      
21 A 1311 1242 0.48      
22 A 1213 1148 1.76      
23 A 1173 1111 0.16      
24 A 1139 1079 5.82      
25 A 1082 1025 1.37      
26 A 1025 971 1.47      
27 A 993 941 5.31      
28 A 924 875 0.39      
29 A 827 783 0.65      
30 A 781 739 3.43      
31 A 564 534 0.09      
32 A 538 509 0.89      
33 A 410 389 0.25      
34 A 320 303 0.43      
35 A 305 288 0.18      
36 A 217 206 0.06      
37 A 204 193 1.96      
38 A 161 152 4.99      
39 A 96 91 1.42      

Unscaled Zero Point Vibrational Energy (zpe) 29206.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 27661.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.21926 0.07363 0.05905

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.521 -0.279 0.121
N2 -2.607 -0.663 -0.108
C3 -0.003 1.630 -0.152
H4 -0.077 1.645 -1.242
H5 -0.783 2.279 0.252
H6 0.964 2.042 0.137
C7 2.301 -0.418 0.054
H8 2.465 -0.296 1.129
H9 2.972 -1.205 -0.297
H10 2.596 0.508 -0.442
C11 0.850 -0.787 -0.241
H12 0.681 -0.829 -1.322
H13 0.636 -1.787 0.149
C14 -0.158 0.199 0.374
H15 -0.018 0.204 1.462

Atom - Atom Distances (Å)
  C1 N2 C3 H4 H5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.17432.45432.76452.66593.40083.82544.11214.60704.22972.45232.68942.63261.46662.0720
N21.17433.47003.60723.48094.48704.91685.23385.60885.34403.46213.50853.44212.64073.1500
C32.45433.47001.09281.09141.09073.09003.38334.11262.84632.56512.80823.48961.53242.1539
H42.76453.60721.09281.76981.77353.40423.98124.27903.01302.78832.58833.77072.16943.0644
H52.66593.48091.09141.76981.76654.10164.23705.15153.87733.50873.77894.30792.17532.5211
H63.40084.48701.09071.77351.76652.80102.95033.84222.31332.85643.23333.84312.17072.4692
C73.82544.91683.09003.40424.10162.80101.09381.09211.09131.52552.16512.15732.55542.7833
H84.11215.23383.38333.98124.23702.95031.09381.76461.76912.17333.07792.55482.77452.5550
H94.60705.60884.11264.27905.15153.84221.09211.76461.75972.16342.53842.44853.49583.7444
H104.22975.34402.84633.01303.87732.31331.09131.76911.75972.18322.49673.07552.88913.2481
C112.45233.46212.56512.78833.50872.85641.52552.17332.16342.18321.09541.09441.53842.1530
H122.68943.50852.80822.58833.77893.23332.16513.07792.53842.49671.09541.75572.15323.0507
H132.63263.44213.48963.77074.30793.84312.15732.55482.44853.07551.09441.75572.15092.4733
C141.46662.64071.53242.16942.17532.17072.55542.77453.49582.88911.53842.15322.15091.0975
H152.07203.15002.15393.06442.52112.46922.78332.55503.74443.24812.15303.05072.47331.0975

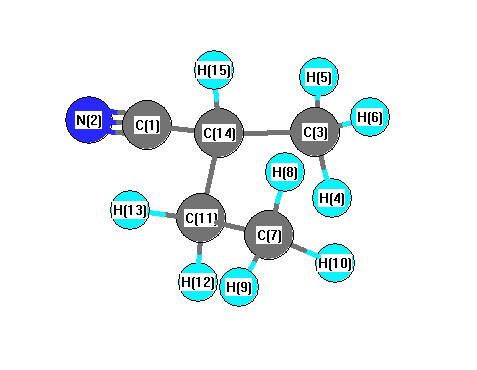
picture of Butanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C3 109.822 C1 C14 C11 109.363
C1 C14 H15 106.941 N2 C1 C14 178.675
C3 C14 C11 113.297 C3 C14 H15 108.841
H4 C3 H5 108.251 H4 C3 H6 108.632
H4 C3 C14 110.331 H5 C3 H6 108.106
H5 C3 C14 110.882 H6 C3 C14 110.558
C7 C11 H12 110.322 C7 C11 H13 109.771
C7 C11 C14 113.032 H8 C7 H9 107.661
H8 C7 H10 108.123 H8 C7 C11 111.072
H9 C7 H10 107.401 H9 C7 C11 110.387
H10 C7 C11 112.019 C11 C14 H15 108.369
H12 C11 H13 106.604 H12 C11 C14 108.501
H13 C11 C14 108.386
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability