Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -114.395302 |
| Energy at 298.15K | -114.396724 |
| HF Energy | -114.395302 |
| Nuclear repulsion energy | 30.780534 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3705 |
3567 |
43.92 |
|
|
|
| 2 |
A' |
2856 |
2749 |
164.15 |
|
|
|
| 3 |
A' |
1544 |
1486 |
29.01 |
|
|
|
| 4 |
A' |
1352 |
1302 |
67.78 |
|
|
|
| 5 |
A' |
1246 |
1199 |
128.06 |
|
|
|
| 6 |
A" |
1132 |
1090 |
133.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5917.4 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 5696.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.741 |
0.000 |
| O2 |
0.011 |
-0.566 |
0.000 |
| H3 |
-1.084 |
0.966 |
0.000 |
| H4 |
0.926 |
-0.883 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3080 | 1.1177 | 1.8643 |
O2 | 1.3080 | | 1.8835 | 0.9675 | H3 | 1.1177 | 1.8835 | | 2.7307 | H4 | 1.8643 | 0.9675 | 2.7307 | |
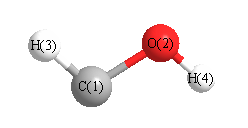 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
109.110 |
|
O2 |
C1 |
H3 |
101.591 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.200 |
|
|
|
| 2 |
O |
-0.393 |
|
|
|
| 3 |
H |
0.171 |
|
|
|
| 4 |
H |
0.423 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.311 |
-1.675 |
0.000 |
1.704 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-10.934 |
-3.814 |
0.000 |
| y |
-3.814 |
-13.663 |
0.000 |
| z |
0.000 |
0.000 |
-11.914 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.855 |
-3.814 |
0.000 |
| y |
-3.814 |
-2.239 |
0.000 |
| z |
0.000 |
0.000 |
0.384 |
|
| Polar |
|---|
| 3z2-r2 | 0.769 |
|---|
| x2-y2 | 2.729 |
|---|
| xy | -3.814 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.495 |
-0.551 |
0.000 |
| y |
-0.551 |
2.504 |
0.000 |
| z |
0.000 |
0.000 |
1.530 |
<r2> (average value of r
2) Å
2
| <r2> |
17.213 |
| (<r2>)1/2 |
4.149 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -114.386211 |
| Energy at 298.15K | -114.387622 |
| HF Energy | -114.386211 |
| Nuclear repulsion energy | 30.703757 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3484 |
3354 |
35.81 |
|
|
|
| 2 |
A' |
2742 |
2640 |
254.05 |
|
|
|
| 3 |
A' |
1497 |
1441 |
65.40 |
|
|
|
| 4 |
A' |
1357 |
1307 |
58.99 |
|
|
|
| 5 |
A' |
1261 |
1214 |
23.71 |
|
|
|
| 6 |
A" |
1043 |
1004 |
43.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5691.9 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 5479.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.122 |
0.740 |
0.000 |
| O2 |
0.122 |
-0.562 |
0.000 |
| H3 |
-0.955 |
1.068 |
0.000 |
| H4 |
-0.749 |
-1.007 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3022 | 1.1260 | 1.9517 |
O2 | 1.3022 | | 1.9542 | 0.9773 | H3 | 1.1260 | 1.9542 | | 2.0856 | H4 | 1.9517 | 0.9773 | 2.0856 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
117.069 |
|
O2 |
C1 |
H3 |
106.953 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.145 |
|
|
|
| 2 |
O |
-0.389 |
|
|
|
| 3 |
H |
0.140 |
|
|
|
| 4 |
H |
0.394 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.145 |
-1.738 |
0.000 |
3.594 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.300 |
-0.025 |
0.000 |
| y |
-0.025 |
-13.142 |
0.000 |
| z |
0.000 |
0.000 |
-11.863 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.202 |
-0.025 |
0.000 |
| y |
-0.025 |
-1.060 |
0.000 |
| z |
0.000 |
0.000 |
0.858 |
|
| Polar |
|---|
| 3z2-r2 | 1.716 |
|---|
| x2-y2 | 0.842 |
|---|
| xy | -0.025 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.463 |
-0.021 |
0.000 |
| y |
-0.021 |
3.006 |
0.000 |
| z |
0.000 |
0.000 |
1.527 |
<r2> (average value of r
2) Å
2
| <r2> |
17.417 |
| (<r2>)1/2 |
4.173 |
