Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -114.161917 |
| Energy at 298.15K | -114.163347 |
| HF Energy | -113.813945 |
| Nuclear repulsion energy | 30.795315 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3778 |
3578 |
80.36 |
|
|
|
| 2 |
A' |
2968 |
2811 |
140.13 |
|
|
|
| 3 |
A' |
1565 |
1482 |
25.99 |
|
|
|
| 4 |
A' |
1385 |
1312 |
73.17 |
|
|
|
| 5 |
A' |
1260 |
1193 |
132.48 |
|
|
|
| 6 |
A" |
1170 |
1108 |
137.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6063.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 5742.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.741 |
0.000 |
| O2 |
0.011 |
-0.568 |
0.000 |
| H3 |
-1.075 |
0.971 |
0.000 |
| H4 |
0.926 |
-0.872 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3099 | 1.1098 | 1.8550 |
O2 | 1.3099 | | 1.8841 | 0.9644 | H3 | 1.1098 | 1.8841 | | 2.7207 | H4 | 1.8550 | 0.9644 | 2.7207 | |
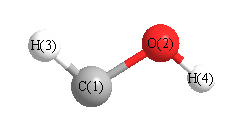 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
108.343 |
|
O2 |
C1 |
H3 |
101.962 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -114.151643 |
| Energy at 298.15K | -114.153063 |
| HF Energy | -113.804190 |
| Nuclear repulsion energy | 30.712962 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3633 |
3441 |
8.68 |
|
|
|
| 2 |
A' |
2848 |
2697 |
222.37 |
|
|
|
| 3 |
A' |
1521 |
1440 |
54.22 |
|
|
|
| 4 |
A' |
1387 |
1314 |
89.09 |
|
|
|
| 5 |
A' |
1293 |
1225 |
28.10 |
|
|
|
| 6 |
A" |
1077 |
1021 |
45.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5879.7 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 5568.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.122 |
0.741 |
0.000 |
| O2 |
0.122 |
-0.565 |
0.000 |
| H3 |
-0.949 |
1.066 |
0.000 |
| H4 |
-0.753 |
-0.988 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3058 | 1.1192 | 1.9369 |
O2 | 1.3058 | | 1.9515 | 0.9711 | H3 | 1.1192 | 1.9515 | | 2.0632 | H4 | 1.9369 | 0.9711 | 2.0632 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
115.784 |
|
O2 |
C1 |
H3 |
106.915 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
