Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -117.150565 |
| Energy at 298.15K | -117.154830 |
| HF Energy | -117.010419 |
| Nuclear repulsion energy | 64.993077 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3272 |
3272 |
23.85 |
|
|
|
| 2 |
A1 |
3175 |
3175 |
1.10 |
|
|
|
| 3 |
A1 |
3167 |
3167 |
20.49 |
|
|
|
| 4 |
A1 |
1545 |
1545 |
3.26 |
|
|
|
| 5 |
A1 |
1293 |
1293 |
0.85 |
|
|
|
| 6 |
A1 |
1052 |
1052 |
0.10 |
|
|
|
| 7 |
A1 |
434 |
434 |
0.04 |
|
|
|
| 8 |
A2 |
773 |
773 |
0.00 |
|
|
|
| 9 |
A2 |
560 |
560 |
0.00 |
|
|
|
| 10 |
B1 |
1016 |
1016 |
26.06 |
|
|
|
| 11 |
B1 |
795 |
795 |
87.56 |
|
|
|
| 12 |
B1 |
536 |
536 |
18.95 |
|
|
|
| 13 |
B2 |
3269 |
3269 |
6.93 |
|
|
|
| 14 |
B2 |
3167 |
3167 |
11.59 |
|
|
|
| 15 |
B2 |
1528 |
1528 |
1.08 |
|
|
|
| 16 |
B2 |
1440 |
1440 |
7.06 |
|
|
|
| 17 |
B2 |
1192 |
1192 |
0.07 |
|
|
|
| 18 |
B2 |
952 |
952 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14581.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14581.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.444 |
| H2 |
0.000 |
0.000 |
1.531 |
| C3 |
0.000 |
1.226 |
-0.196 |
| C4 |
0.000 |
-1.226 |
-0.196 |
| H5 |
0.000 |
2.154 |
0.358 |
| H6 |
0.000 |
-2.154 |
0.358 |
| H7 |
0.000 |
1.292 |
-1.278 |
| H8 |
0.000 |
-1.292 |
-1.278 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0874 | 1.3827 | 1.3827 | 2.1559 | 2.1559 | 2.1527 | 2.1527 |
H2 | 1.0874 | | 2.1181 | 2.1181 | 2.4529 | 2.4529 | 3.0921 | 3.0921 | C3 | 1.3827 | 2.1181 | | 2.4512 | 1.0815 | 3.4250 | 1.0838 | 2.7403 | C4 | 1.3827 | 2.1181 | 2.4512 | | 3.4250 | 1.0815 | 2.7403 | 1.0838 | H5 | 2.1559 | 2.4529 | 1.0815 | 3.4250 | | 4.3085 | 1.8494 | 3.8150 | H6 | 2.1559 | 2.4529 | 3.4250 | 1.0815 | 4.3085 | | 3.8150 | 1.8494 | H7 | 2.1527 | 3.0921 | 1.0838 | 2.7403 | 1.8494 | 3.8150 | | 2.5842 | H8 | 2.1527 | 3.0921 | 2.7403 | 1.0838 | 3.8150 | 1.8494 | 2.5842 | |
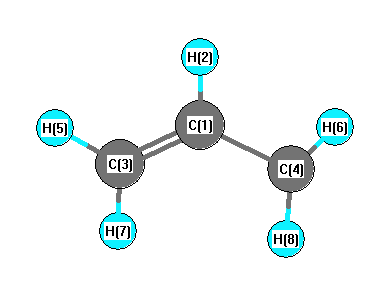 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.583 |
|
C1 |
C3 |
H7 |
121.093 |
| C1 |
C4 |
H6 |
121.583 |
|
C1 |
C4 |
H8 |
121.093 |
| H2 |
C1 |
C3 |
117.577 |
|
H2 |
C1 |
C4 |
117.577 |
| C3 |
C1 |
C4 |
124.846 |
|
H5 |
C3 |
H7 |
117.324 |
| H6 |
C4 |
H8 |
117.324 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -117.150565 |
| Energy at 298.15K | -117.154831 |
| HF Energy | -117.010418 |
| Nuclear repulsion energy | 64.995319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3271 |
3271 |
23.87 |
|
|
|
| 2 |
A' |
3268 |
3268 |
6.95 |
|
|
|
| 3 |
A' |
3174 |
3174 |
0.91 |
|
|
|
| 4 |
A' |
3167 |
3167 |
20.49 |
|
|
|
| 5 |
A' |
3166 |
3166 |
11.73 |
|
|
|
| 6 |
A' |
1545 |
1545 |
3.27 |
|
|
|
| 7 |
A' |
1528 |
1528 |
1.08 |
|
|
|
| 8 |
A' |
1440 |
1440 |
7.06 |
|
|
|
| 9 |
A' |
1293 |
1293 |
0.85 |
|
|
|
| 10 |
A' |
1192 |
1192 |
0.07 |
|
|
|
| 11 |
A' |
1052 |
1052 |
0.10 |
|
|
|
| 12 |
A' |
952 |
952 |
0.07 |
|
|
|
| 13 |
A' |
434 |
434 |
0.04 |
|
|
|
| 14 |
A" |
1016 |
1016 |
26.06 |
|
|
|
| 15 |
A" |
795 |
795 |
87.58 |
|
|
|
| 16 |
A" |
773 |
773 |
0.01 |
|
|
|
| 17 |
A" |
560 |
560 |
0.00 |
|
|
|
| 18 |
A" |
537 |
537 |
18.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14581.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14581.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.444 |
0.000 |
| H2 |
0.000 |
1.531 |
0.000 |
| C3 |
-1.225 |
-0.196 |
0.000 |
| C4 |
1.225 |
-0.196 |
0.000 |
| H5 |
-2.154 |
0.358 |
0.000 |
| H6 |
2.154 |
0.358 |
0.000 |
| H7 |
-1.291 |
-1.278 |
0.000 |
| H8 |
1.292 |
-1.278 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0874 | 1.3828 | 1.3826 | 2.1561 | 2.1559 | 2.1525 | 2.1526 |
H2 | 1.0874 | | 2.1184 | 2.1180 | 2.4534 | 2.4529 | 3.0922 | 3.0921 | C3 | 1.3828 | 2.1184 | | 2.4509 | 1.0816 | 3.4248 | 1.0838 | 2.7397 | C4 | 1.3826 | 2.1180 | 2.4509 | | 3.4249 | 1.0815 | 2.7393 | 1.0838 | H5 | 2.1561 | 2.4534 | 1.0816 | 3.4249 | | 4.3085 | 1.8497 | 3.8146 | H6 | 2.1559 | 2.4529 | 3.4248 | 1.0815 | 4.3085 | | 3.8141 | 1.8494 | H7 | 2.1525 | 3.0922 | 1.0838 | 2.7393 | 1.8497 | 3.8141 | | 2.5829 | H8 | 2.1526 | 3.0921 | 2.7397 | 1.0838 | 3.8146 | 1.8494 | 2.5829 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.587 |
|
C1 |
C3 |
H7 |
121.067 |
| C1 |
C4 |
H6 |
121.589 |
|
C1 |
C4 |
H8 |
121.093 |
| H2 |
C1 |
C3 |
117.597 |
|
H2 |
C1 |
C4 |
117.581 |
| C3 |
C1 |
C4 |
124.822 |
|
H5 |
C3 |
H7 |
117.346 |
| H6 |
C4 |
H8 |
117.318 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
