Jump to
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.101604 |
| Energy at 298.15K | -133.107196 |
| HF Energy | -133.101604 |
| Nuclear repulsion energy | 71.485373 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3738 |
3381 |
0.64 |
|
|
|
| 2 |
A' |
3288 |
2973 |
27.26 |
|
|
|
| 3 |
A' |
3200 |
2894 |
66.39 |
|
|
|
| 4 |
A' |
3177 |
2873 |
26.83 |
|
|
|
| 5 |
A' |
1903 |
1721 |
93.94 |
|
|
|
| 6 |
A' |
1608 |
1454 |
10.86 |
|
|
|
| 7 |
A' |
1581 |
1429 |
41.07 |
|
|
|
| 8 |
A' |
1527 |
1381 |
8.03 |
|
|
|
| 9 |
A' |
1389 |
1256 |
26.54 |
|
|
|
| 10 |
A' |
1158 |
1047 |
27.66 |
|
|
|
| 11 |
A' |
981 |
887 |
5.89 |
|
|
|
| 12 |
A' |
523 |
473 |
20.24 |
|
|
|
| 13 |
A" |
3227 |
2918 |
37.93 |
|
|
|
| 14 |
A" |
1611 |
1457 |
9.00 |
|
|
|
| 15 |
A" |
1253 |
1134 |
8.48 |
|
|
|
| 16 |
A" |
1181 |
1068 |
22.33 |
|
|
|
| 17 |
A" |
744 |
673 |
59.64 |
|
|
|
| 18 |
A" |
196 |
178 |
2.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16142.0 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 14598.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.019 |
-0.653 |
0.000 |
| C2 |
0.000 |
0.446 |
0.000 |
| N3 |
1.228 |
0.214 |
0.000 |
| H4 |
-0.527 |
-1.616 |
0.000 |
| H5 |
-1.659 |
-0.578 |
0.874 |
| H6 |
-1.659 |
-0.578 |
-0.874 |
| H7 |
-0.399 |
1.458 |
0.000 |
| H8 |
1.761 |
1.063 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4990 | 2.4088 | 1.0810 | 1.0863 | 1.0863 | 2.2008 | 3.2674 |
C2 | 1.4990 | | 1.2500 | 2.1284 | 2.1369 | 2.1369 | 1.0882 | 1.8662 | N3 | 2.4088 | 1.2500 | | 2.5359 | 3.1194 | 3.1194 | 2.0487 | 1.0025 | H4 | 1.0810 | 2.1284 | 2.5359 | | 1.7674 | 1.7674 | 3.0771 | 3.5235 | H5 | 1.0863 | 2.1369 | 3.1194 | 1.7674 | | 1.7483 | 2.5493 | 3.8933 | H6 | 1.0863 | 2.1369 | 3.1194 | 1.7674 | 1.7483 | | 2.5493 | 3.8933 | H7 | 2.2008 | 1.0882 | 2.0487 | 3.0771 | 2.5493 | 2.5493 | | 2.1963 | H8 | 3.2674 | 1.8662 | 1.0025 | 3.5235 | 3.8933 | 3.8933 | 2.1963 | |
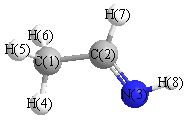 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
122.127 |
|
C1 |
C2 |
H7 |
115.655 |
| C2 |
C1 |
H4 |
110.119 |
|
C2 |
C1 |
H5 |
110.479 |
| C2 |
C1 |
H6 |
110.479 |
|
C2 |
N3 |
H8 |
111.418 |
| N3 |
C2 |
H7 |
122.218 |
|
H4 |
C1 |
H5 |
109.271 |
| H4 |
C1 |
H6 |
109.271 |
|
H5 |
C1 |
H6 |
107.161 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.651 |
|
|
|
| 2 |
C |
0.038 |
|
|
|
| 3 |
N |
-0.534 |
|
|
|
| 4 |
H |
0.239 |
|
|
|
| 5 |
H |
0.218 |
|
|
|
| 6 |
H |
0.218 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.295 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.556 |
1.586 |
0.000 |
2.221 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.161 |
2.512 |
0.000 |
| y |
2.512 |
-17.572 |
0.000 |
| z |
0.000 |
0.000 |
-19.963 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.394 |
2.512 |
0.000 |
| y |
2.512 |
2.490 |
0.000 |
| z |
0.000 |
0.000 |
-1.096 |
|
| Polar |
|---|
| 3z2-r2 | -2.192 |
|---|
| x2-y2 | -2.589 |
|---|
| xy | 2.512 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.594 |
0.230 |
0.000 |
| y |
0.230 |
3.969 |
0.000 |
| z |
0.000 |
0.000 |
2.931 |
<r2> (average value of r
2) Å
2
| <r2> |
49.990 |
| (<r2>)1/2 |
7.070 |
Jump to
S1C1
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.099974 |
| Energy at 298.15K | -133.105538 |
| HF Energy | -133.099974 |
| Nuclear repulsion energy | 71.457321 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3703 |
3349 |
3.29 |
|
|
|
| 2 |
A' |
3266 |
2954 |
83.36 |
|
|
|
| 3 |
A' |
3245 |
2935 |
1.97 |
|
|
|
| 4 |
A' |
3179 |
2875 |
18.04 |
|
|
|
| 5 |
A' |
1904 |
1722 |
100.74 |
|
|
|
| 6 |
A' |
1614 |
1459 |
26.83 |
|
|
|
| 7 |
A' |
1556 |
1407 |
19.00 |
|
|
|
| 8 |
A' |
1532 |
1385 |
3.04 |
|
|
|
| 9 |
A' |
1399 |
1266 |
67.70 |
|
|
|
| 10 |
A' |
1153 |
1043 |
21.86 |
|
|
|
| 11 |
A' |
969 |
877 |
1.28 |
|
|
|
| 12 |
A' |
528 |
477 |
10.22 |
|
|
|
| 13 |
A" |
3234 |
2925 |
30.25 |
|
|
|
| 14 |
A" |
1606 |
1453 |
9.49 |
|
|
|
| 15 |
A" |
1259 |
1139 |
51.90 |
|
|
|
| 16 |
A" |
1184 |
1071 |
15.00 |
|
|
|
| 17 |
A" |
756 |
684 |
16.31 |
|
|
|
| 18 |
A" |
173 |
157 |
0.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16130.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 14588.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.006 |
-0.636 |
0.000 |
| C2 |
0.000 |
0.483 |
0.000 |
| N3 |
1.246 |
0.392 |
0.000 |
| H4 |
-0.526 |
-1.608 |
0.000 |
| H5 |
-1.647 |
-0.566 |
0.873 |
| H6 |
-1.647 |
-0.566 |
-0.873 |
| H7 |
-0.417 |
1.483 |
0.000 |
| H8 |
1.547 |
-0.567 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5045 | 2.4756 | 1.0841 | 1.0858 | 1.0858 | 2.1998 | 2.5533 |
C2 | 1.5045 | | 1.2495 | 2.1559 | 2.1387 | 2.1387 | 1.0840 | 1.8691 | N3 | 2.4756 | 1.2495 | | 2.6721 | 3.1702 | 3.1702 | 1.9891 | 1.0048 | H4 | 1.0841 | 2.1559 | 2.6721 | | 1.7627 | 1.7627 | 3.0934 | 2.3191 | H5 | 1.0858 | 2.1387 | 3.1702 | 1.7627 | | 1.7470 | 2.5445 | 3.3108 | H6 | 1.0858 | 2.1387 | 3.1702 | 1.7627 | 1.7470 | | 2.5445 | 3.3108 | H7 | 2.1998 | 1.0840 | 1.9891 | 3.0934 | 2.5445 | 2.5445 | | 2.8387 | H8 | 2.5533 | 1.8691 | 1.0048 | 2.3191 | 3.3108 | 3.3108 | 2.8387 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.794 |
|
C1 |
C2 |
H7 |
115.434 |
| C2 |
C1 |
H4 |
111.755 |
|
C2 |
C1 |
H5 |
110.270 |
| C2 |
C1 |
H6 |
110.270 |
|
C2 |
N3 |
H8 |
111.553 |
| N3 |
C2 |
H7 |
116.771 |
|
H4 |
C1 |
H5 |
108.646 |
| H4 |
C1 |
H6 |
108.646 |
|
H5 |
C1 |
H6 |
107.119 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.679 |
|
|
|
| 2 |
C |
0.060 |
|
|
|
| 3 |
N |
-0.543 |
|
|
|
| 4 |
H |
0.216 |
|
|
|
| 5 |
H |
0.228 |
|
|
|
| 6 |
H |
0.228 |
|
|
|
| 7 |
H |
0.206 |
|
|
|
| 8 |
H |
0.285 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.130 |
-1.551 |
0.000 |
2.635 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.544 |
-3.625 |
0.000 |
| y |
-3.625 |
-19.170 |
0.000 |
| z |
0.000 |
0.000 |
-19.902 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.008 |
-3.625 |
0.000 |
| y |
-3.625 |
1.553 |
0.000 |
| z |
0.000 |
0.000 |
0.455 |
|
| Polar |
|---|
| 3z2-r2 | 0.909 |
|---|
| x2-y2 | -2.374 |
|---|
| xy | -3.625 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.463 |
-0.137 |
0.000 |
| y |
-0.137 |
3.967 |
0.000 |
| z |
0.000 |
0.000 |
2.910 |
<r2> (average value of r
2) Å
2
| <r2> |
50.003 |
| (<r2>)1/2 |
7.071 |
