Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.596852 |
| Energy at 298.15K | -133.602344 |
| HF Energy | -133.099821 |
| Nuclear repulsion energy | 70.819175 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3502 |
3316 |
1.55 |
|
|
|
| 2 |
A' |
3202 |
3033 |
8.98 |
|
|
|
| 3 |
A' |
3082 |
2919 |
13.19 |
|
|
|
| 4 |
A' |
3068 |
2906 |
58.54 |
|
|
|
| 5 |
A' |
1715 |
1624 |
38.51 |
|
|
|
| 6 |
A' |
1515 |
1435 |
13.77 |
|
|
|
| 7 |
A' |
1463 |
1386 |
24.63 |
|
|
|
| 8 |
A' |
1424 |
1348 |
19.16 |
|
|
|
| 9 |
A' |
1312 |
1242 |
33.79 |
|
|
|
| 10 |
A' |
1091 |
1033 |
28.12 |
|
|
|
| 11 |
A' |
951 |
901 |
5.26 |
|
|
|
| 12 |
A' |
496 |
470 |
17.74 |
|
|
|
| 13 |
A" |
3158 |
2991 |
14.04 |
|
|
|
| 14 |
A" |
1517 |
1437 |
12.14 |
|
|
|
| 15 |
A" |
1142 |
1082 |
6.20 |
|
|
|
| 16 |
A" |
1106 |
1047 |
14.48 |
|
|
|
| 17 |
A" |
697 |
660 |
63.17 |
|
|
|
| 18 |
A" |
191 |
181 |
1.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15315.2 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14505.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.039 |
-0.623 |
0.000 |
| C2 |
0.000 |
0.454 |
0.000 |
| N3 |
1.247 |
0.167 |
0.000 |
| H4 |
-0.551 |
-1.597 |
0.000 |
| H5 |
-1.682 |
-0.538 |
0.880 |
| H6 |
-1.682 |
-0.538 |
-0.880 |
| H7 |
-0.375 |
1.487 |
0.000 |
| H8 |
1.799 |
1.026 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4963 | 2.4183 | 1.0896 | 1.0937 | 1.0937 | 2.2120 | 3.2815 |
C2 | 1.4963 | | 1.2792 | 2.1237 | 2.1423 | 2.1423 | 1.0989 | 1.8871 | N3 | 2.4183 | 1.2792 | | 2.5187 | 3.1387 | 3.1387 | 2.0907 | 1.0204 | H4 | 1.0896 | 2.1237 | 2.5187 | | 1.7825 | 1.7825 | 3.0891 | 3.5210 | H5 | 1.0937 | 2.1423 | 3.1387 | 1.7825 | | 1.7608 | 2.5662 | 3.9161 | H6 | 1.0937 | 2.1423 | 3.1387 | 1.7825 | 1.7608 | | 2.5662 | 3.9161 | H7 | 2.2120 | 1.0989 | 2.0907 | 3.0891 | 2.5662 | 2.5662 | | 2.2217 | H8 | 3.2815 | 1.8871 | 1.0204 | 3.5210 | 3.9161 | 3.9161 | 2.2217 | |
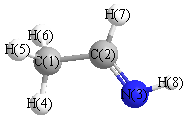 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.021 |
|
C1 |
C2 |
H7 |
116.085 |
| C2 |
C1 |
H4 |
109.413 |
|
C2 |
C1 |
H5 |
110.643 |
| C2 |
C1 |
H6 |
110.643 |
|
C2 |
N3 |
H8 |
109.784 |
| N3 |
C2 |
H7 |
122.894 |
|
H4 |
C1 |
H5 |
109.450 |
| H4 |
C1 |
H6 |
109.450 |
|
H5 |
C1 |
H6 |
107.205 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.594939 |
| Energy at 298.15K | -133.600372 |
| HF Energy | -133.098181 |
| Nuclear repulsion energy | 70.692392 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3458 |
3275 |
6.33 |
|
|
|
| 2 |
A' |
3177 |
3009 |
25.60 |
|
|
|
| 3 |
A' |
3138 |
2972 |
24.91 |
|
|
|
| 4 |
A' |
3075 |
2912 |
8.31 |
|
|
|
| 5 |
A' |
1704 |
1614 |
38.34 |
|
|
|
| 6 |
A' |
1520 |
1440 |
29.49 |
|
|
|
| 7 |
A' |
1456 |
1379 |
23.43 |
|
|
|
| 8 |
A' |
1433 |
1358 |
9.37 |
|
|
|
| 9 |
A' |
1310 |
1241 |
56.35 |
|
|
|
| 10 |
A' |
1090 |
1032 |
22.03 |
|
|
|
| 11 |
A' |
929 |
880 |
2.98 |
|
|
|
| 12 |
A' |
497 |
471 |
8.46 |
|
|
|
| 13 |
A" |
3159 |
2992 |
10.32 |
|
|
|
| 14 |
A" |
1512 |
1432 |
12.55 |
|
|
|
| 15 |
A" |
1169 |
1107 |
52.72 |
|
|
|
| 16 |
A" |
1094 |
1036 |
9.83 |
|
|
|
| 17 |
A" |
687 |
651 |
8.49 |
|
|
|
| 18 |
A" |
157 |
149 |
0.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15283.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14474.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.027 |
-0.616 |
0.000 |
| C2 |
0.000 |
0.482 |
0.000 |
| N3 |
1.274 |
0.370 |
0.000 |
| H4 |
-0.555 |
-1.601 |
0.000 |
| H5 |
-1.672 |
-0.535 |
0.880 |
| H6 |
-1.672 |
-0.535 |
-0.880 |
| H7 |
-0.391 |
1.503 |
0.000 |
| H8 |
1.532 |
-0.620 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5031 | 2.5038 | 1.0925 | 1.0937 | 1.0937 | 2.2123 | 2.5591 |
C2 | 1.5031 | | 1.2791 | 2.1556 | 2.1450 | 2.1450 | 1.0939 | 1.8870 | N3 | 2.5038 | 1.2791 | | 2.6897 | 3.2049 | 3.2049 | 2.0143 | 1.0235 | H4 | 1.0925 | 2.1556 | 2.6897 | | 1.7770 | 1.7770 | 3.1087 | 2.3064 | H5 | 1.0937 | 2.1450 | 3.2049 | 1.7770 | | 1.7596 | 2.5624 | 3.3234 | H6 | 1.0937 | 2.1450 | 3.2049 | 1.7770 | 1.7596 | | 2.5624 | 3.3234 | H7 | 2.2123 | 1.0939 | 2.0143 | 3.1087 | 2.5624 | 2.5624 | | 2.8648 | H8 | 2.5591 | 1.8870 | 1.0235 | 2.3064 | 3.3234 | 3.3234 | 2.8648 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.111 |
|
C1 |
C2 |
H7 |
115.940 |
| C2 |
C1 |
H4 |
111.316 |
|
C2 |
C1 |
H5 |
110.393 |
| C2 |
C1 |
H6 |
110.393 |
|
C2 |
N3 |
H8 |
109.576 |
| N3 |
C2 |
H7 |
115.949 |
|
H4 |
C1 |
H5 |
108.755 |
| H4 |
C1 |
H6 |
108.755 |
|
H5 |
C1 |
H6 |
107.110 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
