return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (2-Butanethiol, 3-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-594.939930
Energy at 298.15K-594.953017
HF Energy-593.891539
Nuclear repulsion energy316.727795
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3185 3017 19.59      
2 A 3183 3015 20.88      
3 A 3174 3006 21.70      
4 A 3166 2998 27.61      
5 A 3160 2993 49.71      
6 A 3154 2987 0.42      
7 A 3085 2922 31.93      
8 A 3079 2916 15.03      
9 A 3075 2912 20.72      
10 A 3072 2909 11.59      
11 A 3063 2901 1.95      
12 A 2681 2539 11.53      
13 A 1563 1480 4.20      
14 A 1544 1462 25.98      
15 A 1541 1459 8.65      
16 A 1530 1449 3.50      
17 A 1524 1443 3.69      
18 A 1518 1438 3.49      
19 A 1458 1380 10.68      
20 A 1445 1369 15.07      
21 A 1438 1362 6.66      
22 A 1407 1333 0.83      
23 A 1376 1303 0.99      
24 A 1348 1277 5.53      
25 A 1293 1225 26.08      
26 A 1230 1165 2.99      
27 A 1207 1143 5.69      
28 A 1187 1124 0.83      
29 A 1118 1059 1.89      
30 A 1073 1016 9.88      
31 A 1006 953 2.96      
32 A 993 940 0.74      
33 A 959 908 1.48      
34 A 949 899 1.34      
35 A 921 872 4.63      
36 A 818 774 2.23      
37 A 721 682 1.30      
38 A 491 465 0.34      
39 A 419 397 0.36      
40 A 390 369 0.06      
41 A 385 364 0.41      
42 A 342 324 1.08      
43 A 268 254 0.74      
44 A 242 229 3.44      
45 A 238 226 0.48      
46 A 195 184 1.80      
47 A 178 169 19.66      
48 A 62 59 2.75      

Unscaled Zero Point Vibrational Energy (zpe) 35725.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 33835.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.12345 0.07135 0.04851

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.194 -1.642 0.161
H2 -2.084 -2.048 -0.326
H3 -0.360 -2.312 -0.053
H4 -1.367 -1.657 1.242
C5 0.534 1.861 0.004
H6 -0.271 2.489 0.392
H7 1.470 2.228 0.431
H8 0.589 1.985 -1.081
S9 1.759 -0.610 -0.133
H10 2.669 0.133 0.522
C11 0.309 0.396 0.358
H12 0.185 0.294 1.443
C13 -0.933 -0.211 -0.313
H14 -0.743 -0.229 -1.395
C15 -2.172 0.648 -0.057
H16 -2.325 0.797 1.017
H17 -2.107 1.627 -0.532
H18 -3.061 0.150 -0.452

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 S9 H10 C11 H12 C13 H14 C15 H16 H17 H18
C11.09321.09051.09503.90864.23894.70604.22783.14164.26592.53942.70111.52932.14932.49952.82203.46462.6597
H21.09321.76561.76824.71564.93785.61154.89674.10805.29763.48783.71042.16732.49932.71013.15523.68092.4080
H31.09051.76561.76604.26784.82224.91924.51902.71943.93462.81923.05432.19292.50753.47043.83064.33603.6765
H41.09501.76821.76604.18534.37124.87834.74153.57184.47302.79312.50132.16663.06272.76502.64383.80523.0004
C53.90864.71564.26784.18531.09221.09261.09312.76132.79531.52392.15572.55832.82082.96593.21442.70444.0073
H64.23894.93784.82224.37121.09221.76061.77893.74163.76982.17222.47552.86813.28732.68422.73362.22853.7374
H74.70605.61154.91924.87831.09261.76061.76632.90842.41642.17072.53283.50413.77813.99994.09823.75245.0626
H84.22784.89674.51904.74151.09311.77891.76632.99983.21272.16193.06462.78042.60343.23463.78282.77484.1342
S93.14164.10802.71943.57182.76133.74162.90842.99981.34451.83182.40412.72732.82864.12824.47054.48464.8903
H104.26595.29763.93464.47302.79533.76982.41643.21271.34452.38022.65413.71313.93064.90255.06255.11405.8123
C112.53943.48782.81922.79311.52392.17222.17072.16191.83182.38021.09711.53642.13812.52802.74502.85403.4748
H122.70113.71043.05432.50132.15572.47552.53283.06462.40412.65411.09712.14243.03222.81642.59553.30643.7619
C131.52932.16732.19292.16662.55832.86813.50412.78042.72733.71311.53642.14241.09911.52952.17372.19272.1633
H142.14932.49932.50753.06272.82083.28733.77812.60342.82863.93062.13813.03221.09912.14523.06212.46012.5311
C152.49952.71013.47042.76502.96592.68423.99993.23464.12824.90252.52802.81641.52952.14521.09521.09091.0930
H162.82203.15523.83062.64383.21442.73364.09823.78284.47055.06252.74502.59552.17373.06211.09521.77131.7660
H173.46463.68094.33603.80522.70442.22853.75242.77484.48465.11402.85403.30642.19272.46011.09091.77131.7610
H182.65972.40803.67653.00044.00733.73745.06264.13424.89035.81233.47483.76192.16332.53111.09301.76601.7610

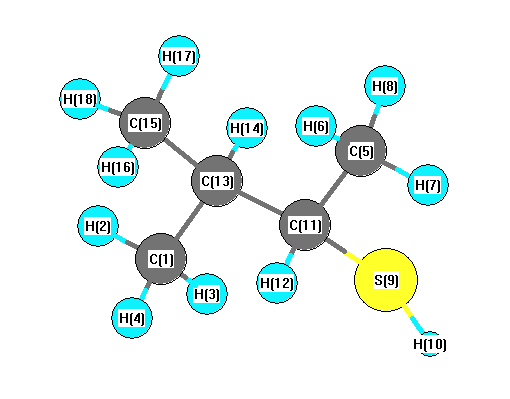
picture of 2-Butanethiol, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C13 C11 111.852 C1 C13 H14 108.608
C1 C13 C15 109.604 H2 C1 H3 107.914
H2 C1 H4 107.815 H2 C1 C13 110.362
H3 C1 H4 107.812 H3 C1 C13 112.580
H4 C1 C13 110.195 C5 C11 S9 110.410
C5 C11 H12 109.588 C5 C11 C13 113.432
H6 C5 H7 107.376 H6 C5 H8 108.984
H6 C5 C11 111.192 H7 C5 H8 107.817
H7 C5 C11 111.044 H8 C5 C11 110.311
S9 C11 H12 107.679 S9 C11 C13 107.817
H10 S9 C11 95.856 C11 C13 H14 107.283
C11 C13 C15 111.092 H12 C11 C13 107.718
C13 C15 H16 110.735 C13 C15 H17 112.531
C13 C15 H18 110.049 H14 C13 C15 108.280
H16 C15 H17 108.246 H16 C15 H18 107.615
H17 C15 H18 107.486
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability