return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7 (n-Propyl radical)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A"
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-118.100579
Energy at 298.15K 
HF Energy-117.652616
Nuclear repulsion energy75.913642
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3311 3136 14.35      
2 A 3199 3030 15.59      
3 A 3175 3007 27.51      
4 A 3169 3002 34.40      
5 A 3107 2943 18.08      
6 A 3081 2918 24.05      
7 A 3024 2864 30.66      
8 A 1546 1464 5.79      
9 A 1541 1460 9.27      
10 A 1522 1441 2.32      
11 A 1508 1428 2.59      
12 A 1447 1371 5.38      
13 A 1393 1319 0.84      
14 A 1307 1238 0.23      
15 A 1208 1144 0.48      
16 A 1121 1062 0.14      
17 A 1084 1026 1.13      
18 A 943 893 2.10      
19 A 913 865 0.33      
20 A 772 731 1.42      
21 A 470 445 51.27      
22 A 374 354 6.69      
23 A 261 247 0.18      
24 A 138 130 0.53      

Unscaled Zero Point Vibrational Energy (zpe) 19805.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18757.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.09209 0.30050 0.26085

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.297 -0.295 -0.040
C2 0.076 0.557 0.058
C3 -1.220 -0.246 -0.041
H4 -2.095 0.403 0.033
H5 -1.273 -0.779 -0.994
H6 -1.281 -0.986 0.761
H7 0.080 1.110 1.011
H8 0.100 1.324 -0.724
H9 2.251 0.127 -0.328
H10 1.298 -1.295 0.376

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.49212.51783.46442.78372.78712.13522.12641.08221.0832
C21.49211.52872.17742.17062.17251.10121.09522.25052.2418
C32.51781.52871.09241.09211.09302.15382.16163.50312.7597
H43.46442.17741.09241.76841.76682.48772.49784.37033.8101
H52.78372.17061.09211.76841.76653.06852.52503.69872.9582
H62.78712.17251.09301.76681.76652.51263.07363.86062.6264
H72.13521.10122.15382.48773.06852.51261.74752.73382.7704
H82.12641.09522.16162.49782.52503.07361.74752.49363.0828
H91.08222.25053.50314.37033.69873.86062.73382.49361.8509
H101.08322.24182.75973.81012.95822.62642.77043.08281.8509

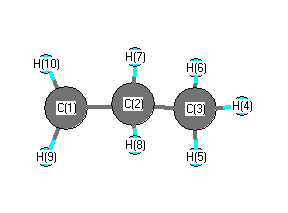
picture of n-Propyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.911 C1 C2 H7 109.931
C1 C2 H8 109.594 C2 C1 H9 121.085
C2 C1 H10 120.208 C2 C3 H4 111.256
C2 C3 H5 110.734 C2 C3 H6 110.830
C3 C2 H7 108.877 C3 C2 H8 109.830
H4 C3 H5 108.104 H4 C3 H6 107.890
H5 C3 H6 107.895 H7 C2 H8 105.425
H9 C1 H10 117.466
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability